Current Issue

2026, Volume 40,  Issue 11

Cover
Cover
2026, 40(11)
PDF (7)
Abstract:
Contents
Contents
2026, 40(11): 1-2.
Abstract:
Calculations and Simulations in High Pressure Science
2026, 40(11): 110101. doi: 10.11858/gywlxb.20261193
Abstract:
Crystal Structure Prediction at High Pressure: Methodological Development, Data-Driven Acceleration, and Exploration of Matter under Extreme Conditions
WANG Zhenyu, LI Honglin, LUO Xiaoshan, ZHANG Wei, LYU Jian, LIU Hanyu, WANG Yanchao
2026, 40(11): 110102. doi: 10.11858/gywlxb.20261158
Abstract:

High pressure can profoundly alter the atomic arrangements, electronic structures, and chemical bonding of condensed matter, providing an important means of tuning material structures and properties and expanding the accessible materials space. Crystal structure prediction (CSP) enables the prediction of stable crystal structures under different pressure conditions based solely on chemical composition, and has become a key theoretical approach for discovering novel high-pressure phases and understanding matter under extreme condition. This review focuses on the global optimization problem on high-dimensional potential-energy and enthalpy landscapes under pressure constraints. We systematically review conventional crystal structure prediction methods, as well as recent applications of machine-learning interatomic potentials and crystal generative models to high-pressure CSP. Representative advances in theory-guided high-pressure materials discovery are further summarized across experimentally accessible and ultrahigh-pressure regimes, including high-temperature superconductivity in hydride compounds and boron-carbon systems, as well as materials relevant to the interiors of the Earth and ice giant planets. Finally, current challenges and future perspectives in high-pressure crystal structure prediction are discussed.

Simulation Study on the Physical Properties of Ultrahigh-Pressure Hydrous Magnesium Silicate Phases at High Temperatures and Pressures: Exploring the Possibility of Water Presence in the Deep Interior in Terrestrial Super-Earths
REN Hang, ZHANG Wenqi, LIU Lei
2026, 40(11): 110103. doi: 10.11858/gywlxb.20261061
HTML PDF (68)
Abstract:

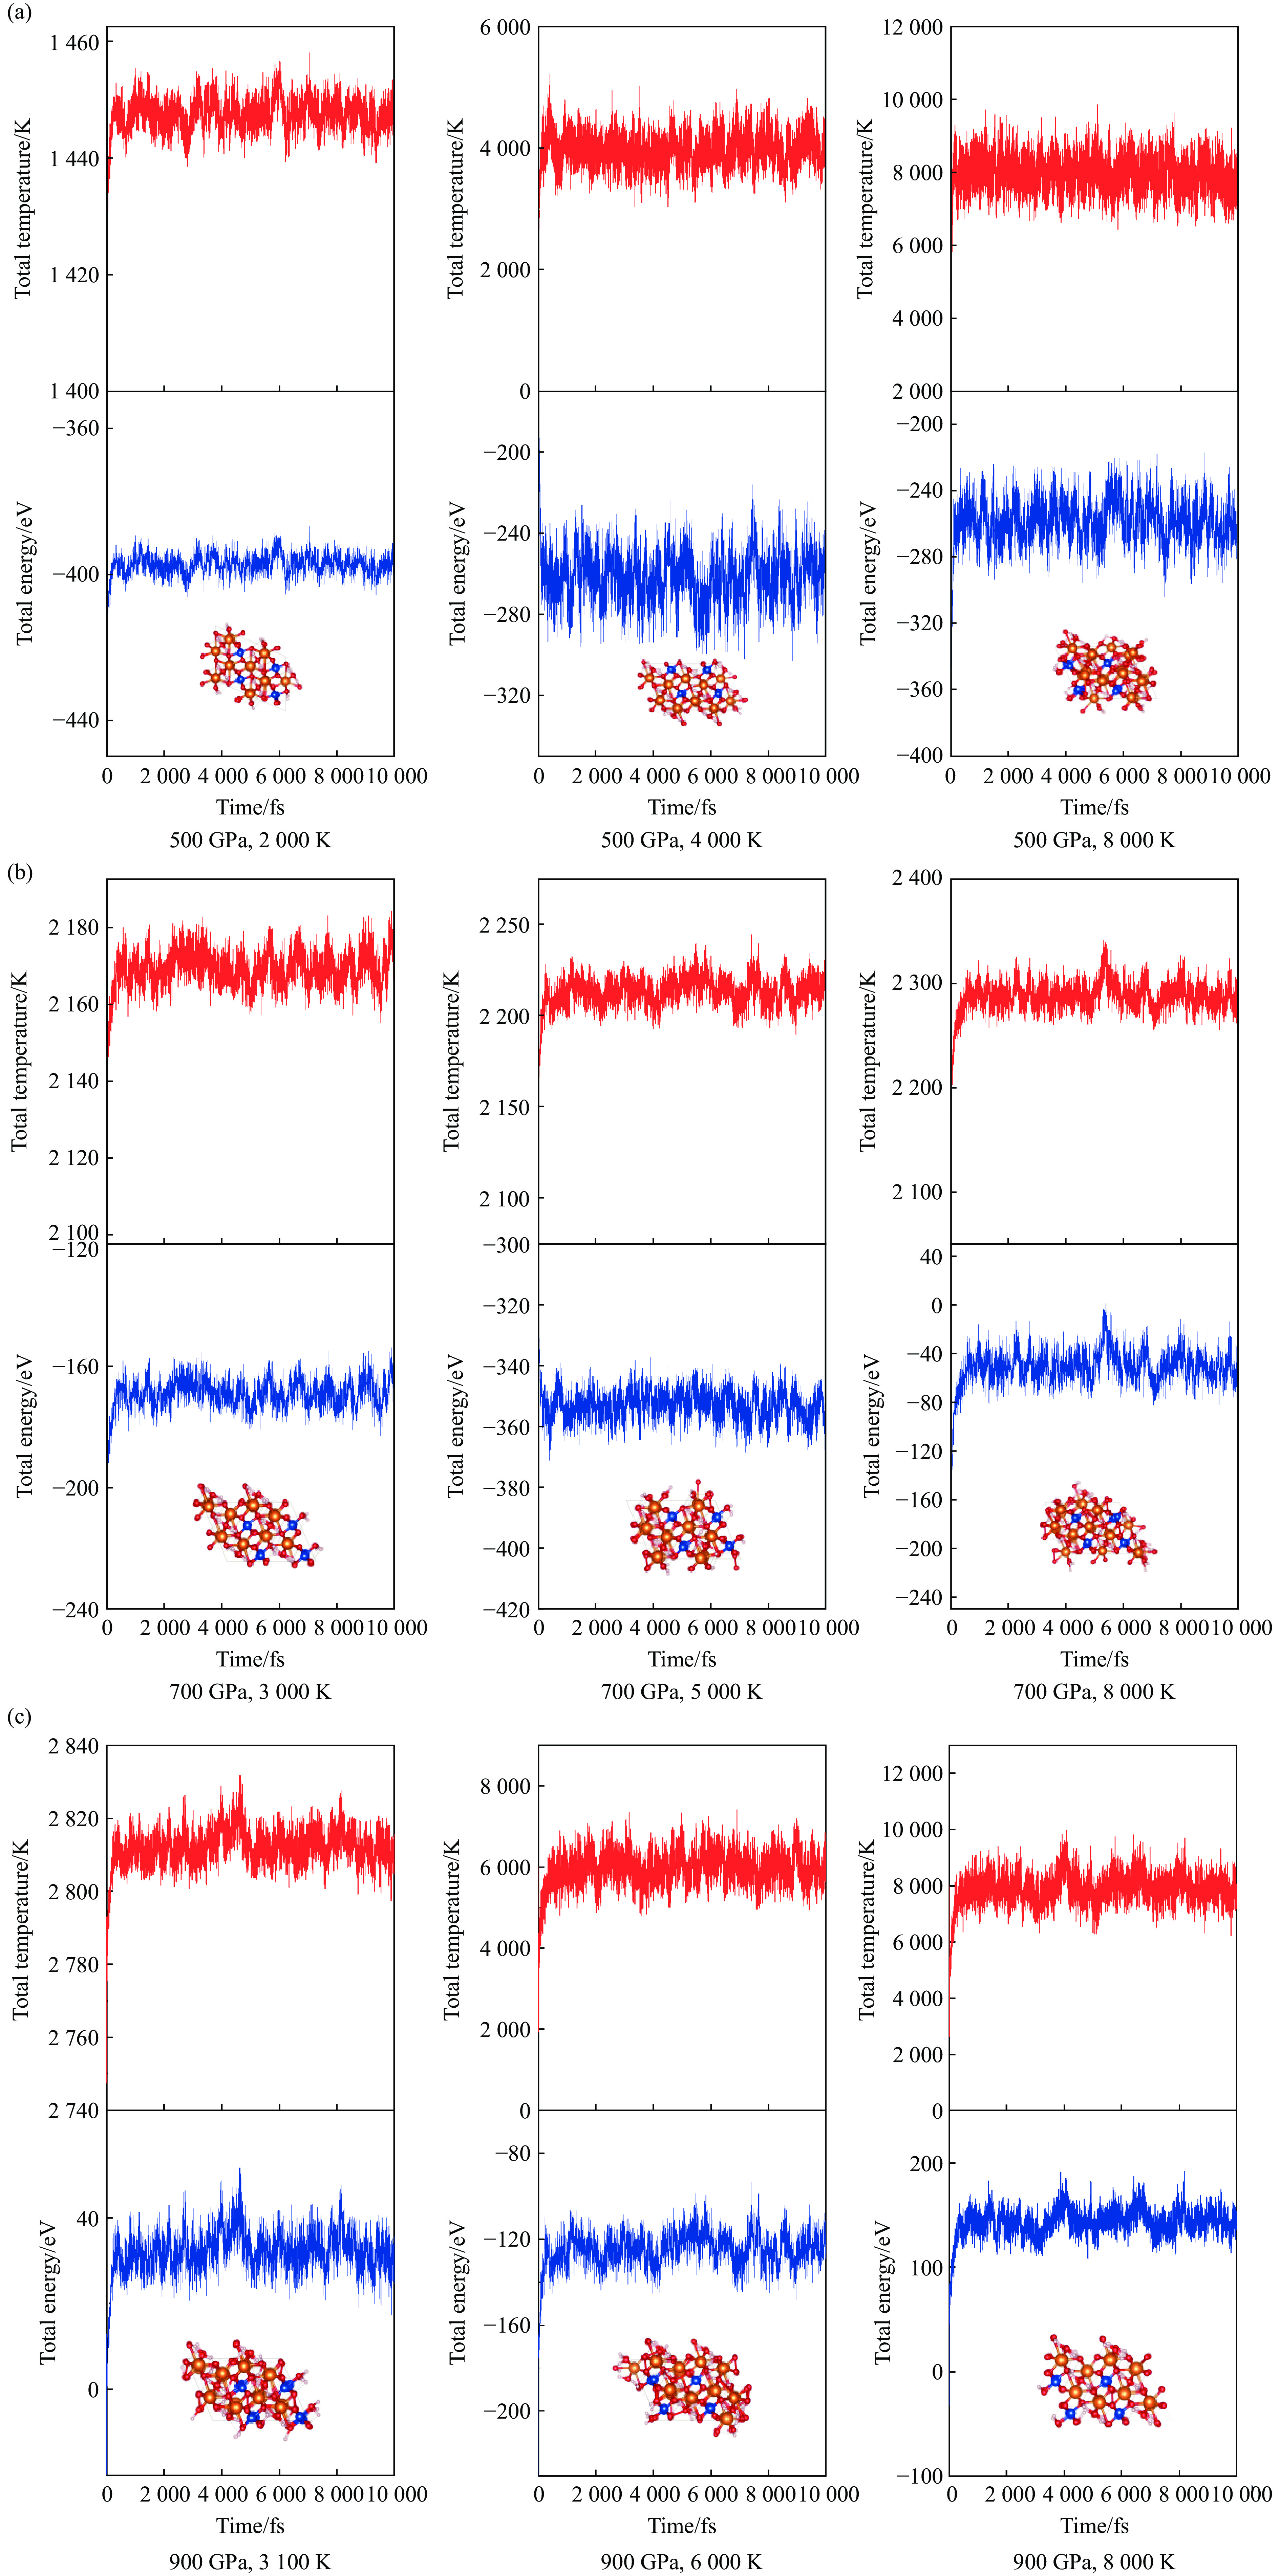
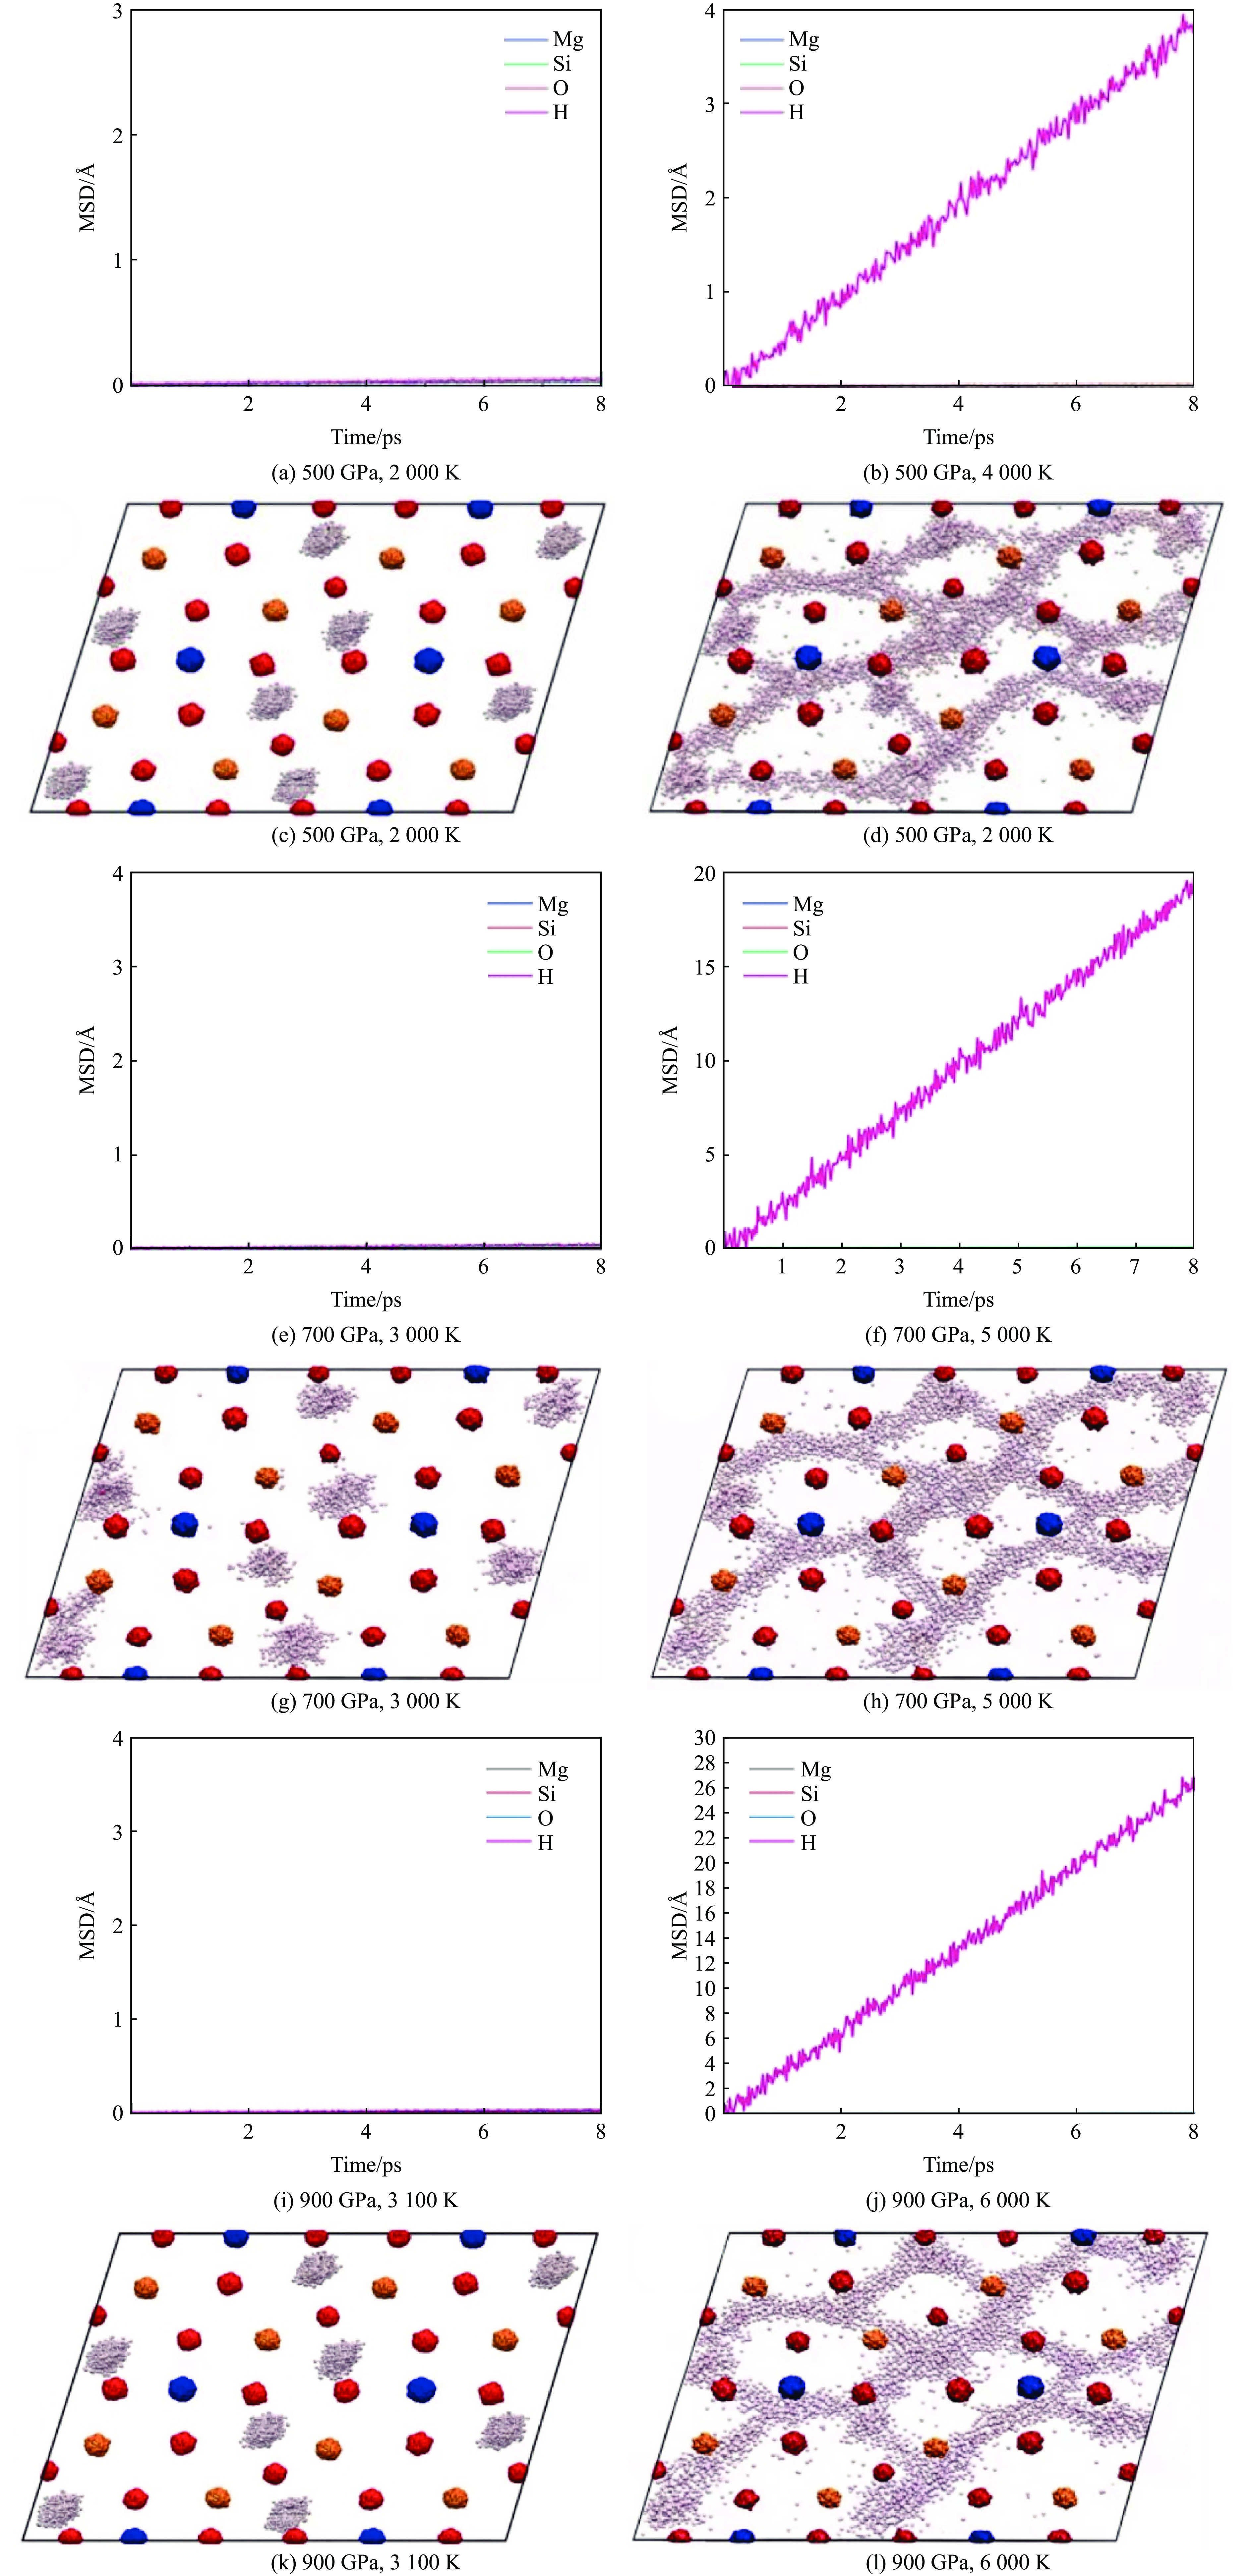
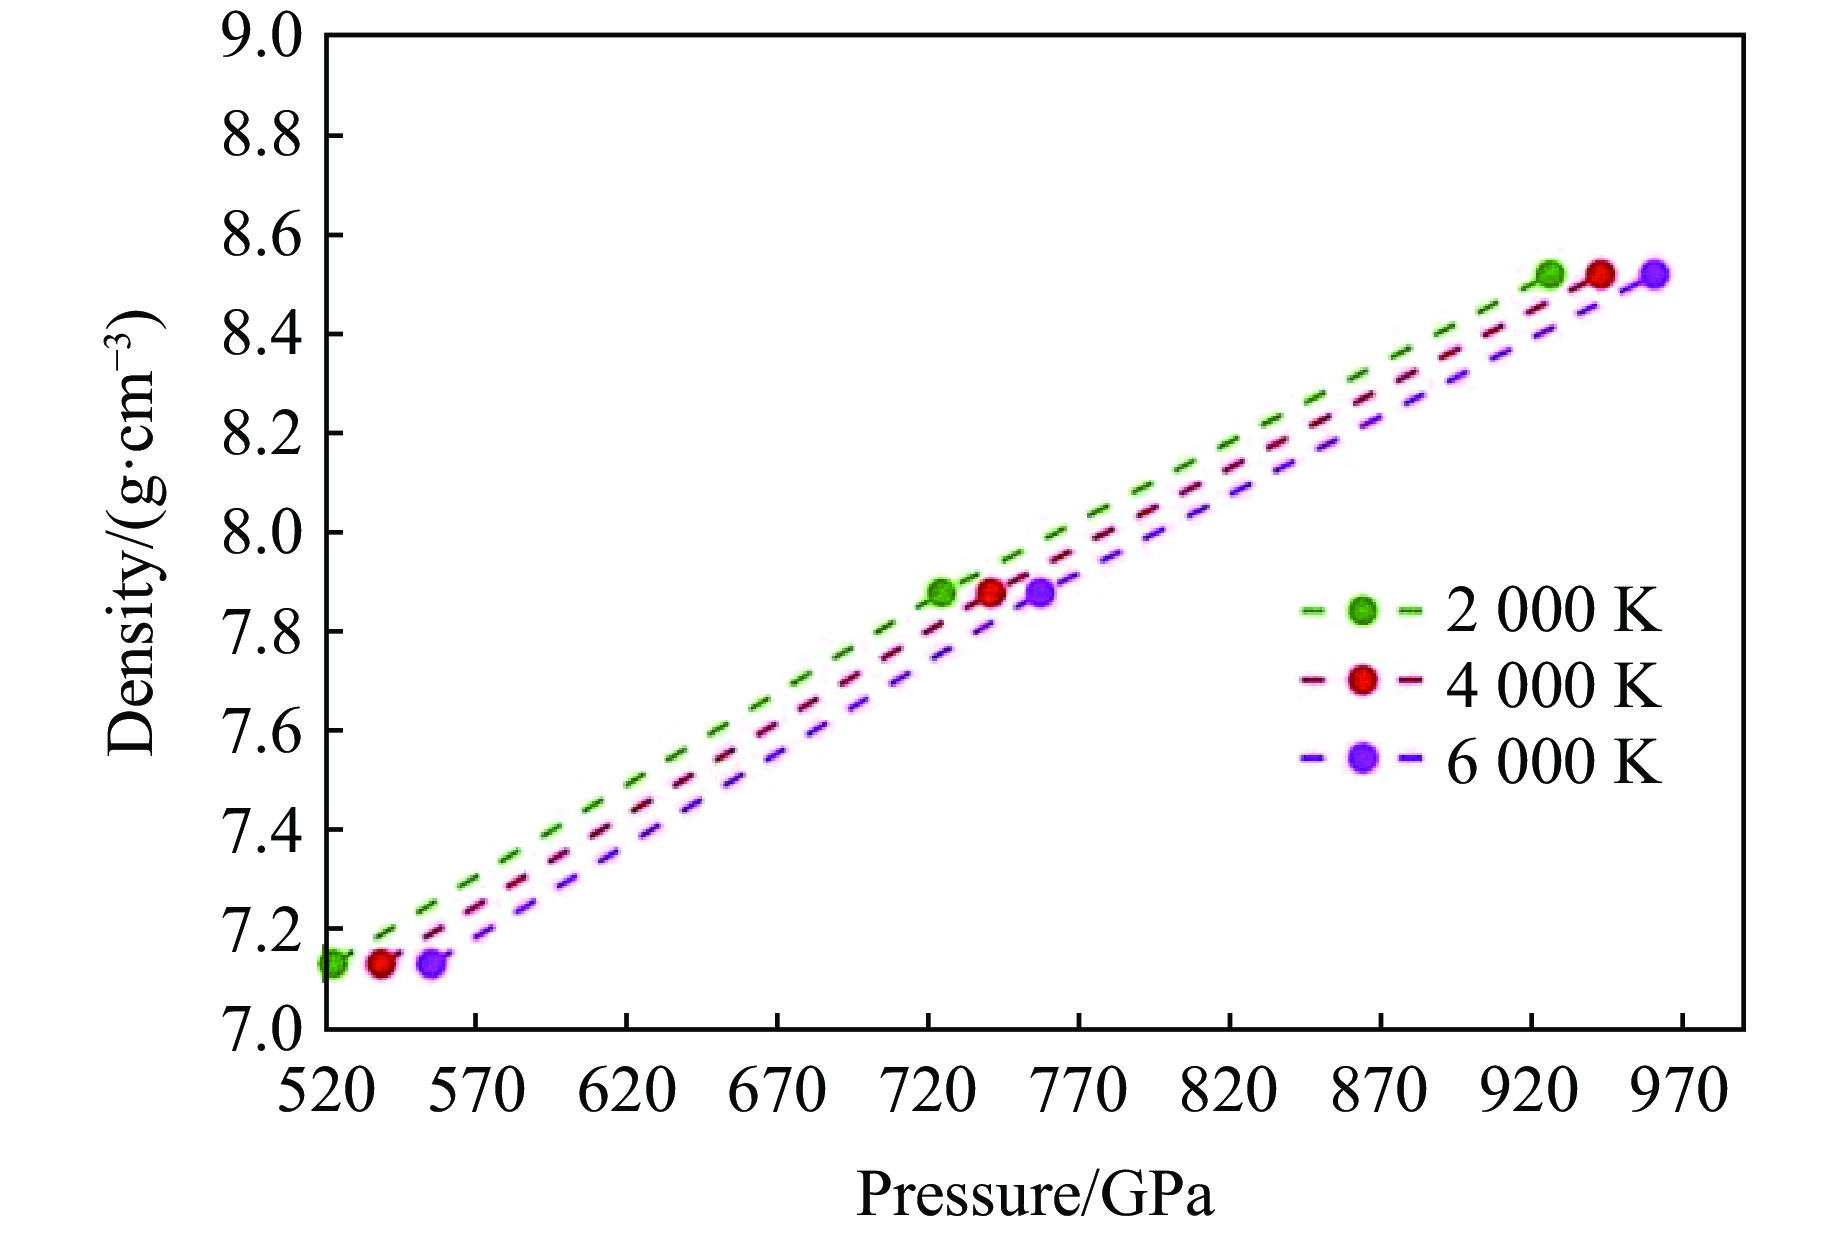
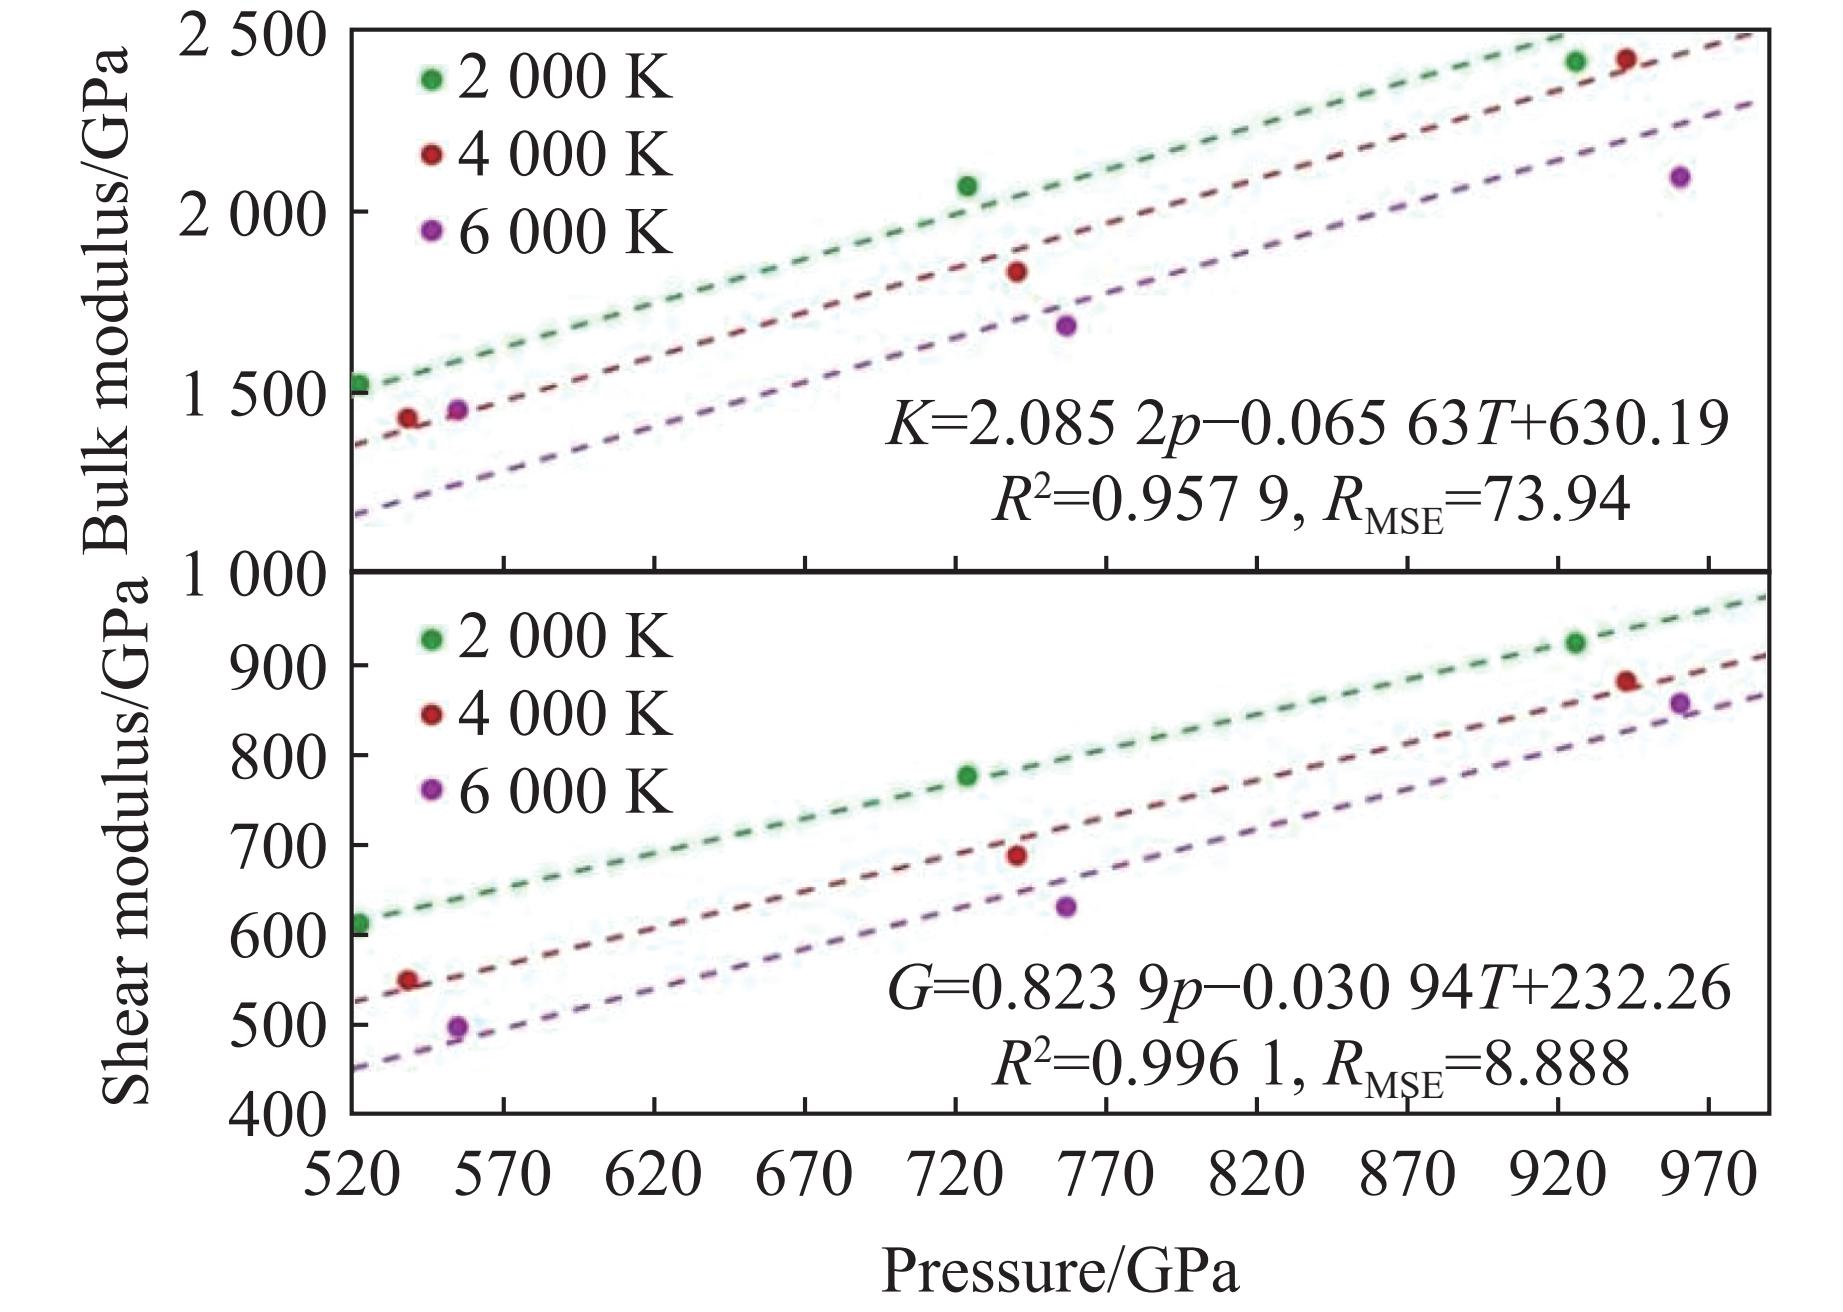
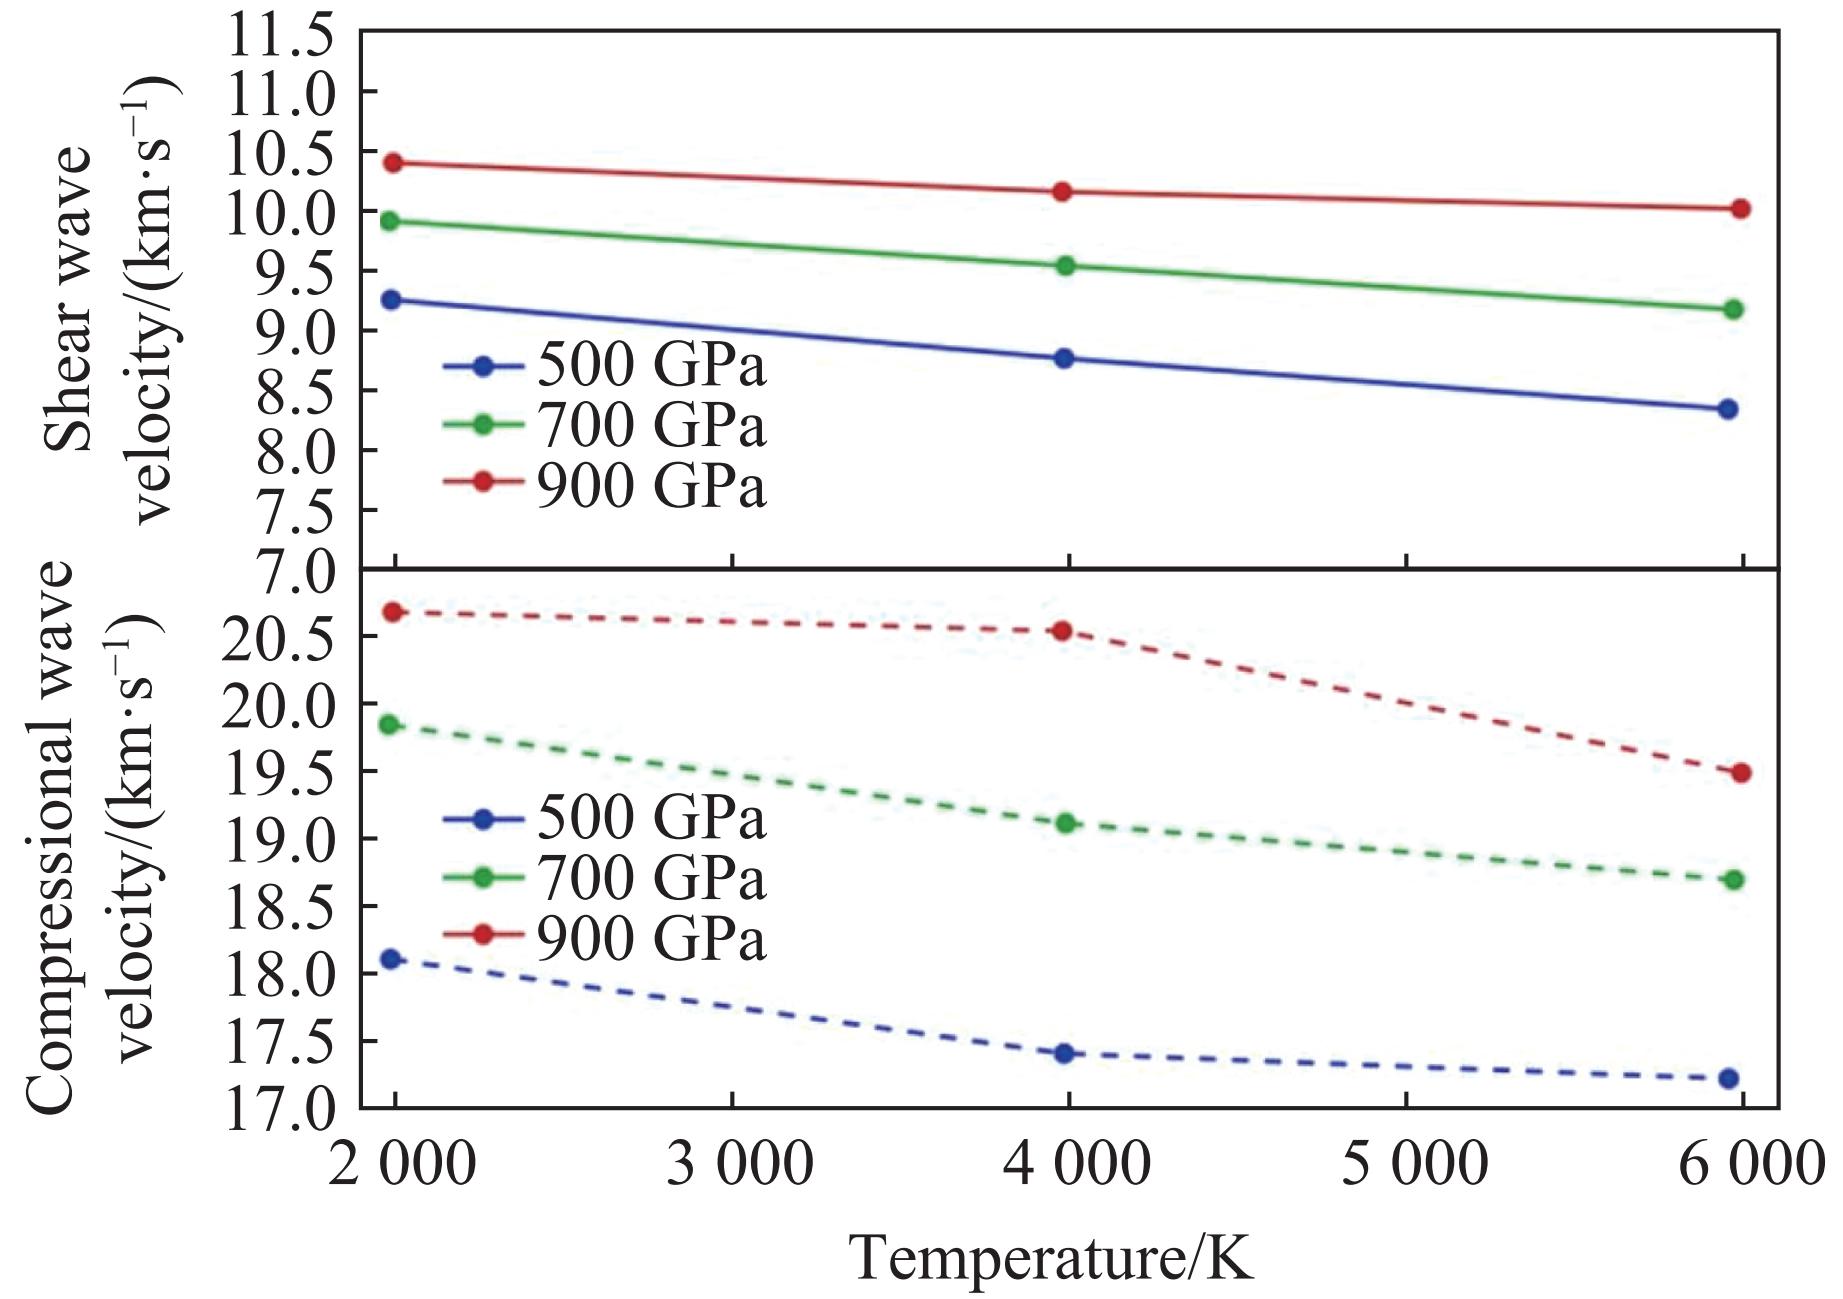
Hydrous magnesium silicate is considered as a potential water-rich reservoir in the early Earth’s interior. Investigating its behavior under extreme high-temperature and high-pressure conditions is crucial for understanding the internal structure models and water presence potential of super-Earths. By using first-principles molecular dynamics simulations, this study systematically investigates the stability and elastic properties of β-Mg2SiO5H2 within the pressure range of 500–900 GPa and temperature range of 2000–6000 K. The results indicate that the system remains thermodynamically stable across the entire studied pressure-temperature range, with no structural phase transitions observed. Calculations of the mean square displacement reveal the transition interval for the superionic state: at 500 GPa and 2000 K, 700 GPa and 3000 K, and 900 GPa and 3100 K, all atoms remain confined within the lattice, with the system in a normal state; when the temperature increases to 4000 K at 500 GPa, 5000 K at 700 GPa, and 6000 K at 900 GPa, the mean square displacement of H atoms exhibits a linear increase, while the framework atoms (Mg, Si, O) remain localized. The proton trajectories form a diffuse network, exhibiting the characteristics of a superionic state with a “solid framework+liquid-like ions”. Simulation results show that the density of β-Mg2SiO5H2 increases linearly with pressure. The shear modulus decreases nearly linearly with increasing temperature, while the bulk modulus shows a significant reduction at 900 GPa and 6000 K, likely in response to the superionic state transition. The variations in shear and compressional wave velocity are primarily controlled by pressure and temperature, increasing with pressure and decreasing with temperature. Under high pressure (900 GPa), the superionic state leads to a notable decrease in the compressional wave velocity of β-Mg2SiO5H2, suggesting that the superionic transition induces structural softening. This study confirms that β-Mg2SiO5H2 can remain stable under the deep mantle pressure-temperature conditions of super-Earths with masses 5–8 times that of Earth and can transition to a superionic state under specific conditions. Its high water mass fraction of up to 11.4% and efficient proton transport capability have significant implications for deep water cycles and the habitability of terrestrial planets, providing key theoretical insights for understanding planetary interior dynamics.

Research Progress on Ion-Partitioning Pathways in High-Pressure Saline Water Systems
WEI Xinmiao, LI Xinyang, LIU Zhao, LI Fangfei, CUI Tian
2026, 40(11): 110104. doi: 10.11858/gywlxb.20261125
HTML PDF (23)
Abstract:

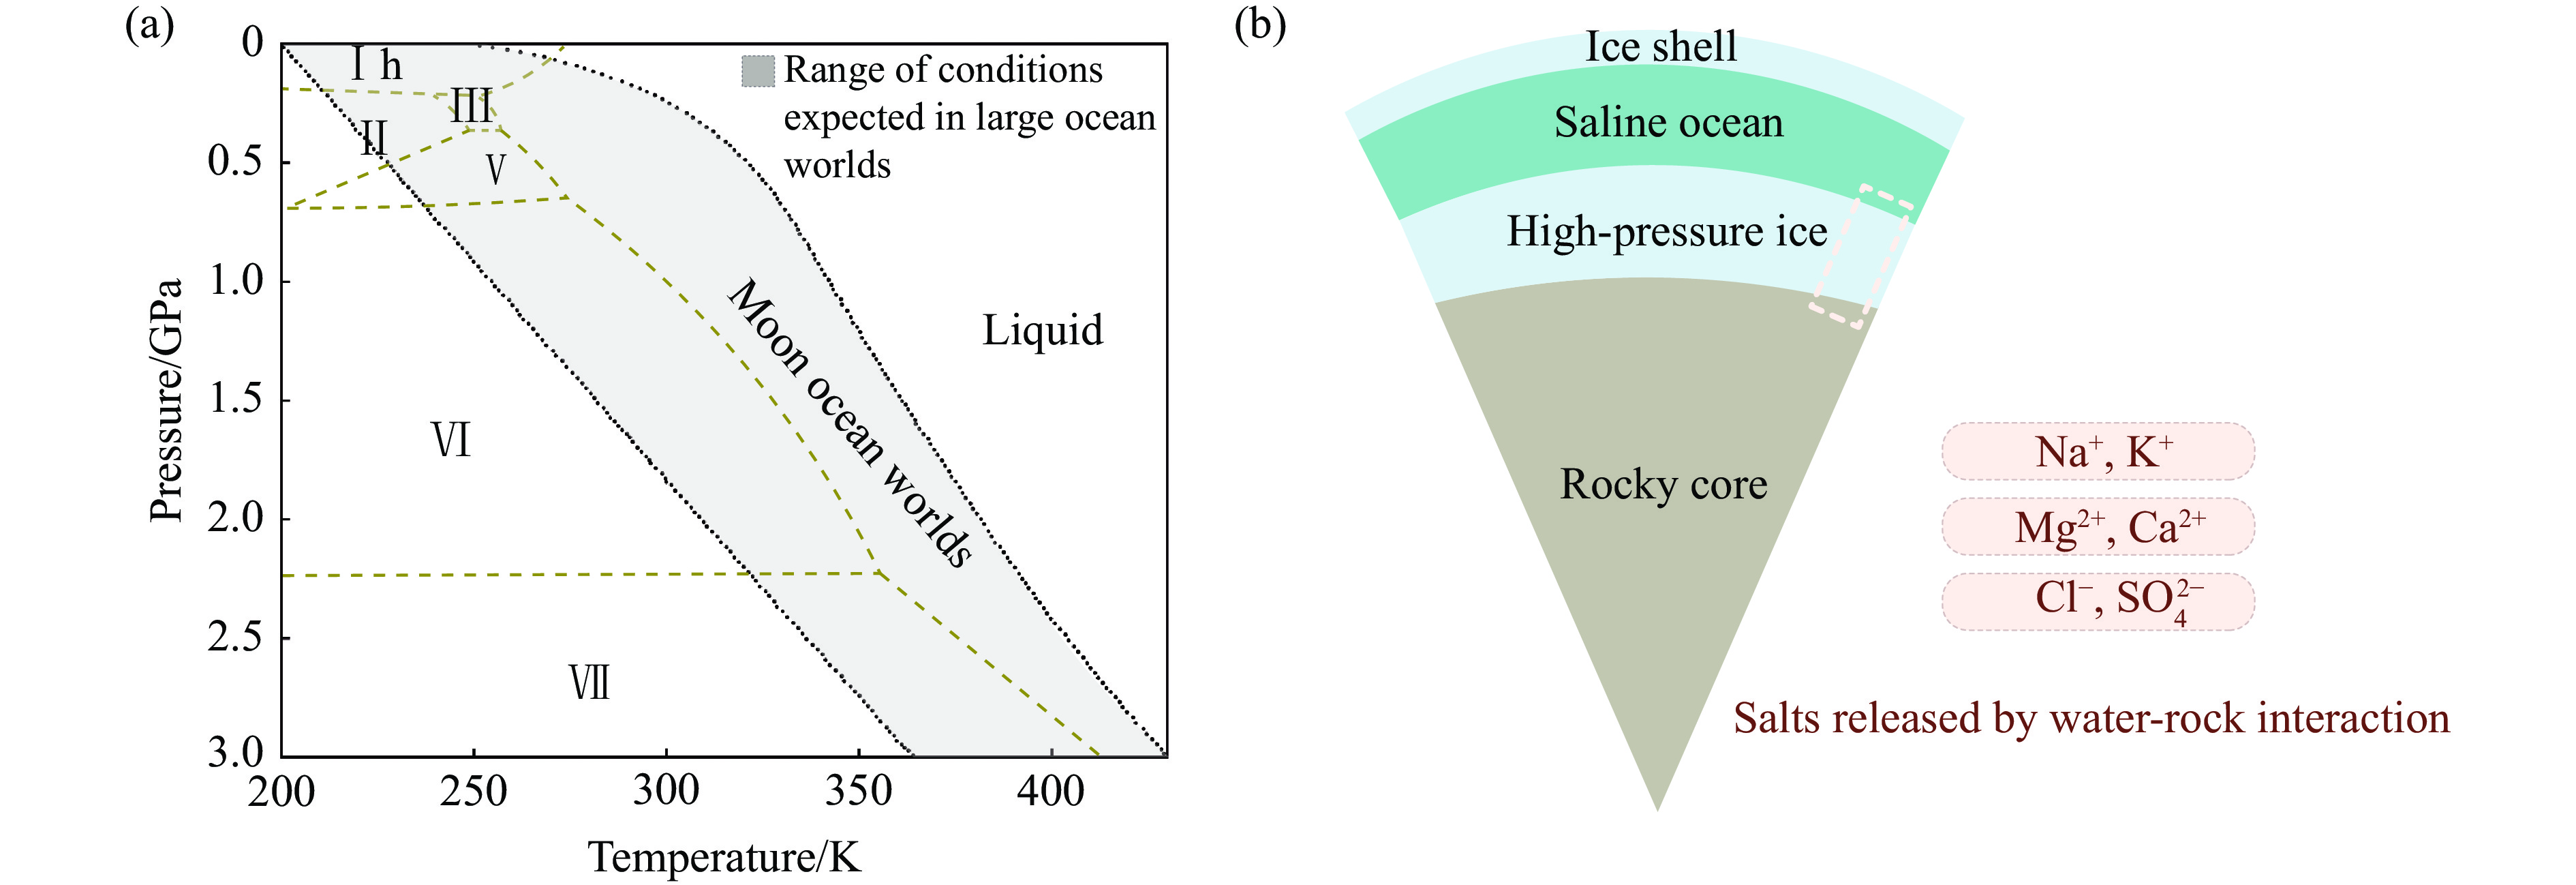
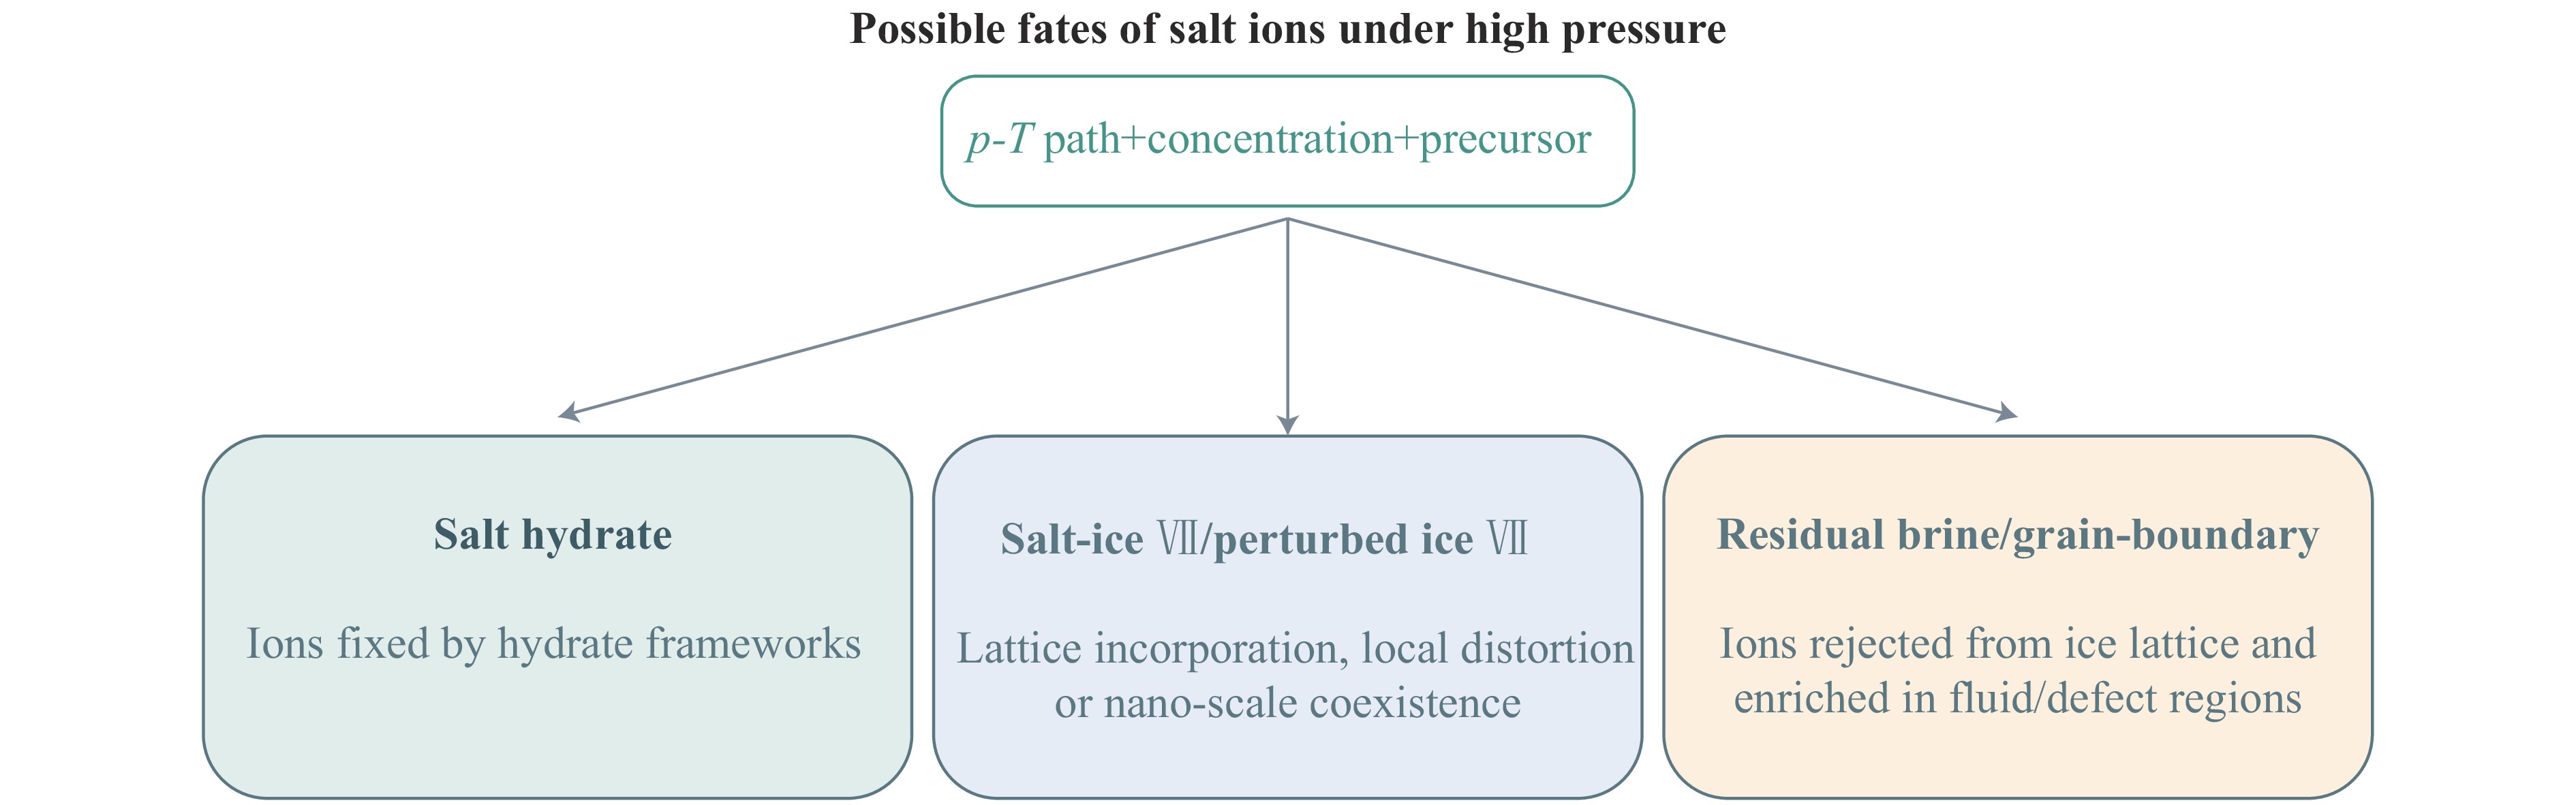
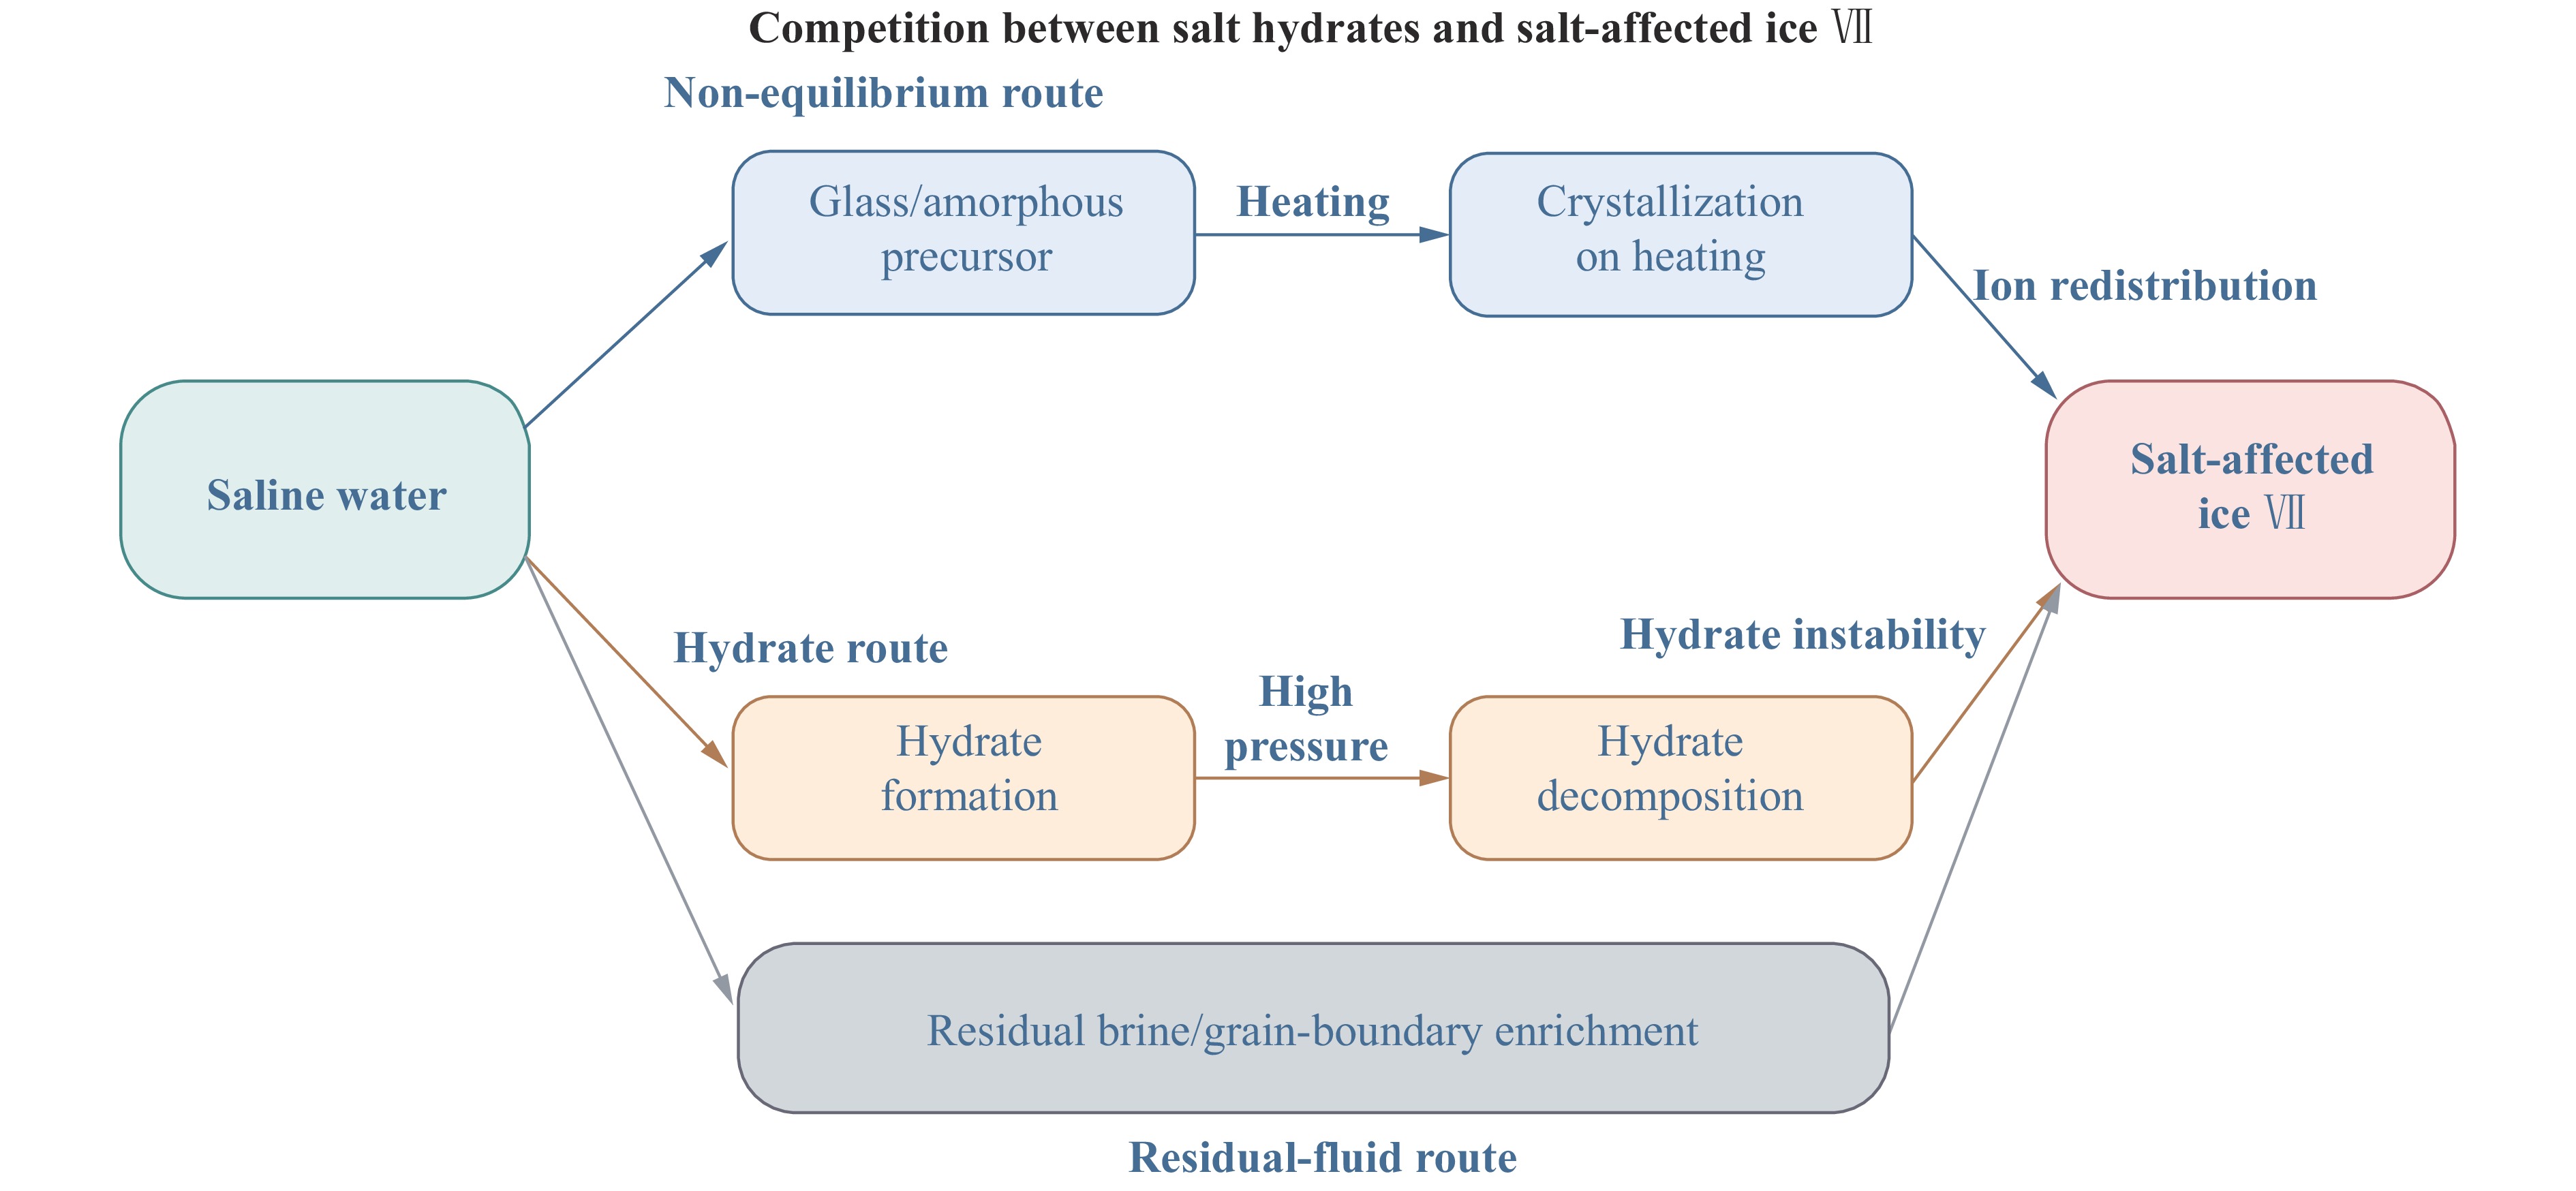
Phase-transition behavior in high-pressure saline water systems is fundamental to understanding the deep hydrospheres of icy moons, the ice mantles of water-rich planets, and high-pressure ice physics. Compared with pure H2O systems, salt ions not only modify the formation pathways of high-pressure ice phases such as ice Ⅵ and ice Ⅶ, but may also be redistributed as salt hydrates, salt-affected ice Ⅶ, or residual brines. In this review, we address the question of where salt ions reside in high-pressure aqueous systems by establishing a comparative framework based on two aspects: phase-transition pathway controls and evidence-assessment criteria. Concentration, water activity, pressure-temperature path, precursor state, and hydrate stability mainly govern the partitioning of salt ions among ice phases, hydrates, and residual fluids, whereas evidence type and evidence strength define the reliability and wording boundaries of different conclusions. On this basis, we review recent progress in representative systems including LiCl, NaCl, KCl, KBr, CaCl2, and MgCl2. Existing studies show that the LiCl system provides representative evidence for salt-affected ice Ⅶ formed through a non-equilibrium pathway; the NaCl system exhibits strong condition dependence and remains controversial with respect to extensive lattice incorporation; KCl/KBr and some NaCl systems preferentially form salt hydrates; the CaCl2 system reveals a stepwise mechanism involving hydrate formation, hydrate decomposition, and subsequent perturbation of ice Ⅶ; and the MgCl2 system indicates that the relationship between multiple hydrates and potential MgCl2-ice Ⅶ still requires confirmation by continuous in situ experiments. It should be noted that this review focuses on representative chloride and halide systems for which relatively systematic high-pressure experimental data are available, whereas sulfate, carbonate, and multicomponent salt systems are discussed only briefly in the context of planetary implications and future research directions.

Theoretical Study on Structural Stability and Superionic Phase Transition of UH5 under High Pressure
DING Yuqing, JIA Xixi, ZHANG Wenhui, WANG Hui
2026, 40(11): 110105. doi: 10.11858/gywlxb.20251224
HTML PDF (94)
Abstract:

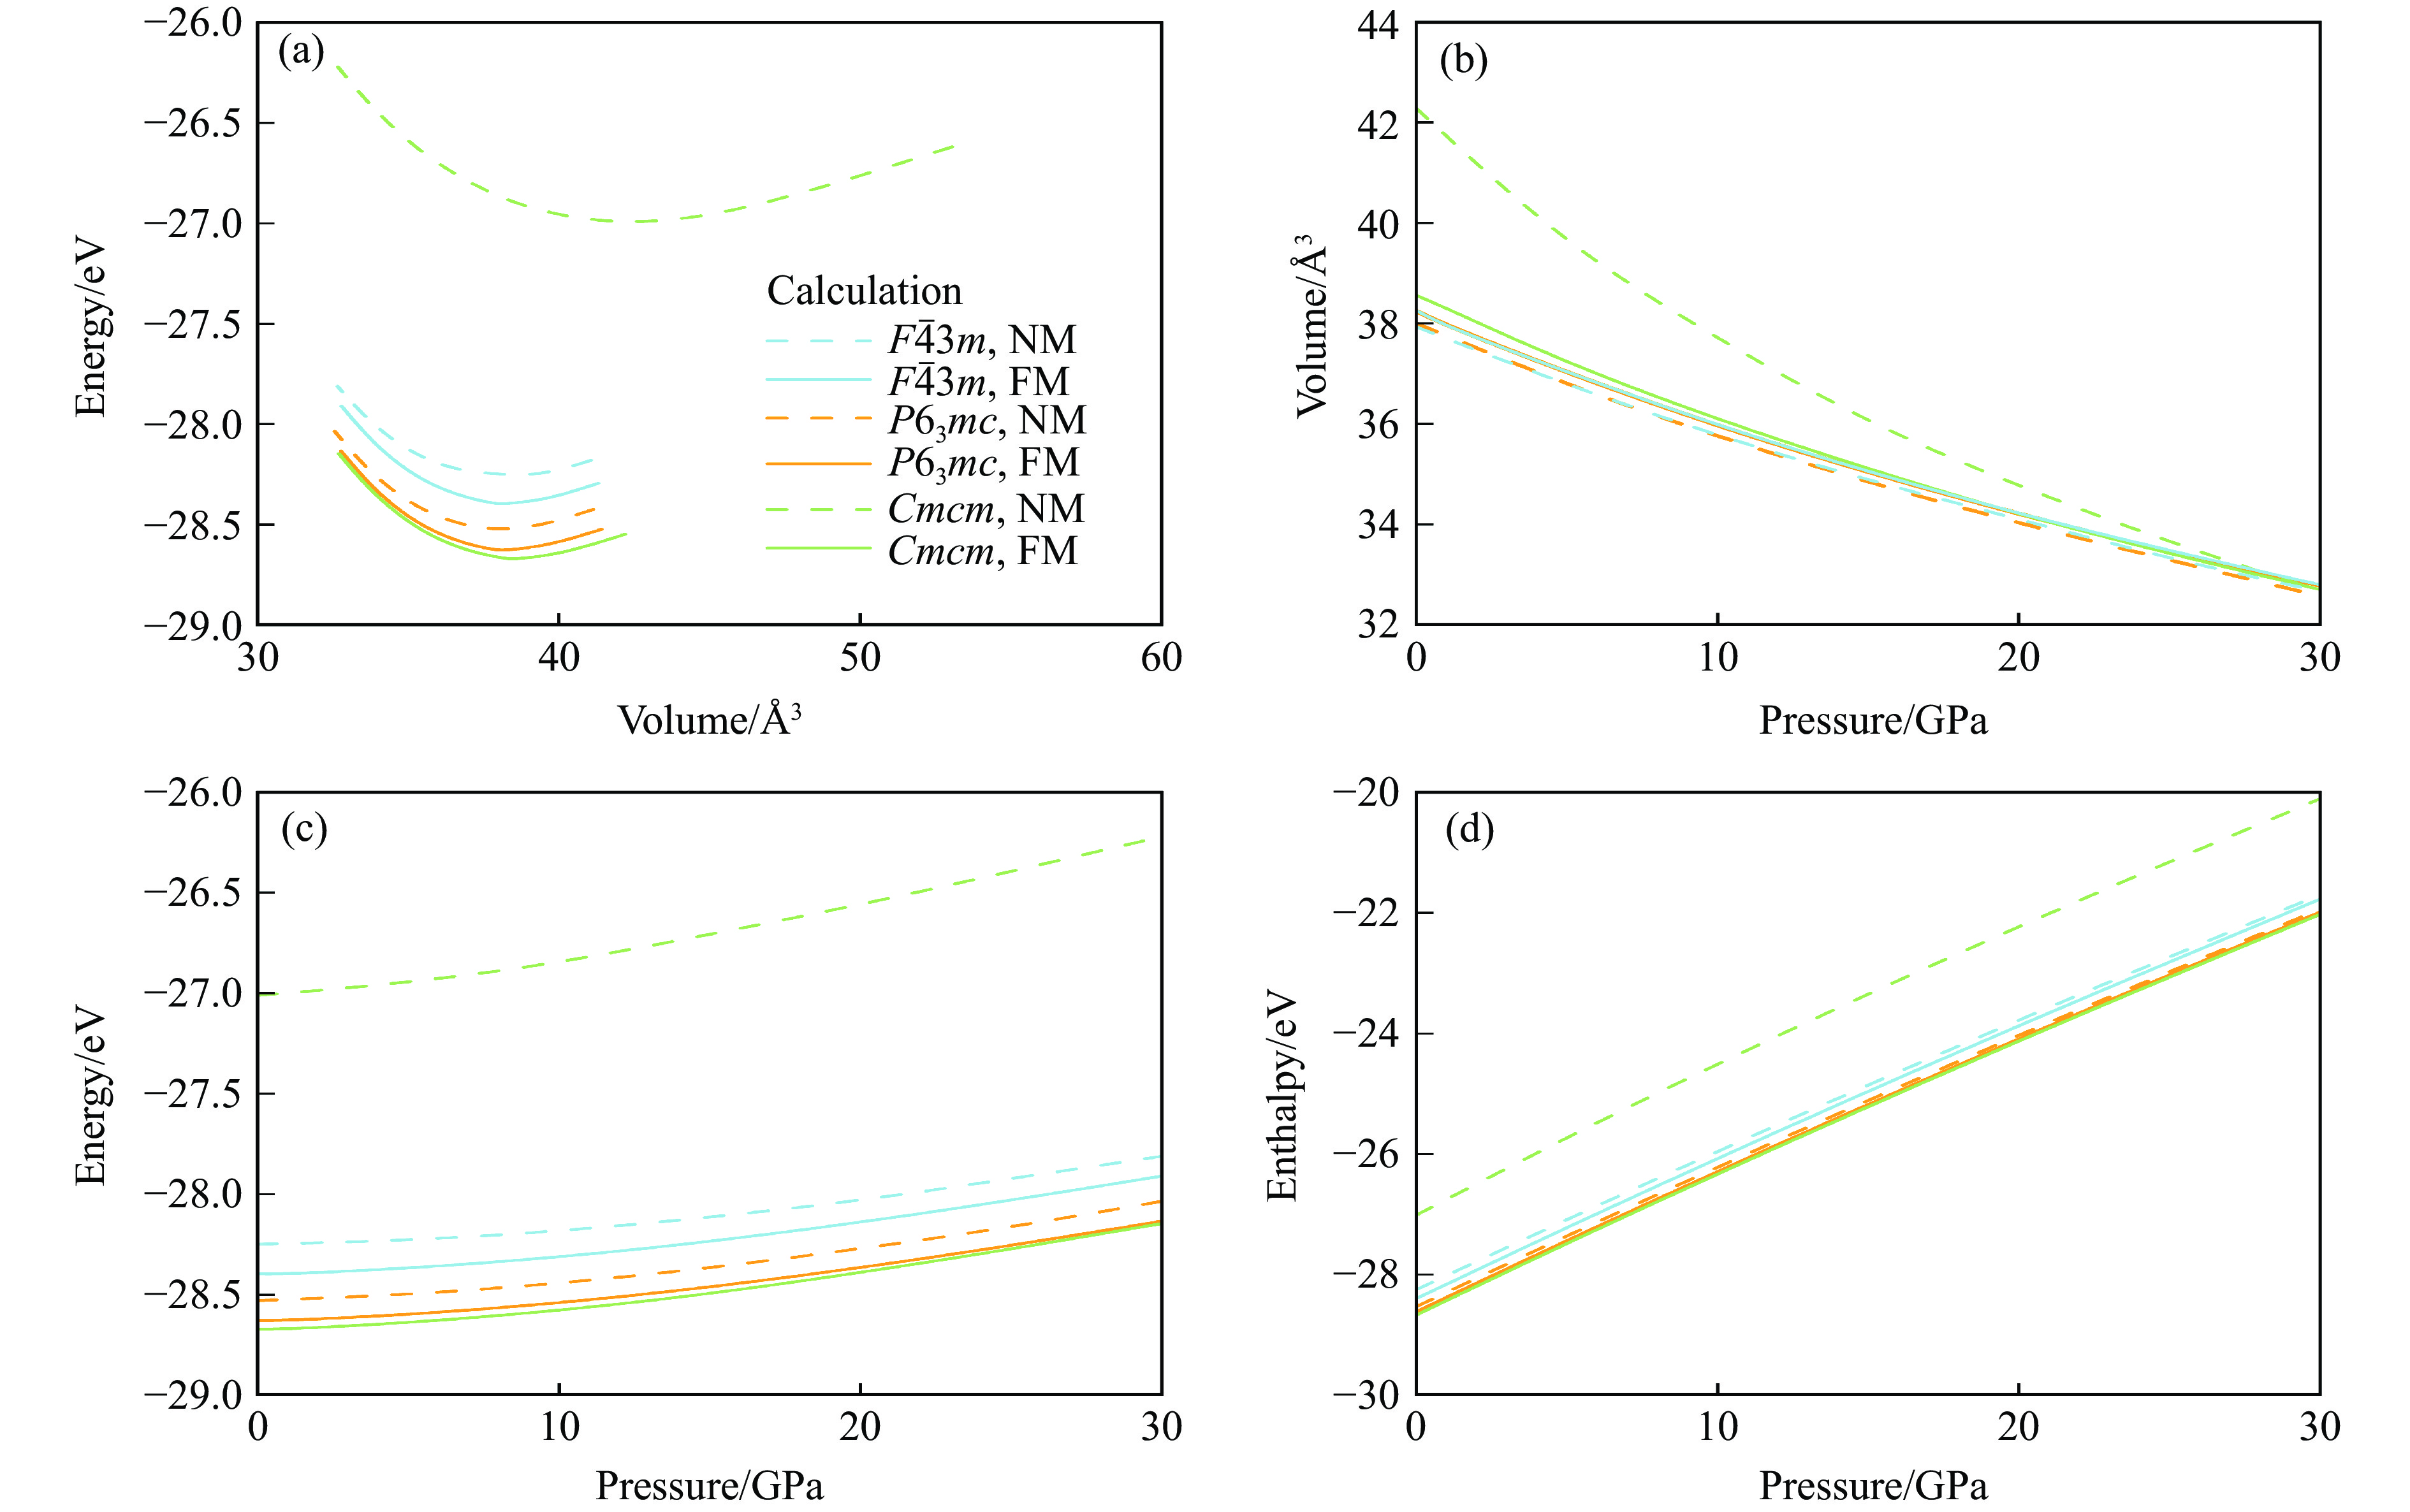
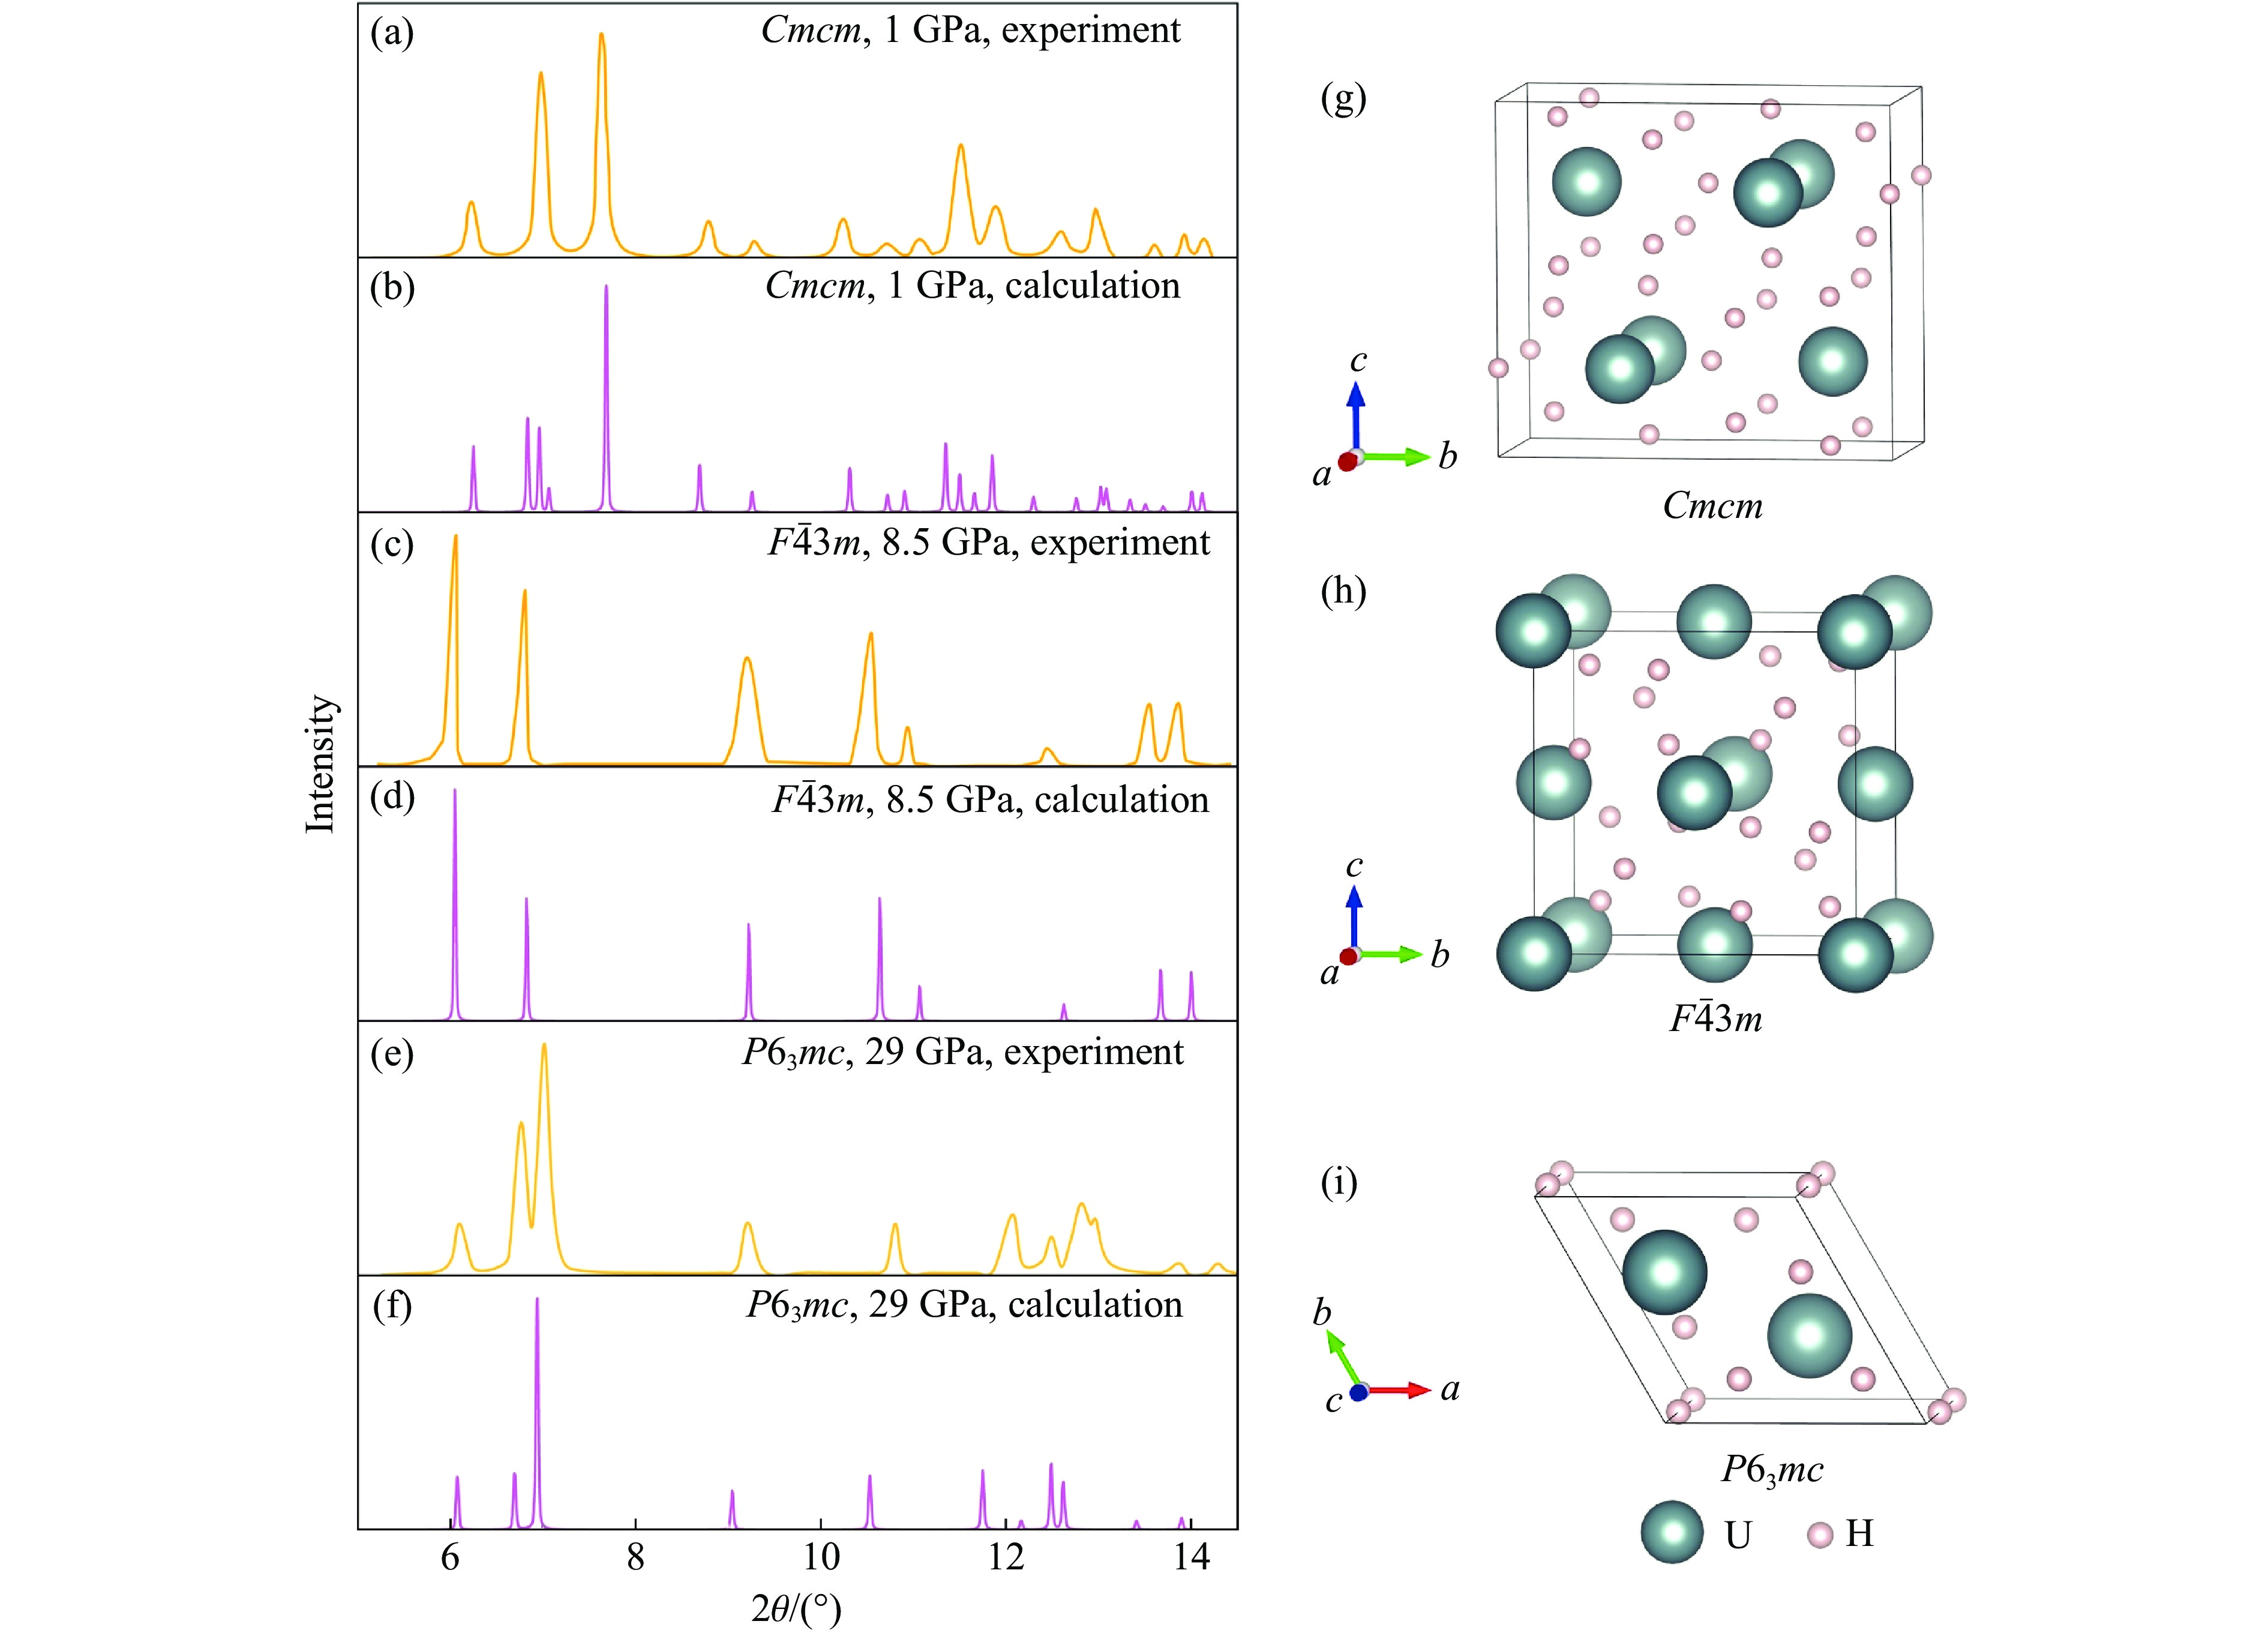
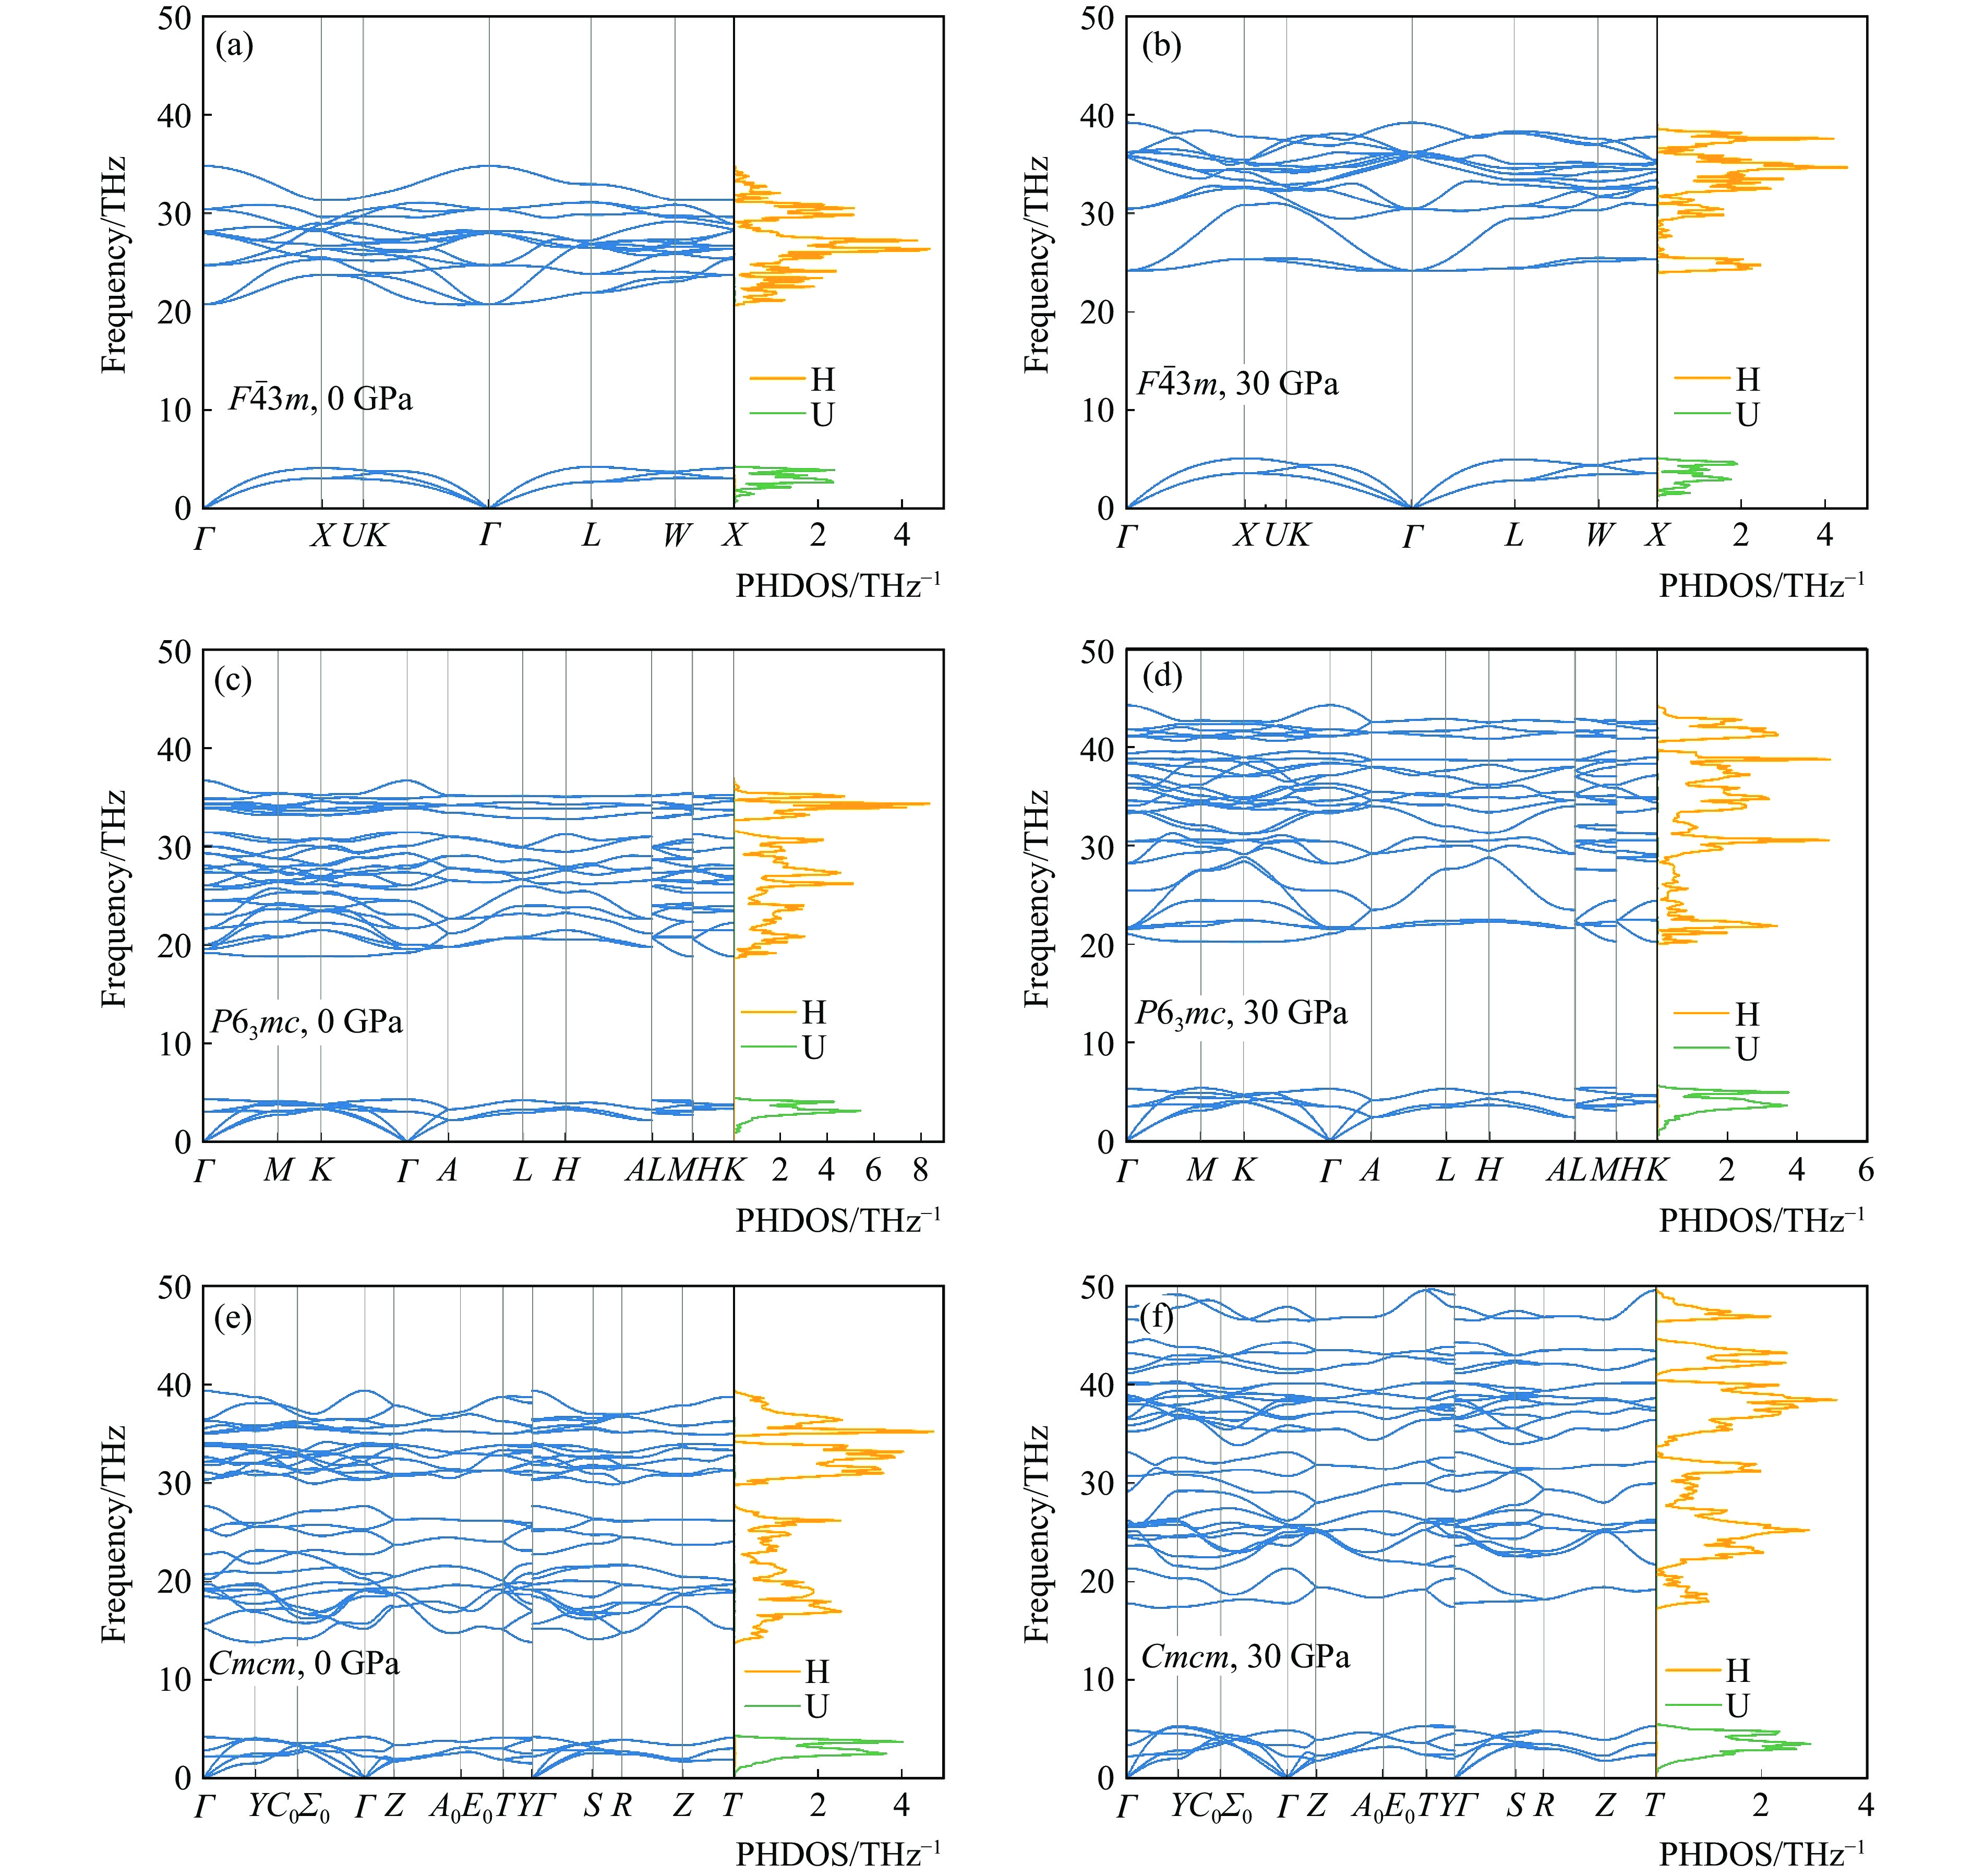
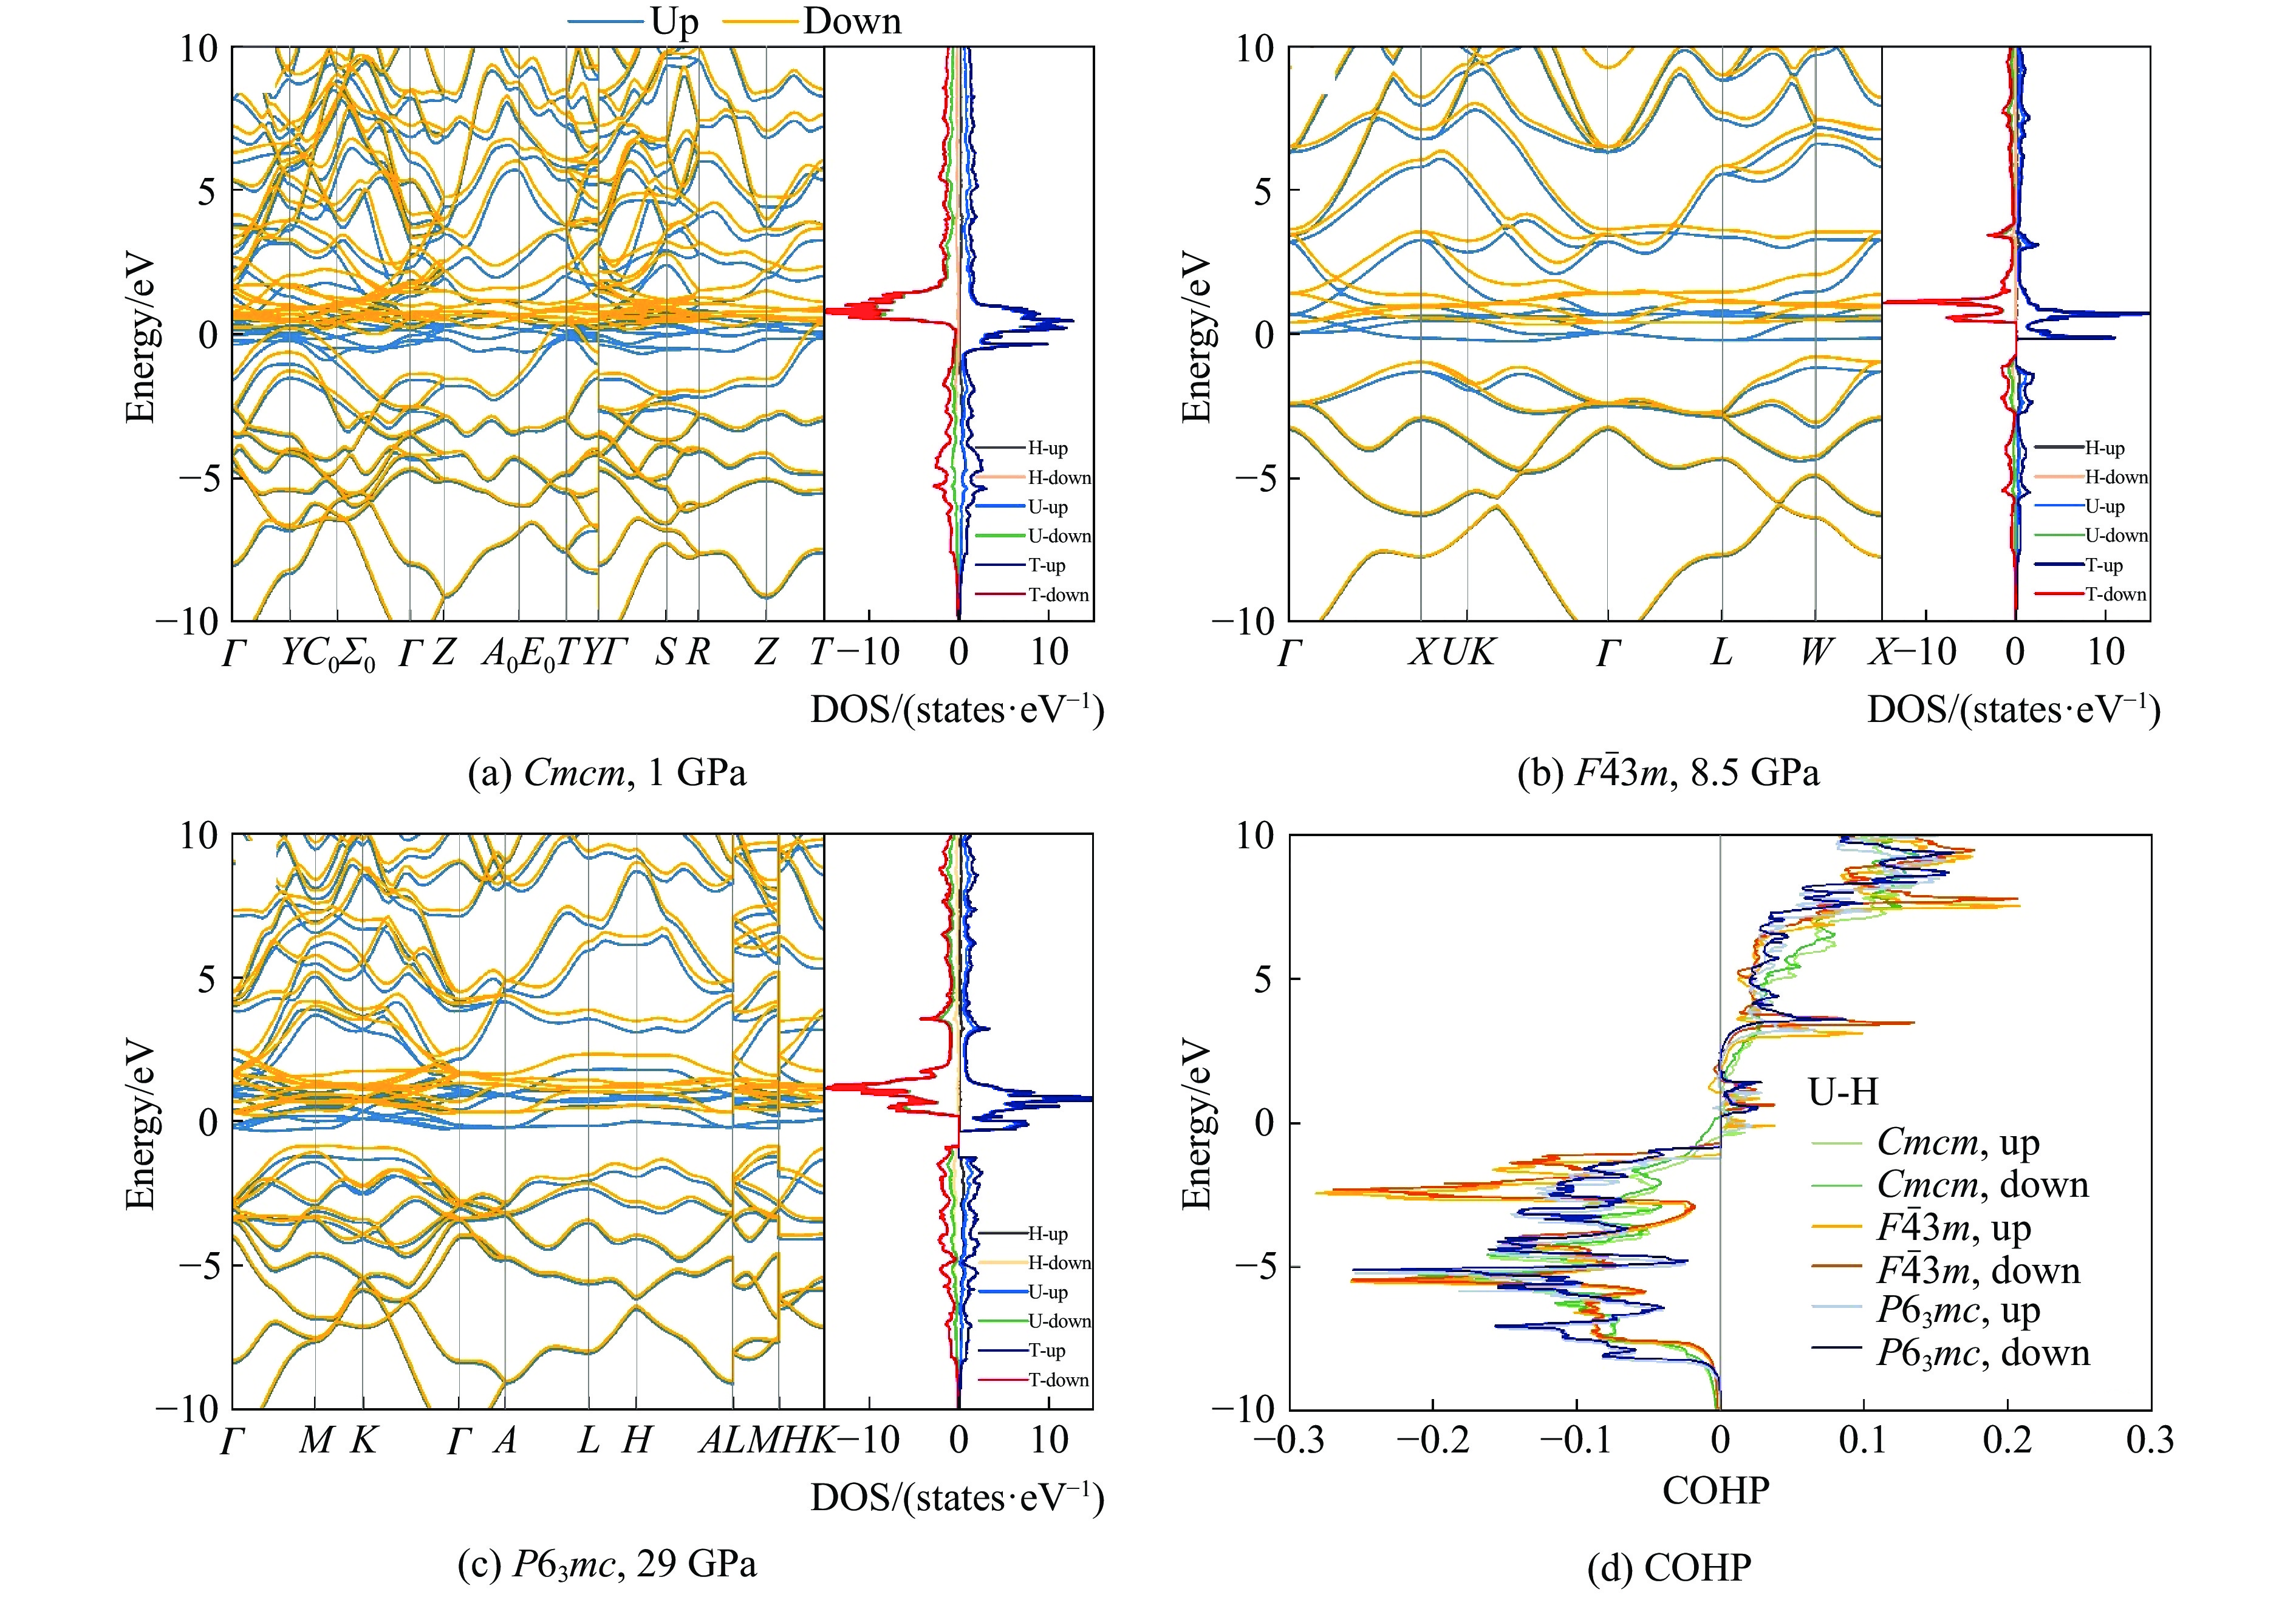
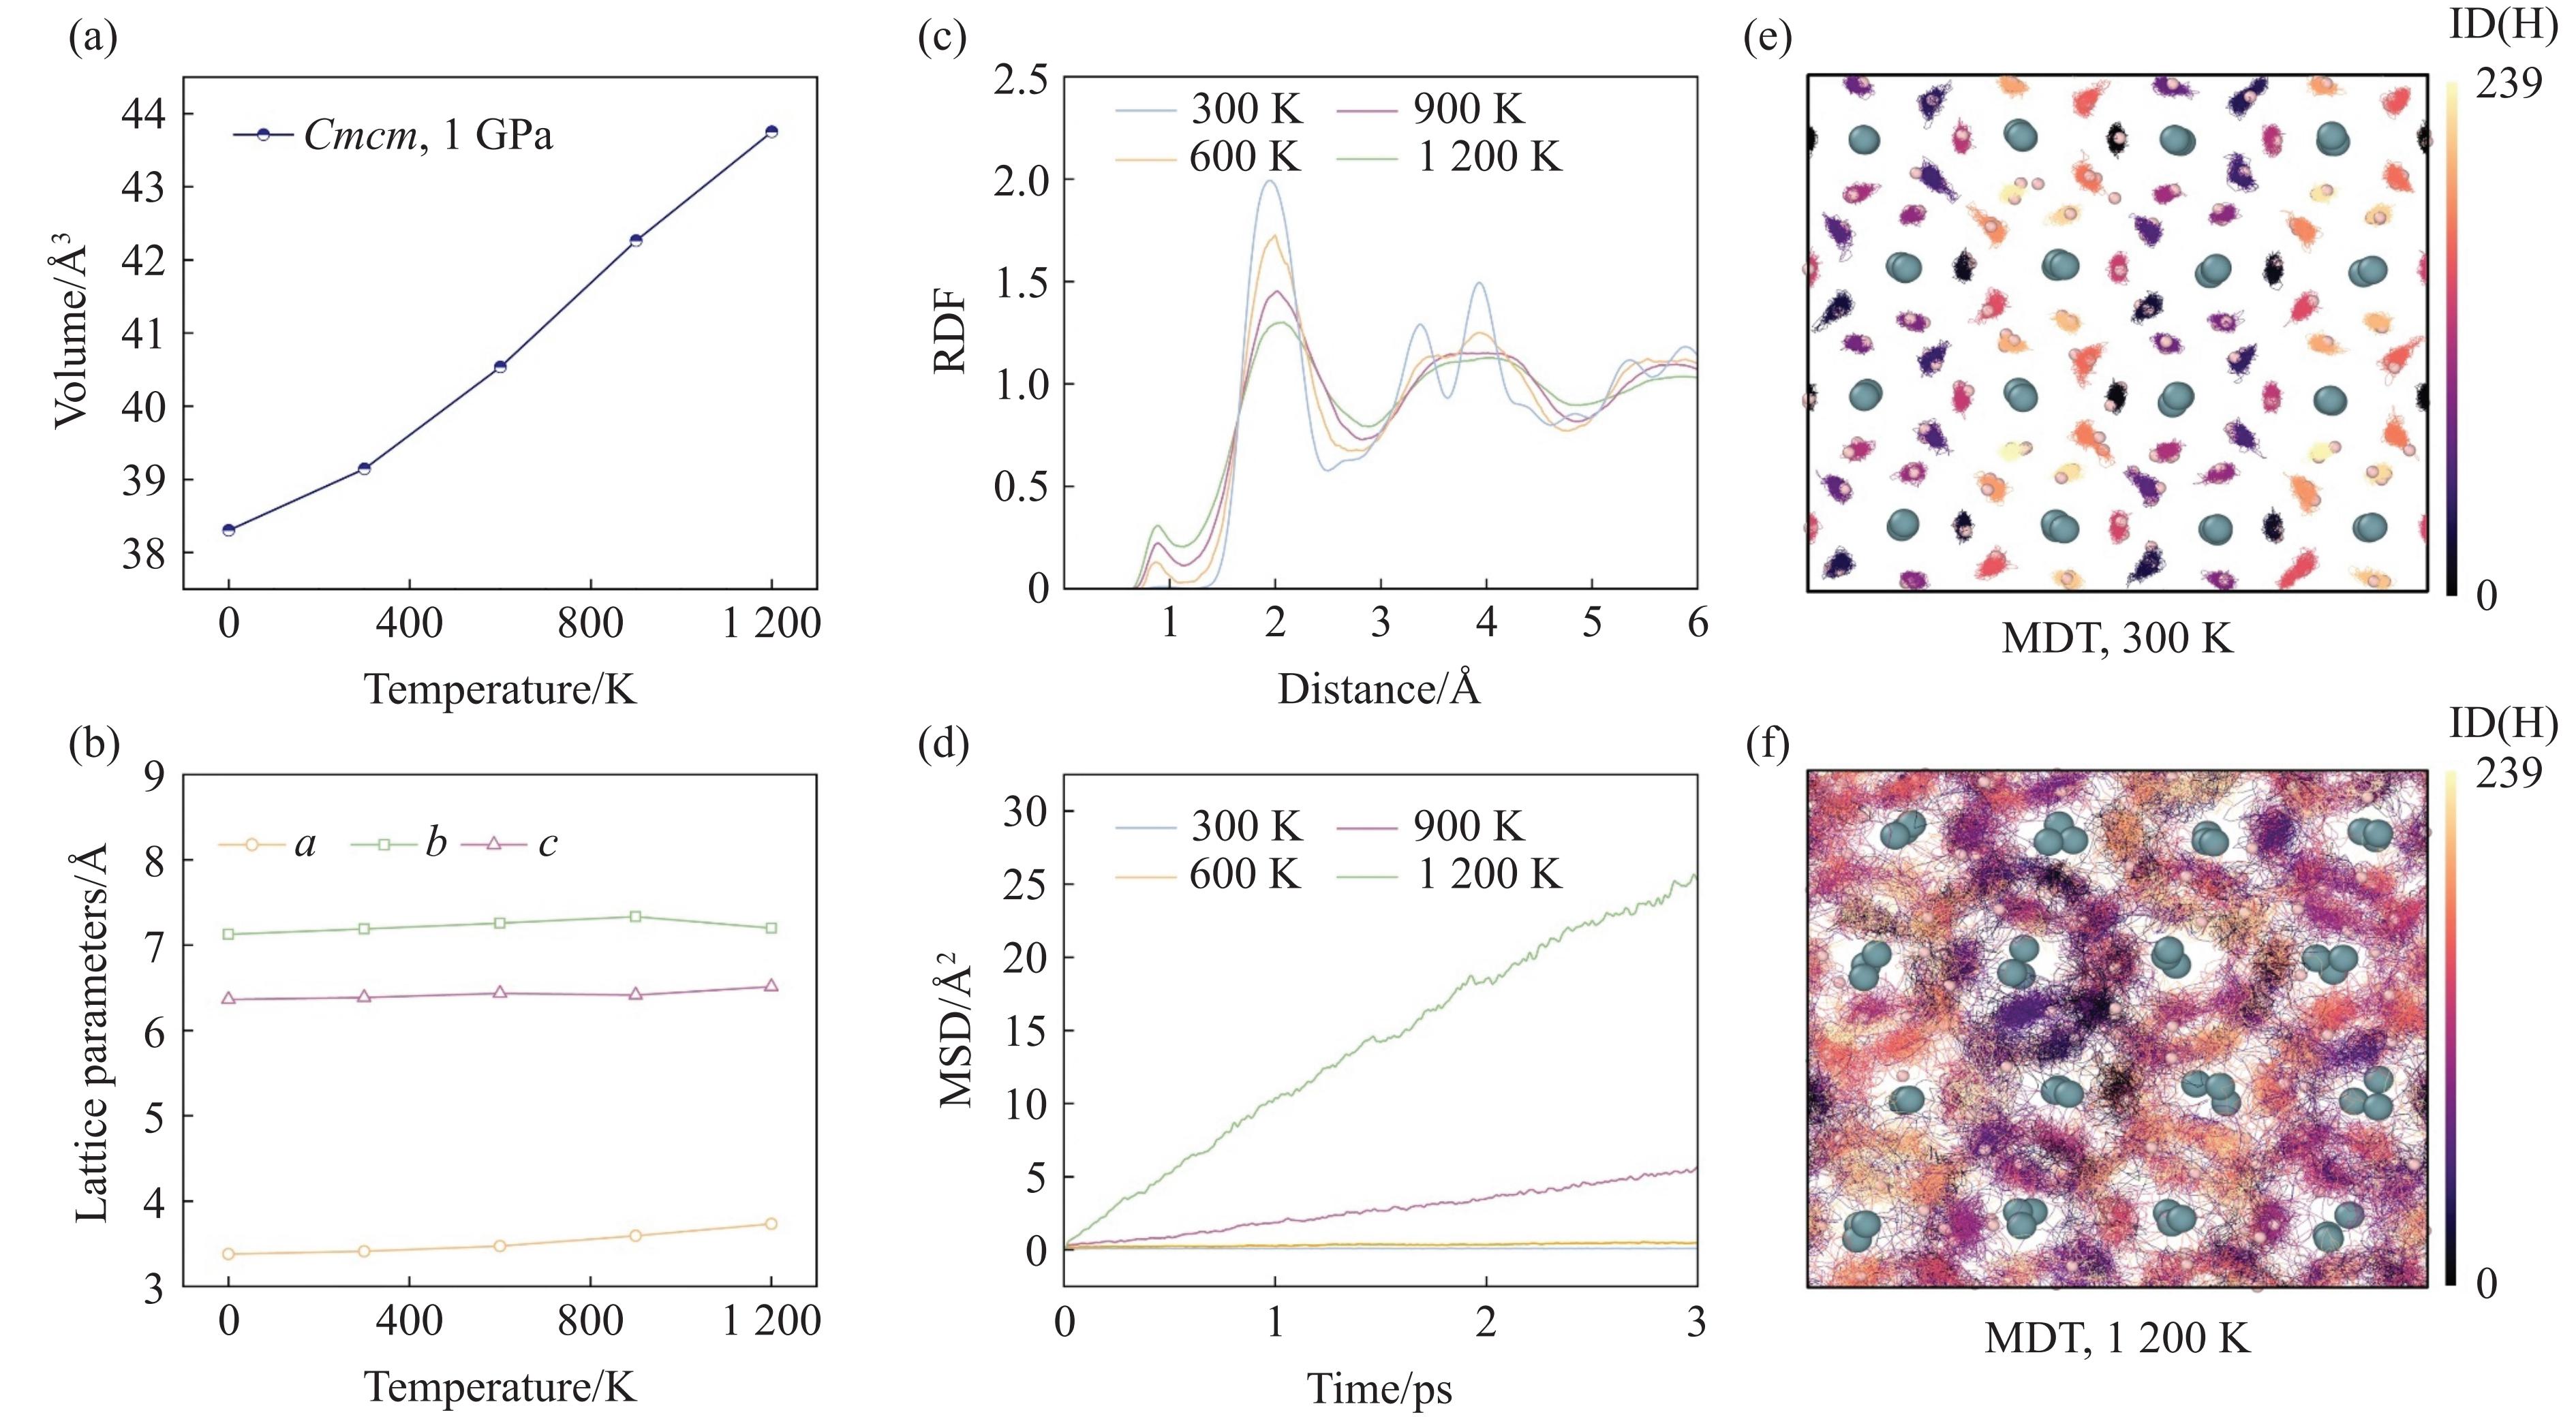
The thermodynamic, mechanical, and dynamical stability, along with the electronic properties of UH5 within 30 GPa, are systematically investigated using first-principles calculations. The experimentally synthesized orthorhombic, hexagonal, and cubic phases are all found to be magnetic materials, with spin polarizations of 82%, 100%, and 100%, respectively, and their thermodynamic stability decreases sequentially. Elastic constant and phonon calculations demonstrate that all three phases are mechanically and dynamically stable. Chemical bonding analysis indicates that this stability primarily originates from the prevalent covalent U-H interaction within the lattice. Furthermore, it is predicted that the orthorhombic phase, which has been experimentally quenched to 1 GPa, transforms into a superionic state at 1200 K, where hydrogen ions undergo rapid diffusion within the uranium sublattice interstices, achieving a diffusion coefficient of 1.2×10⁻⁴ cm2/s.

Structural Evolution, Dual Role of Hydrogen and Superconductivity in Lithium-Rich Li-N-H Compounds under High Pressure
LI Qiuyue, HAN Shuai, YANG Guochun
2026, 40(11): 110106. doi: 10.11858/gywlxb.20261017
HTML PDF (95)
Abstract:

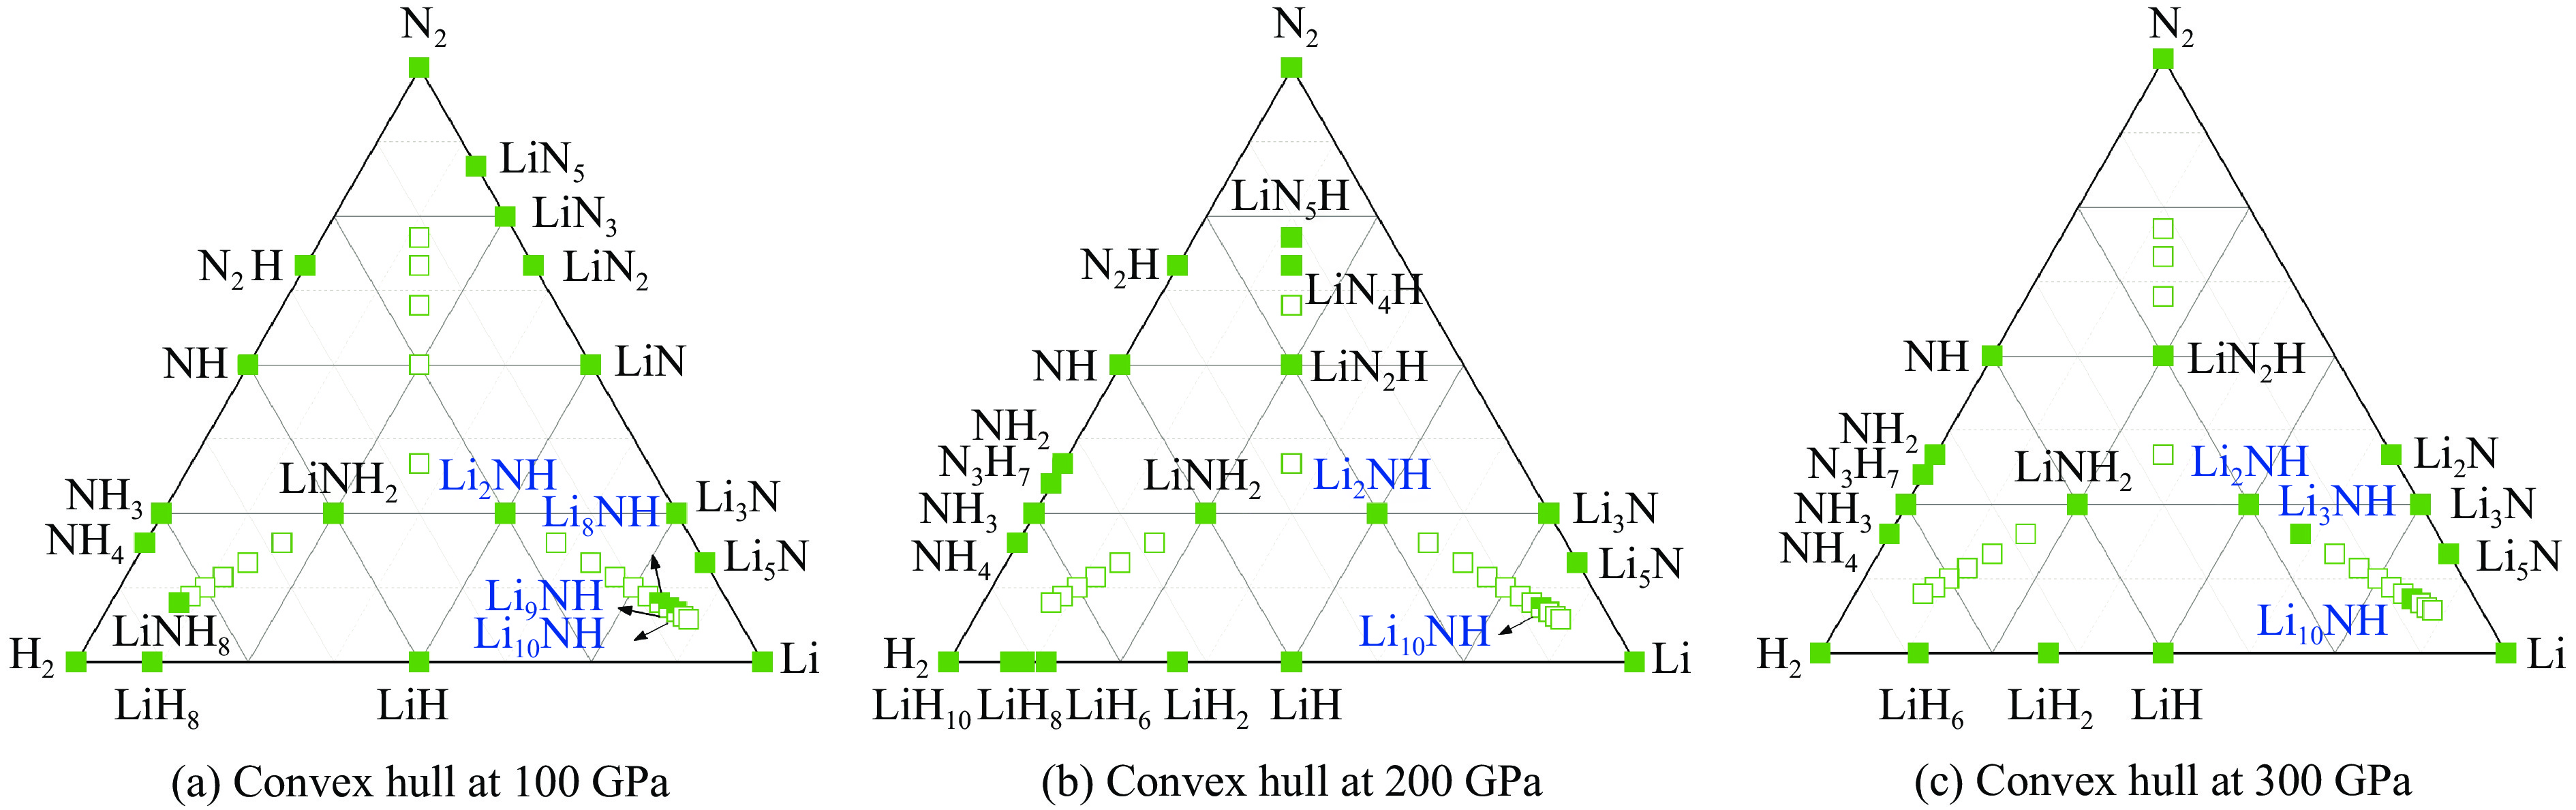
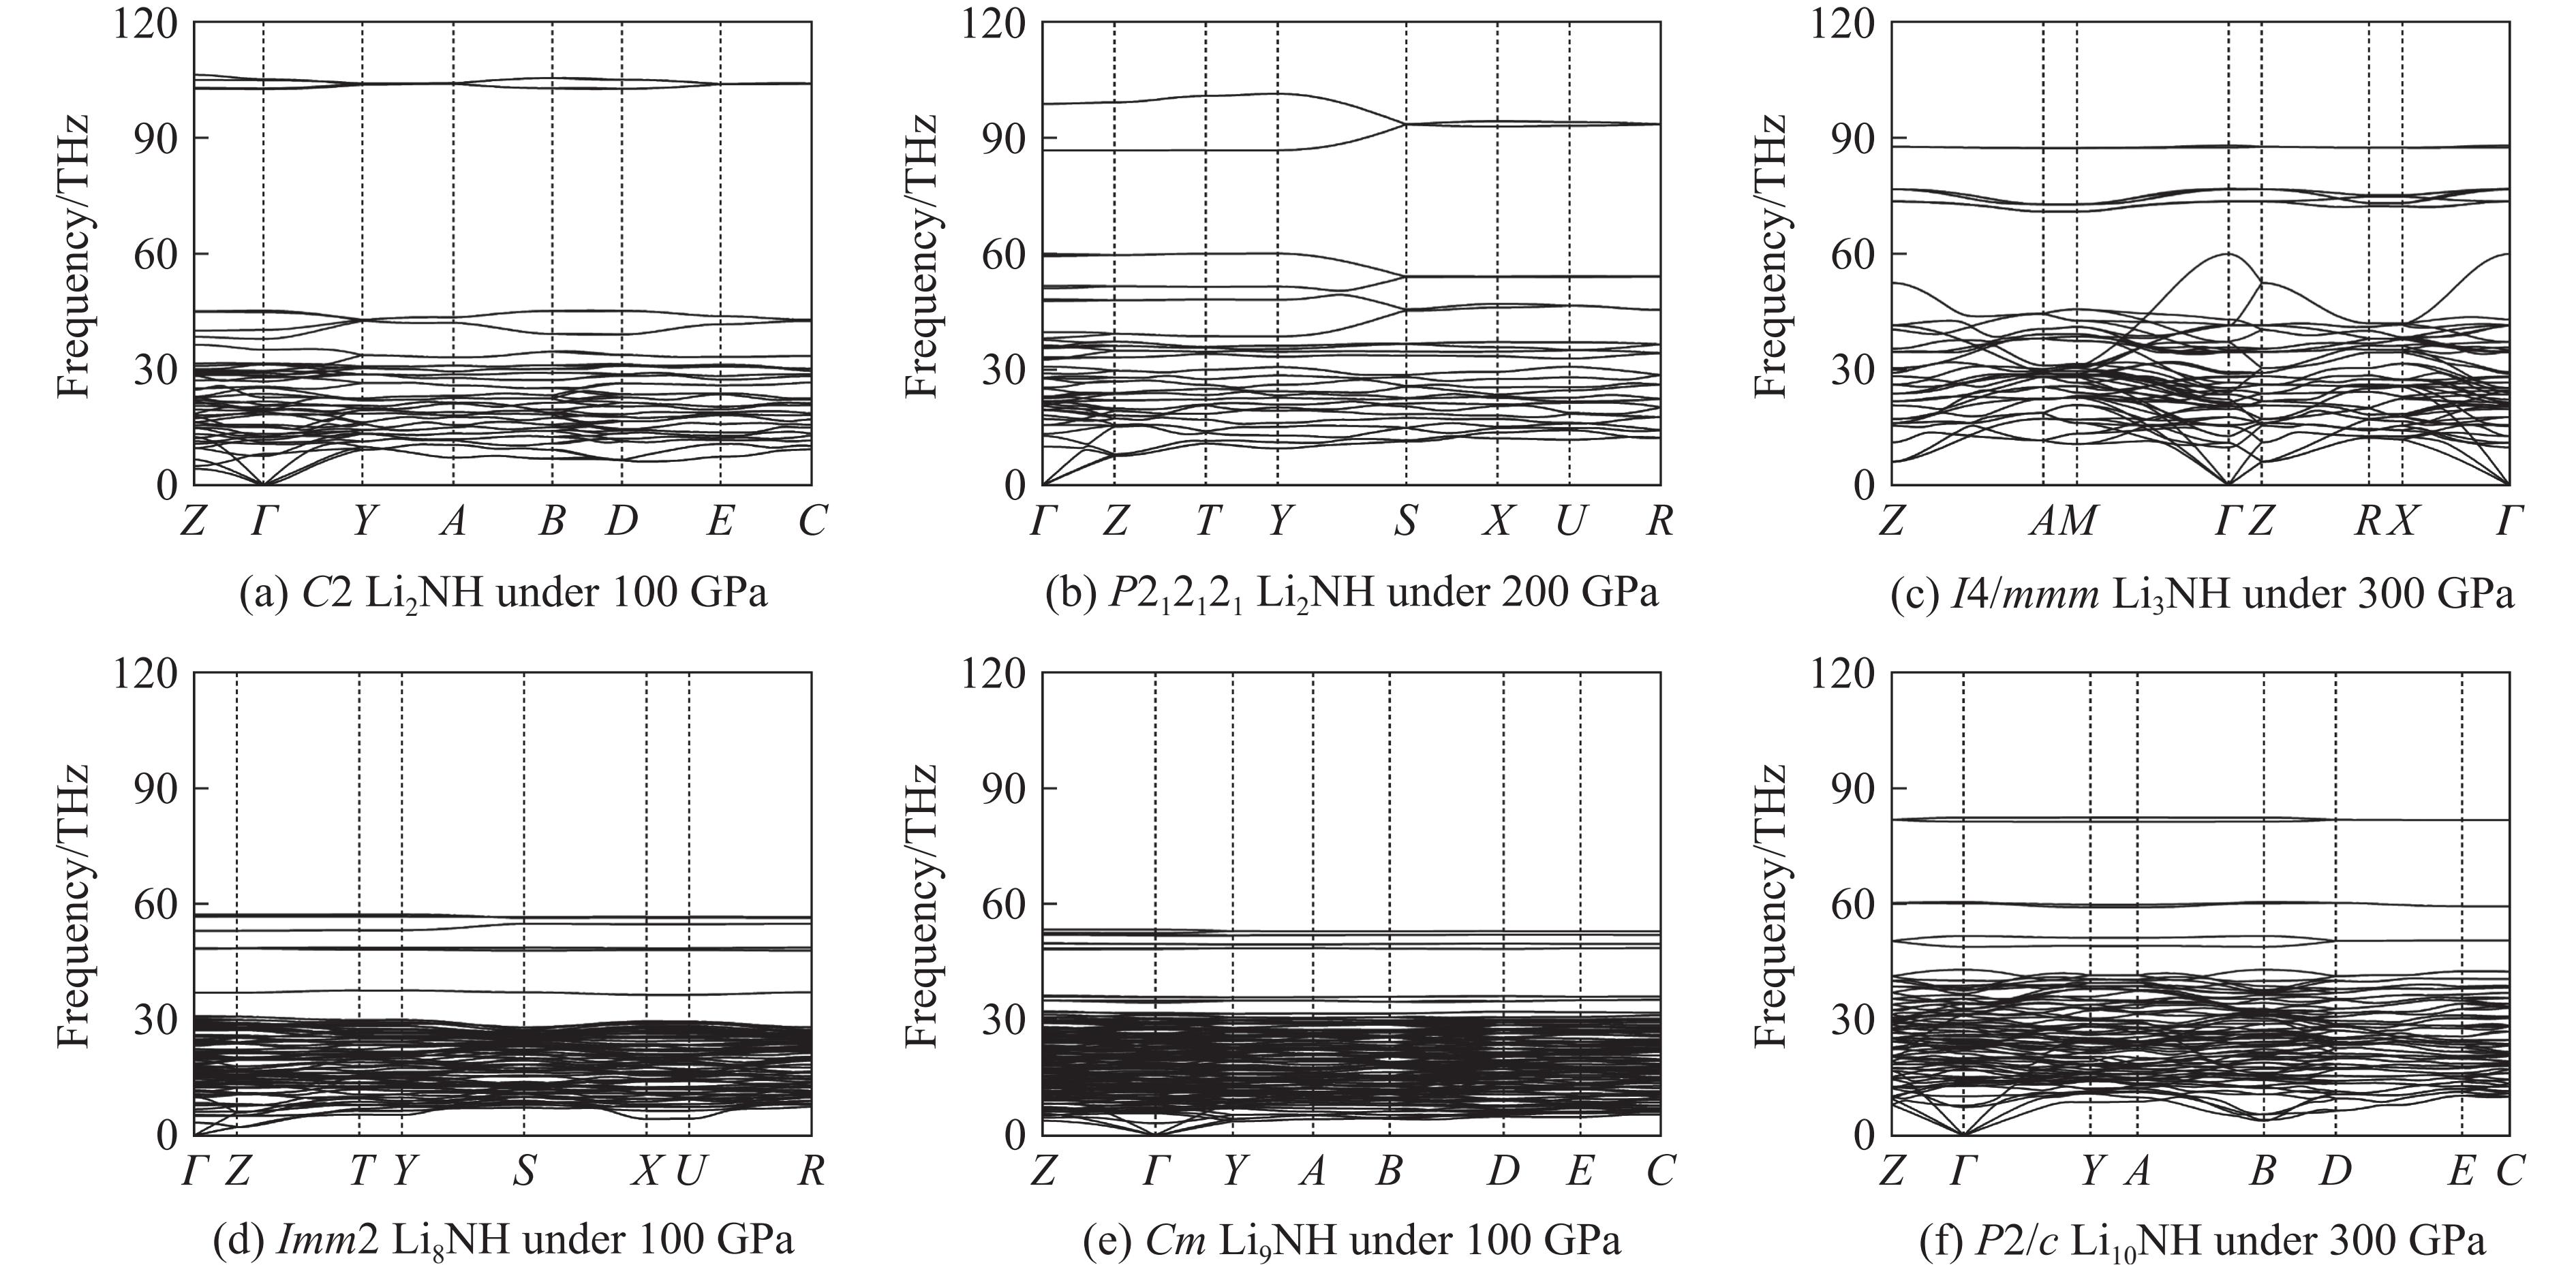
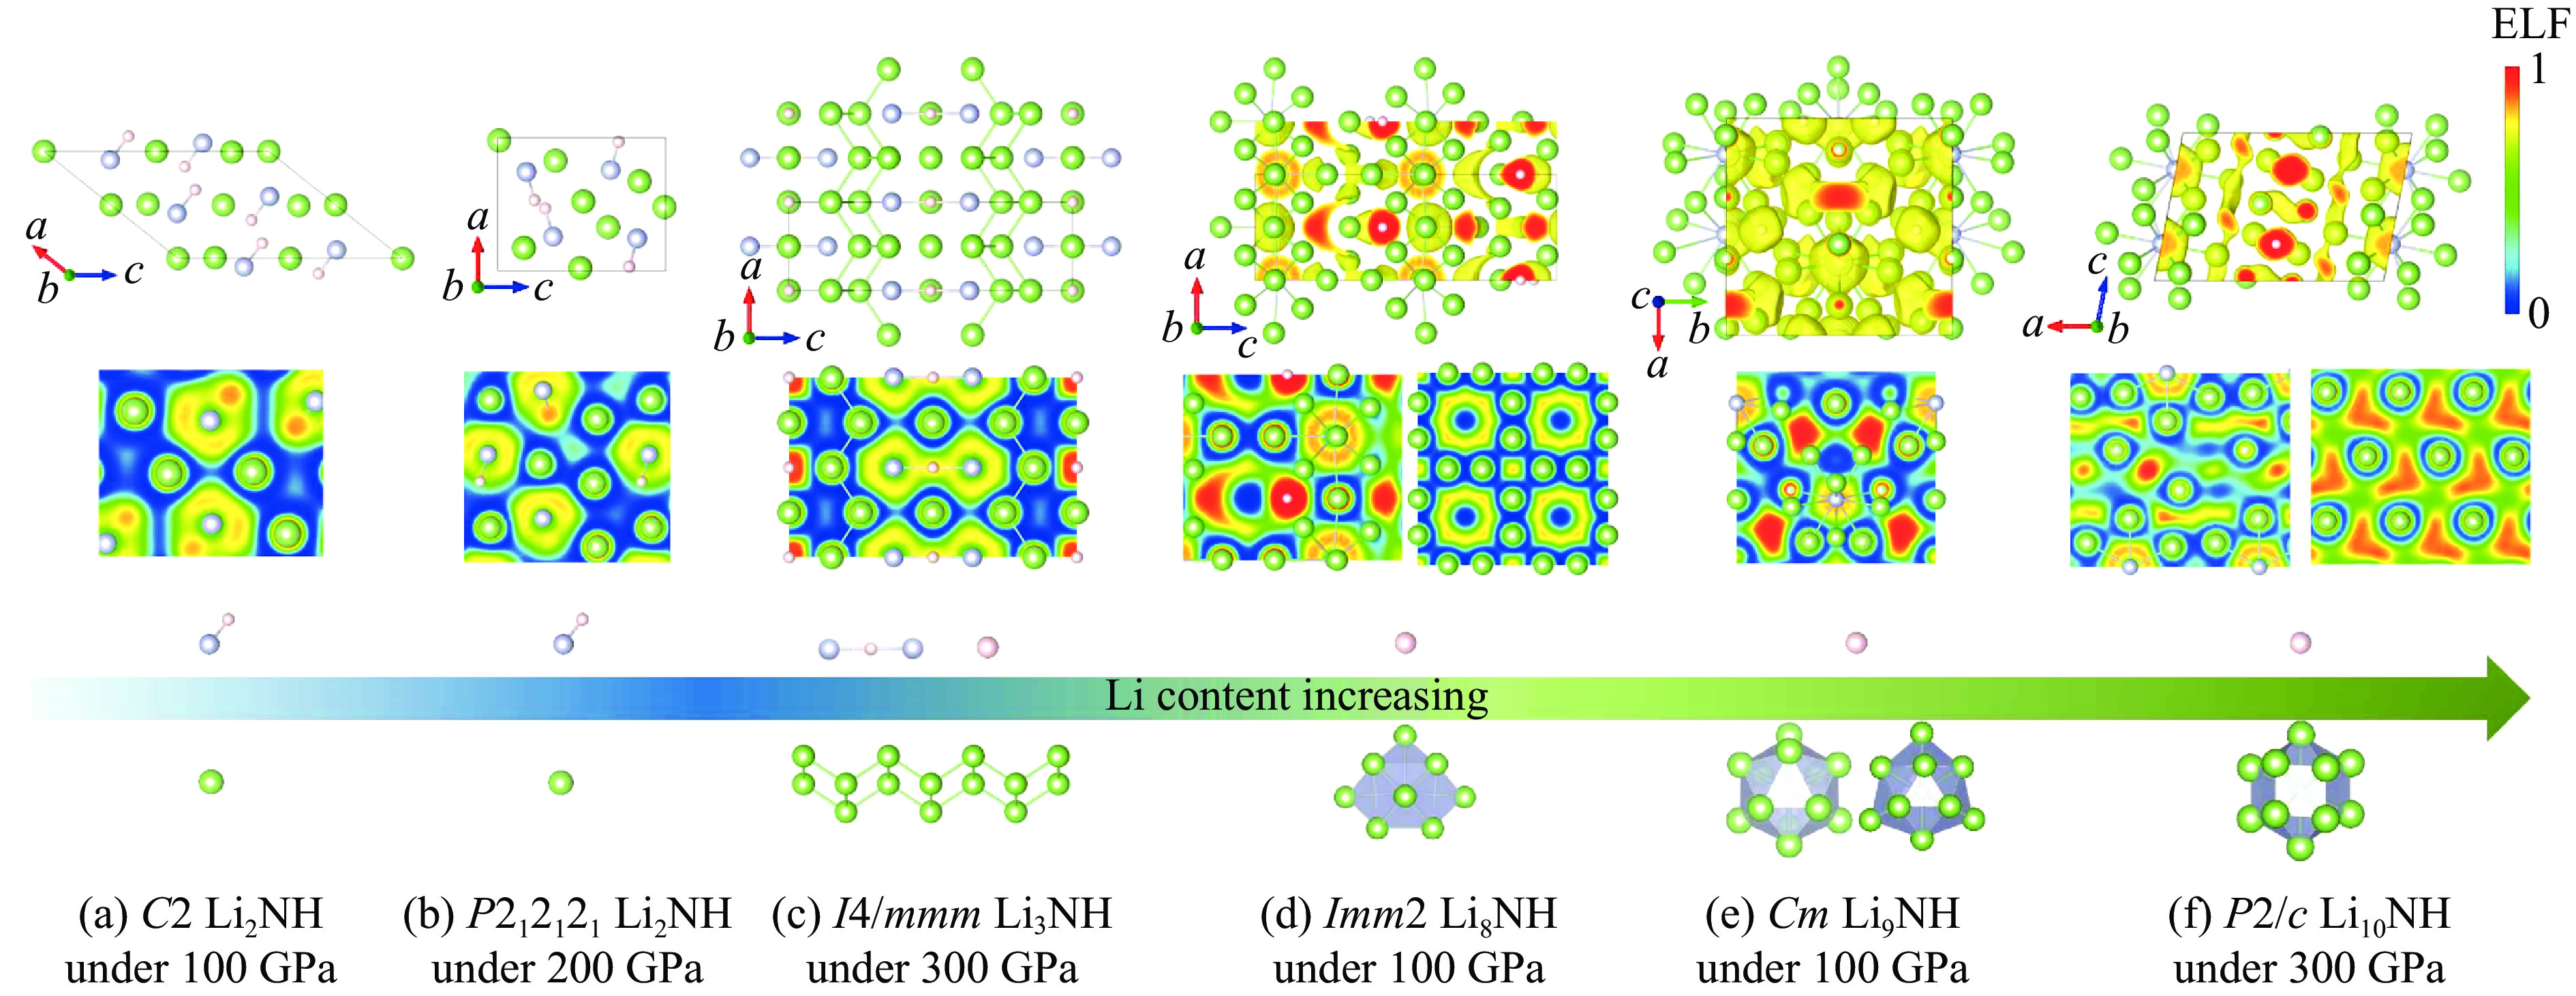
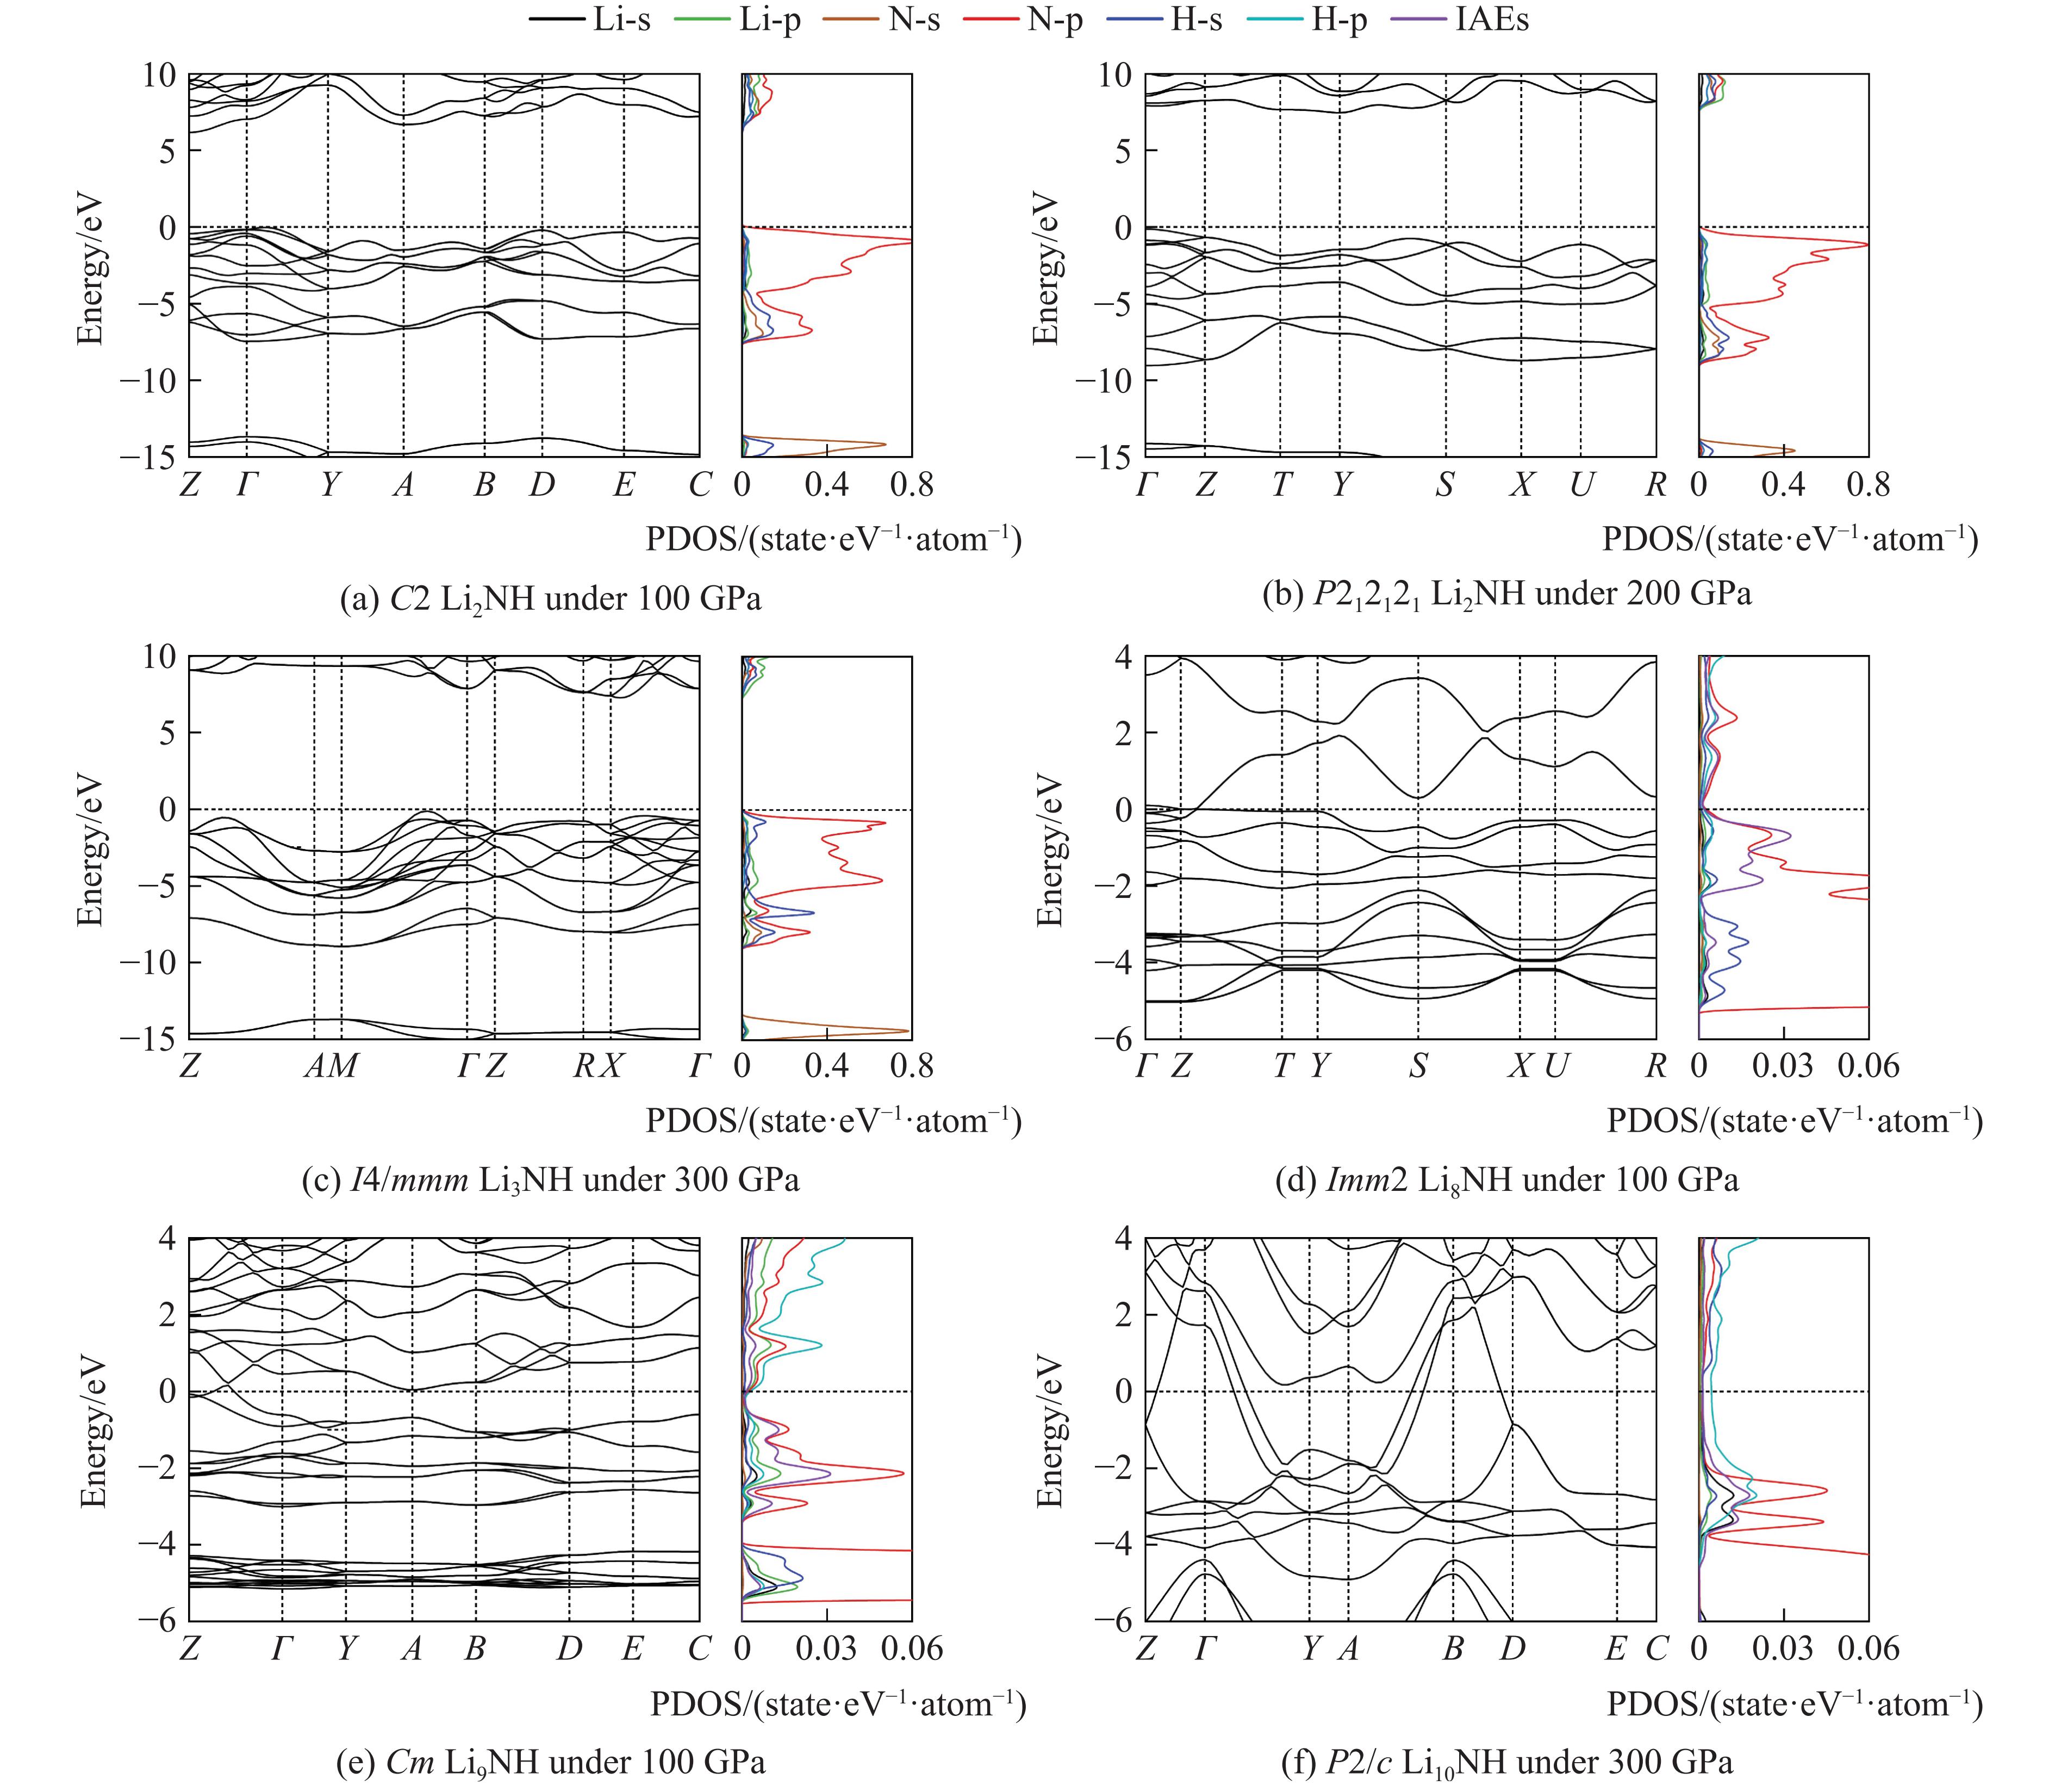
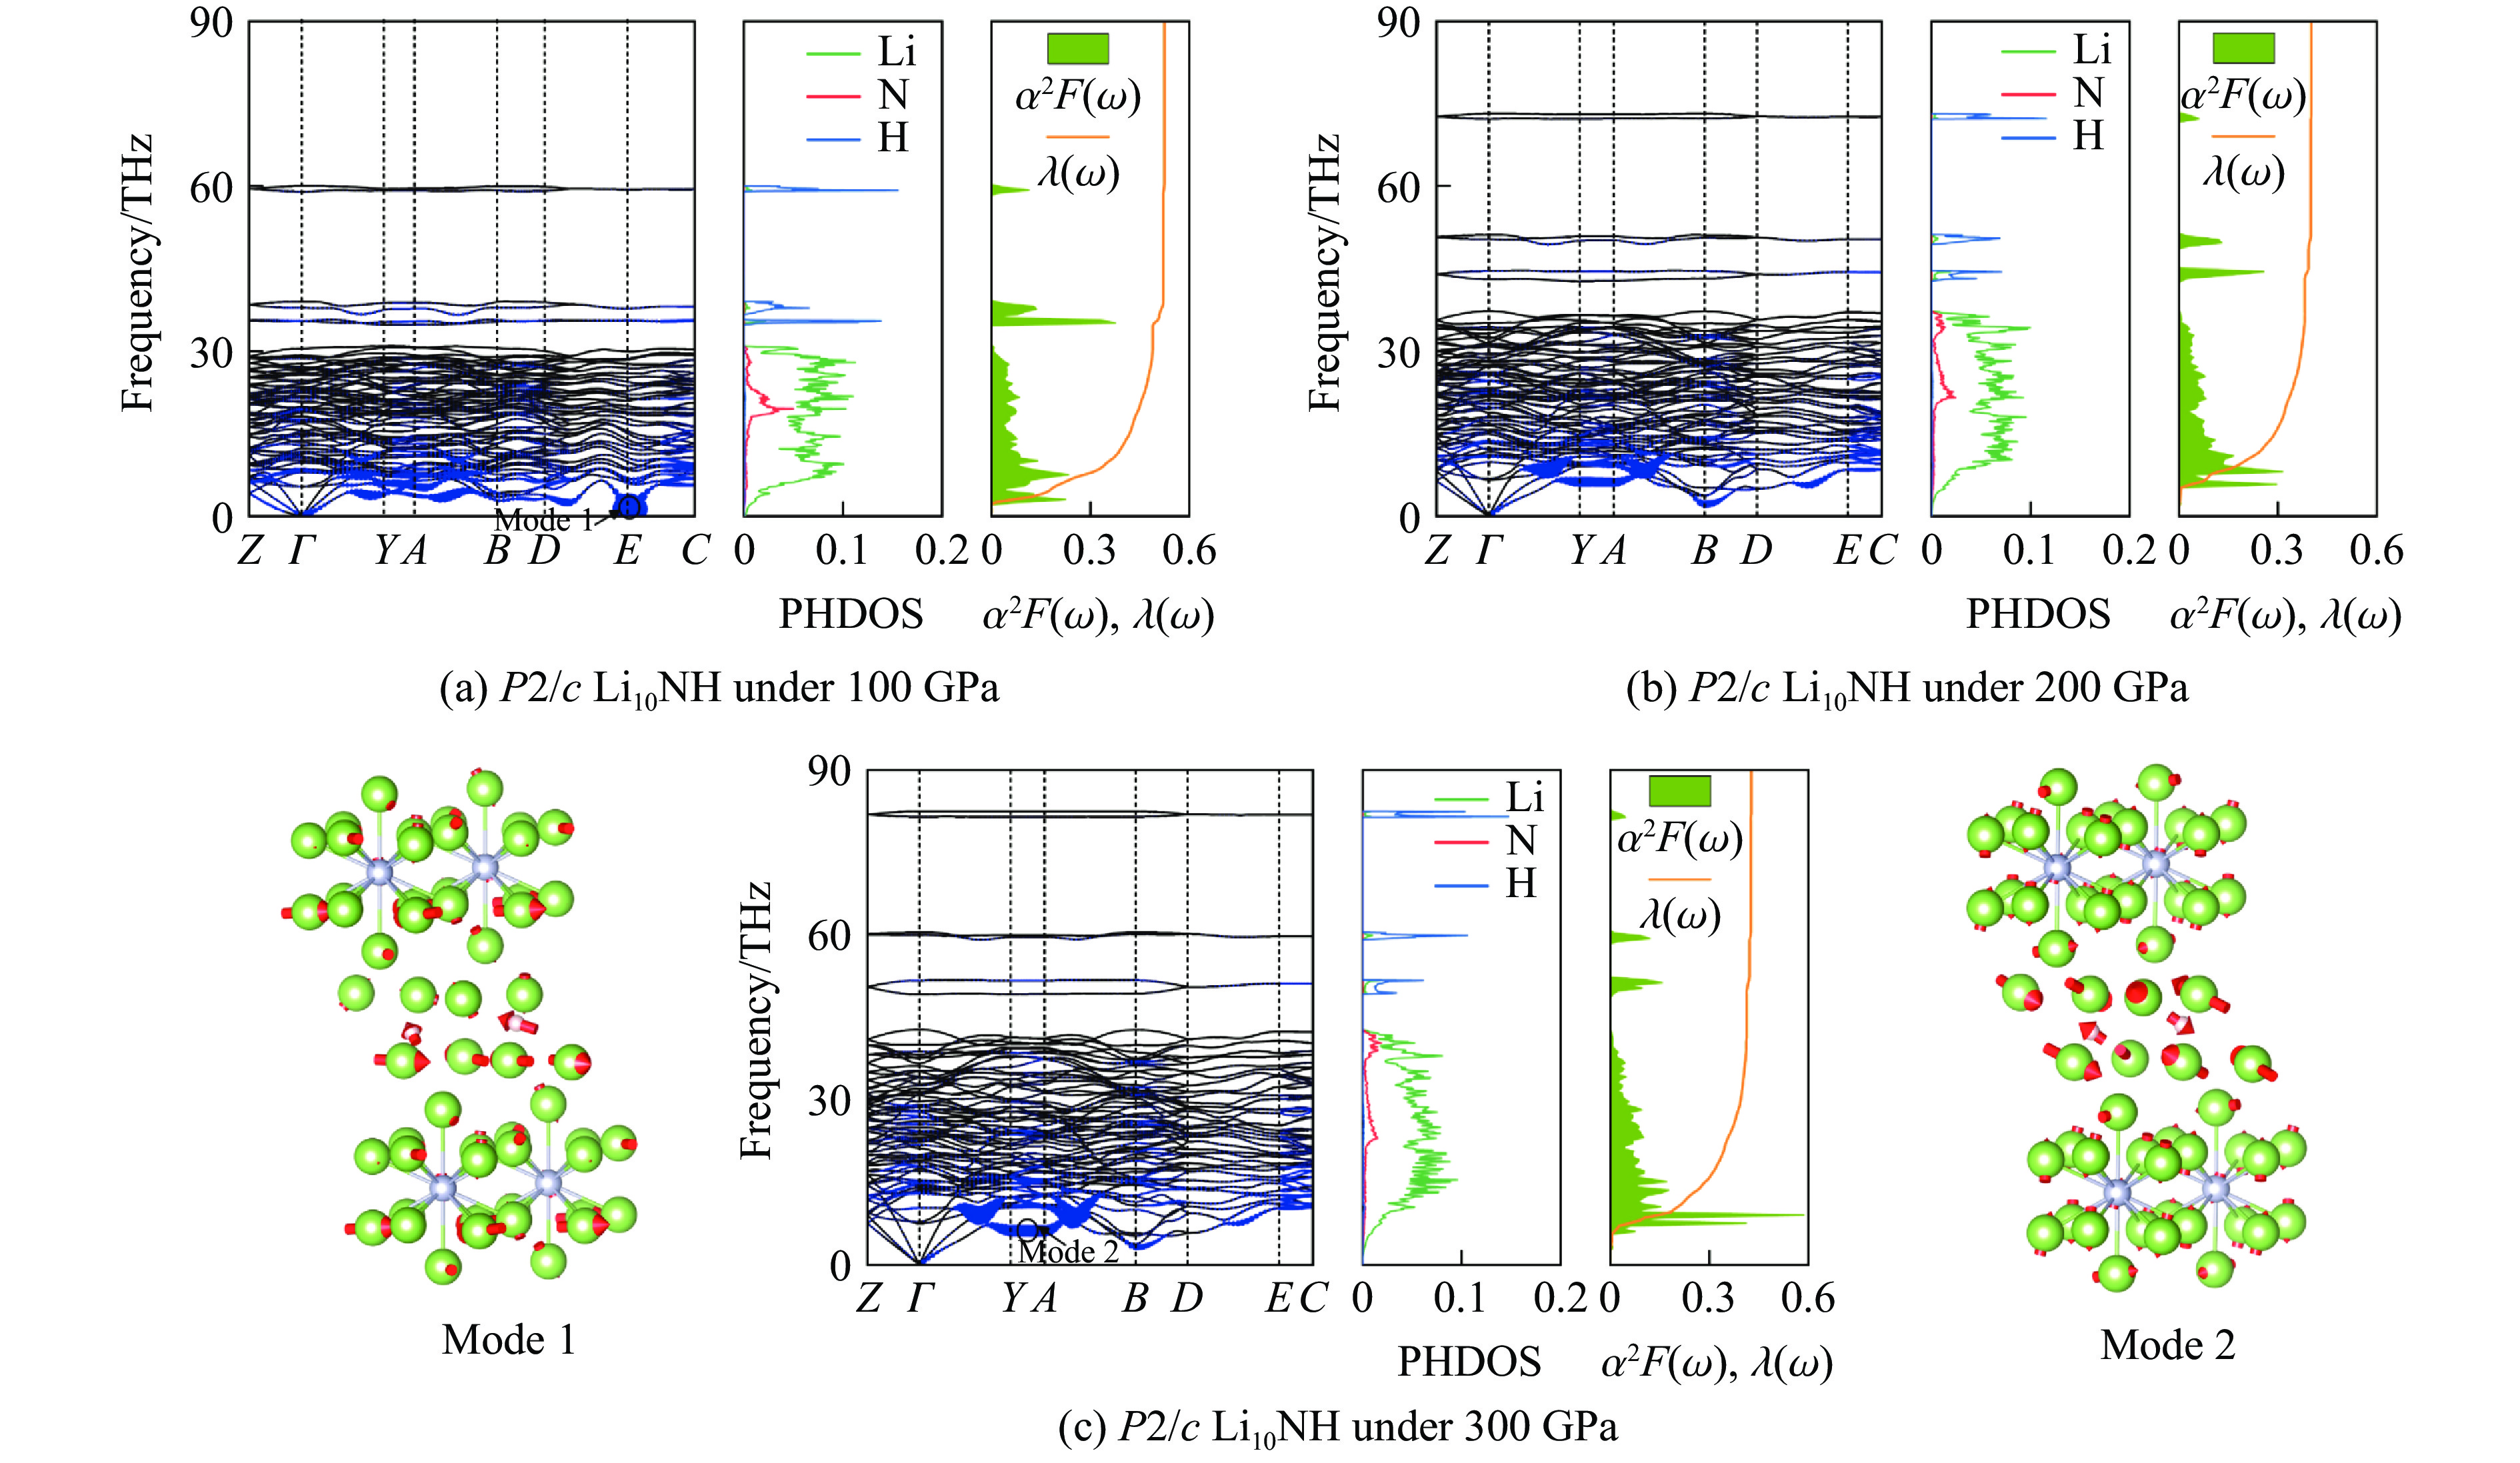
Using first-principles calculations combined with CALYPSO structure prediction method, we systematically investigated the crystal structures, electronic properties, and superconducting behavior of the Li-N-H ternary system under pressure ranging from 100 to 300 GPa. Six thermodynamically stable or metastable Li-rich compounds were identified: C2 Li2NH, P212121 Li2NH, I4/mmm Li3NH, Imm2 Li8NH, Cm Li9NH, and P2/c Li10NH. The results reveal a distinct evolution in the chemical role of hydrogen with increasing lithium content. In phases with lower Li content, H atoms tend to form covalent bonds with N atoms, thereby achieving a stable closed-shell electronic configuration. As the Li content increases, H atoms progressively occupy lattice interstitial sites, acting as electron acceptors that trap excess interstitial anionic electrons (IAEs). This transformation effectively tunes the quantity, degree of localization, and spatial topology of IAEs. Correspondingly, C2 Li2NH, P212121 Li2NH, and I4/mmm Li3NH are insulating non-electrides, while Imm2 Li8NH, Cm Li9NH, and P2/c Li10NH are electrides, which exhibit metallic behavior. Notably, P2/c Li10NH demonstrates superconductivity with a predicted transition temperature of 4.8 K at 100 GPa, mainly originating from the strong electron-phonon coupling between H-p orbital electrons and low frequency phonon modes dominated by Li atoms. This work elucidates the dual functional role of hydrogen in high-pressure Li-N-H systems—from covalent N―H bonding coordination to IAEs capturing—and provides theoretical insights for the rational design of novel high-pressure electrides and superconducting materials.

Crystal Structure and Physical Properties of Sr2He Compound under High Pressure
WANG Qingmu, ZHANG Pan, SHI Jingming, LI Yinwei
2026, 40(11): 110107. doi: 10.11858/gywlxb.20251084
HTML PDF (111)
Abstract:

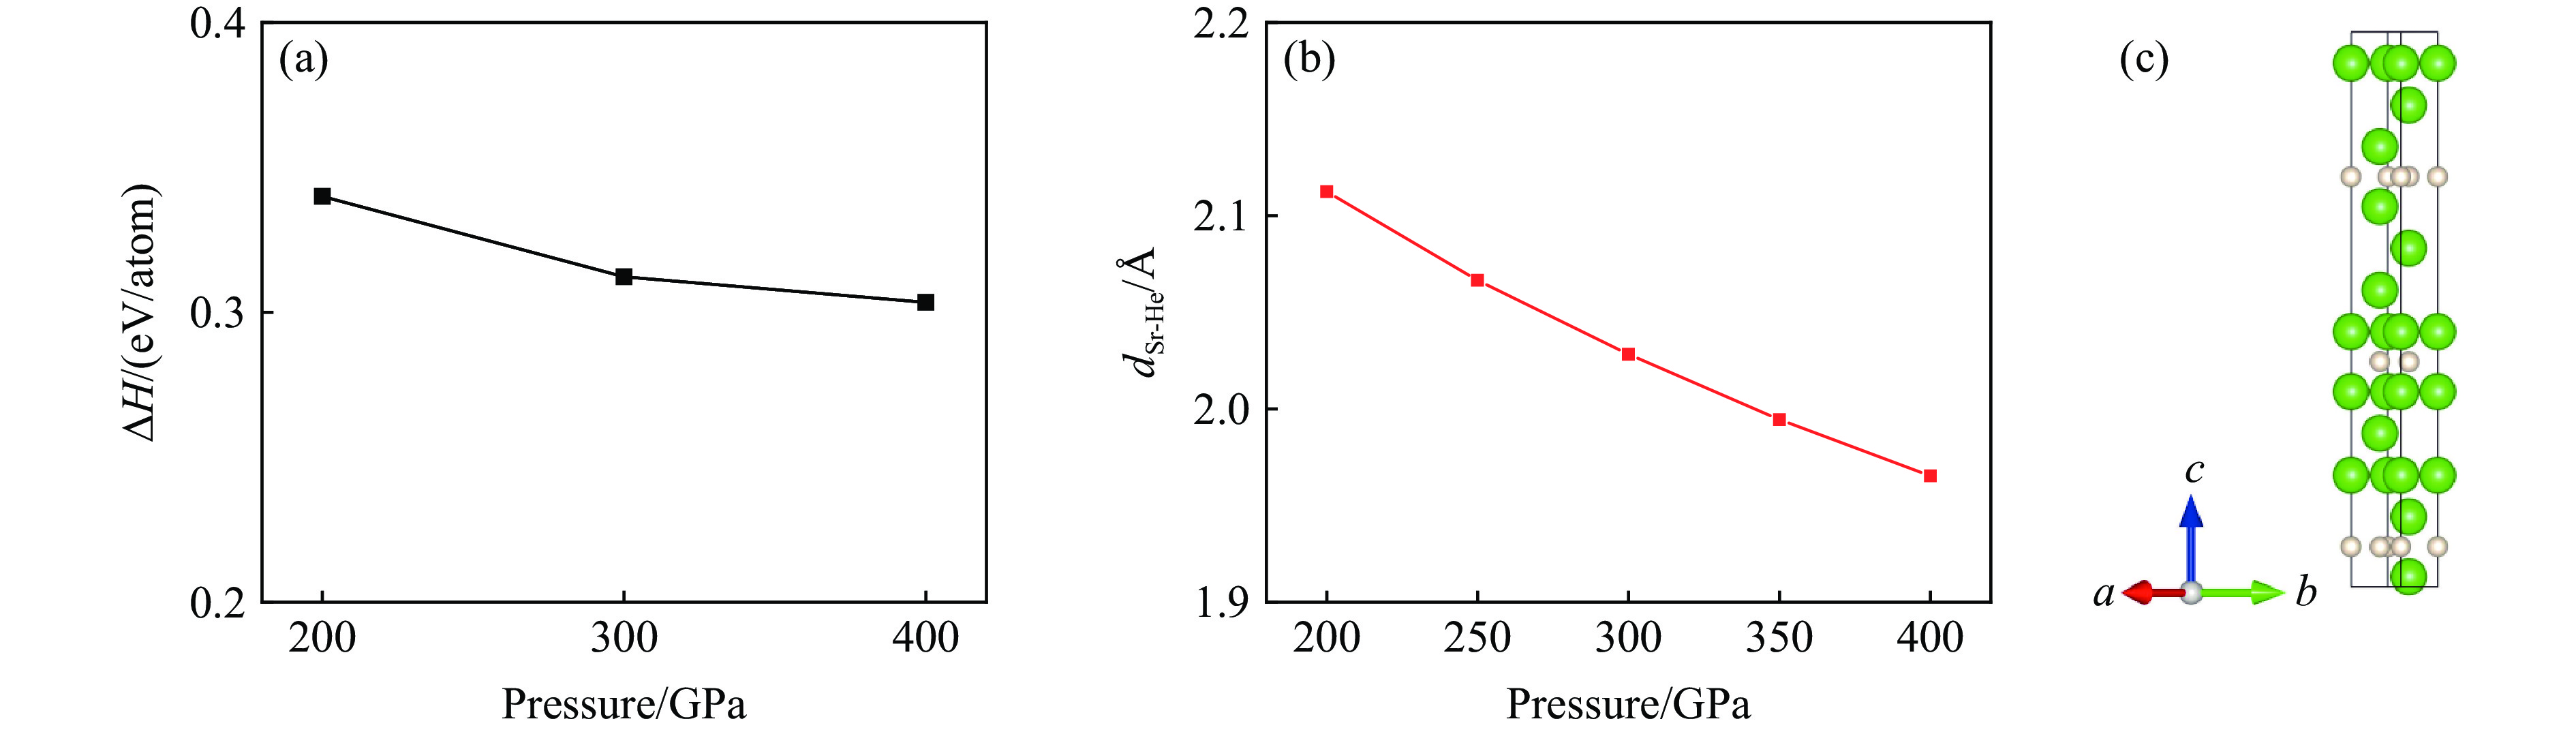
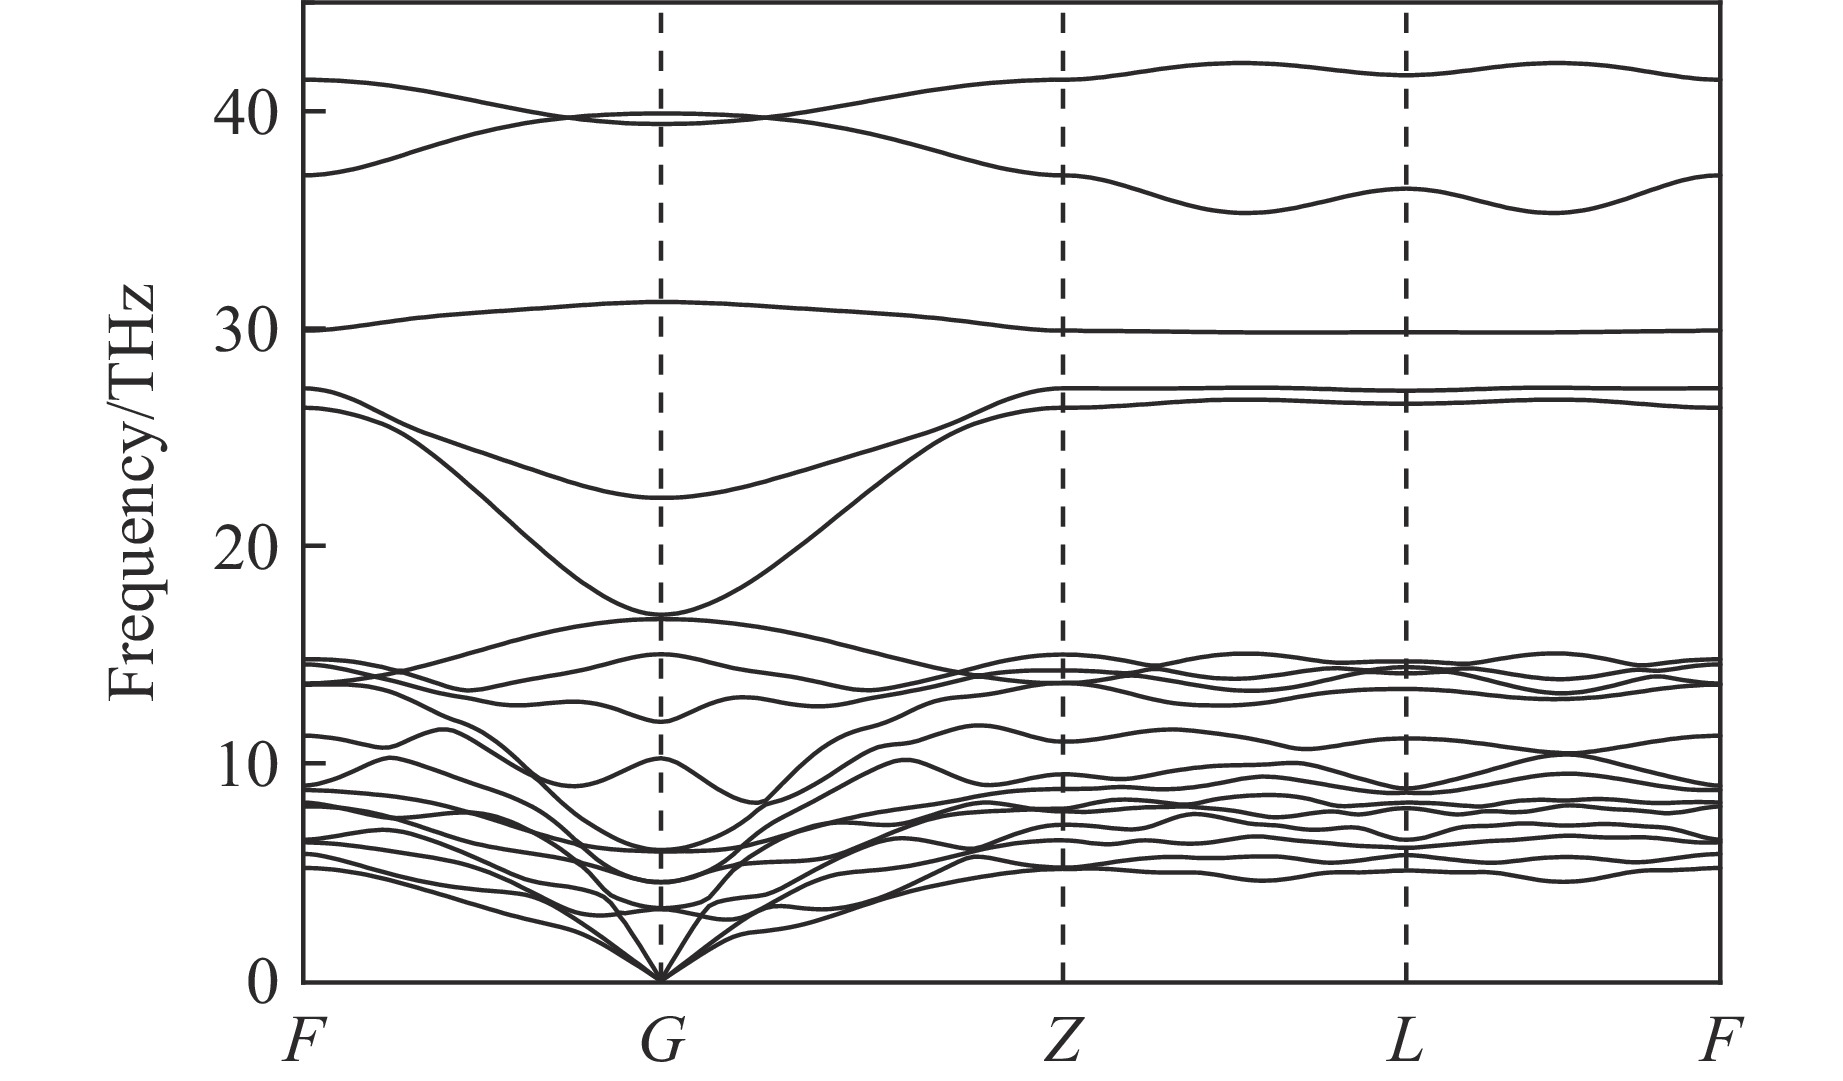
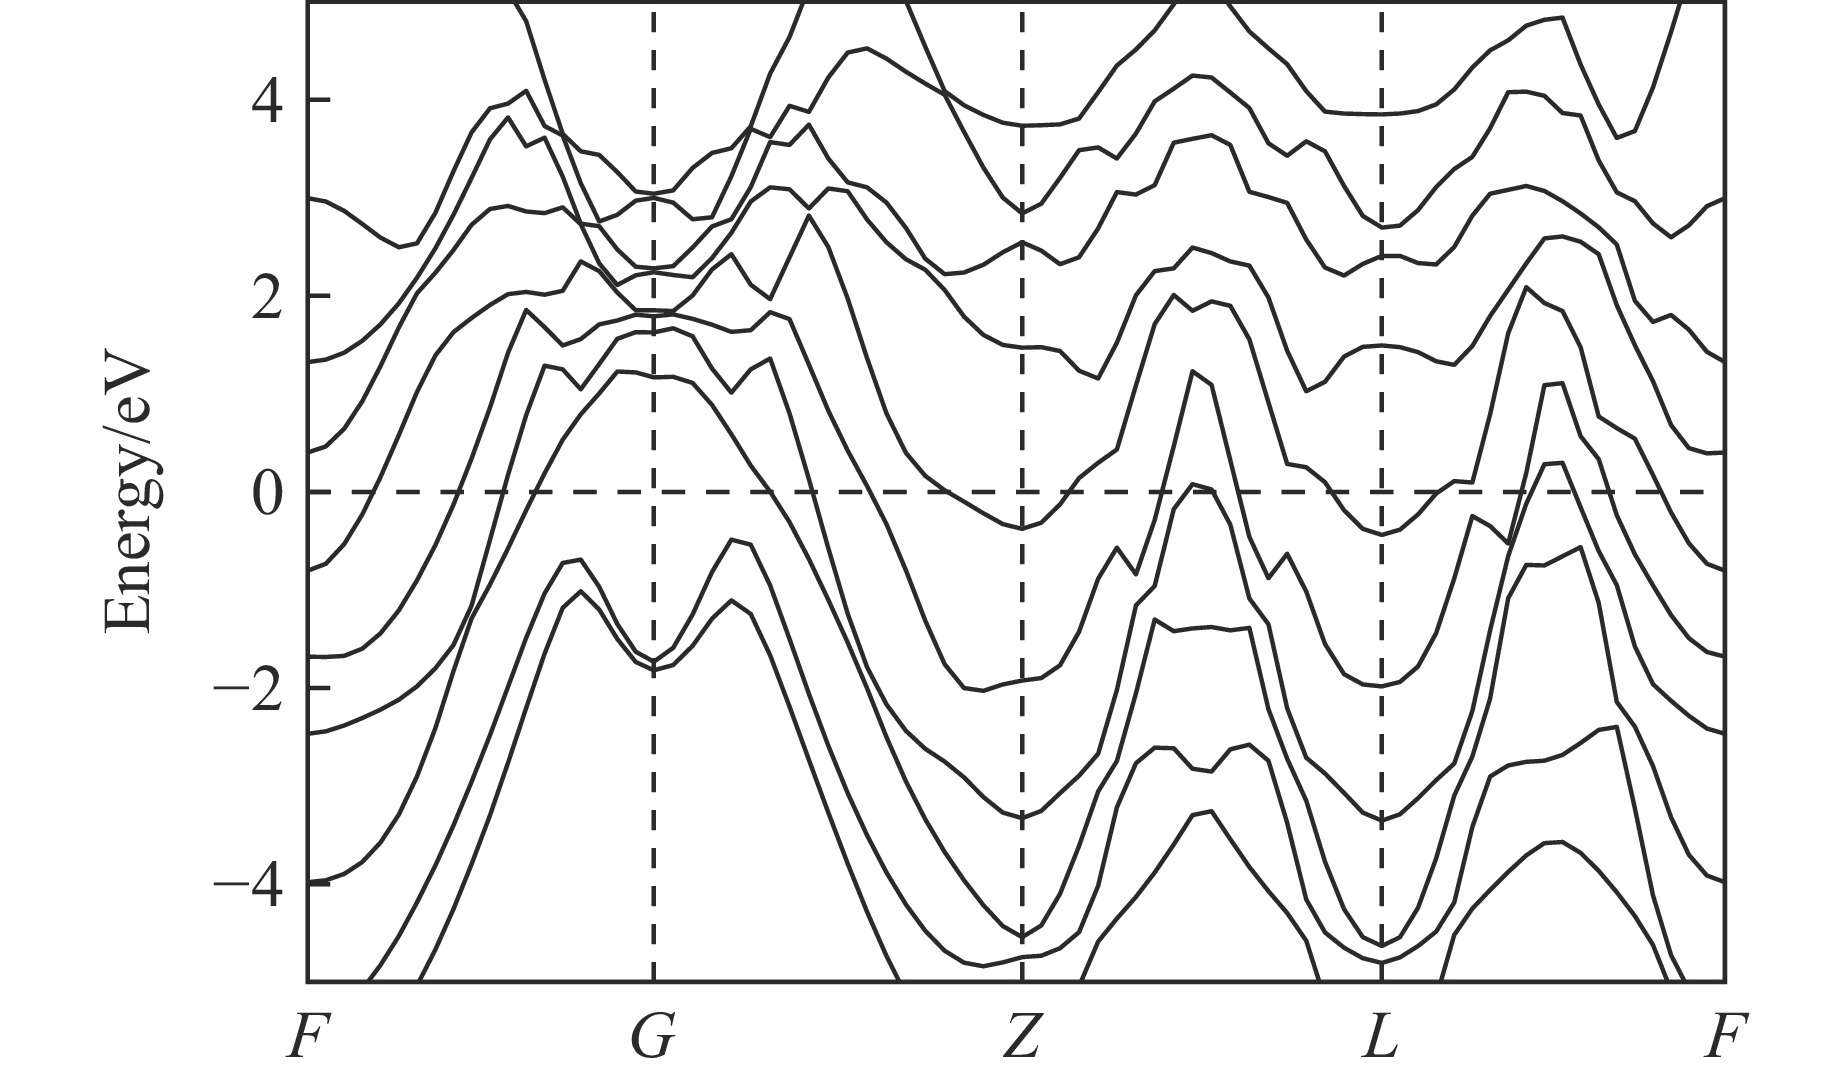
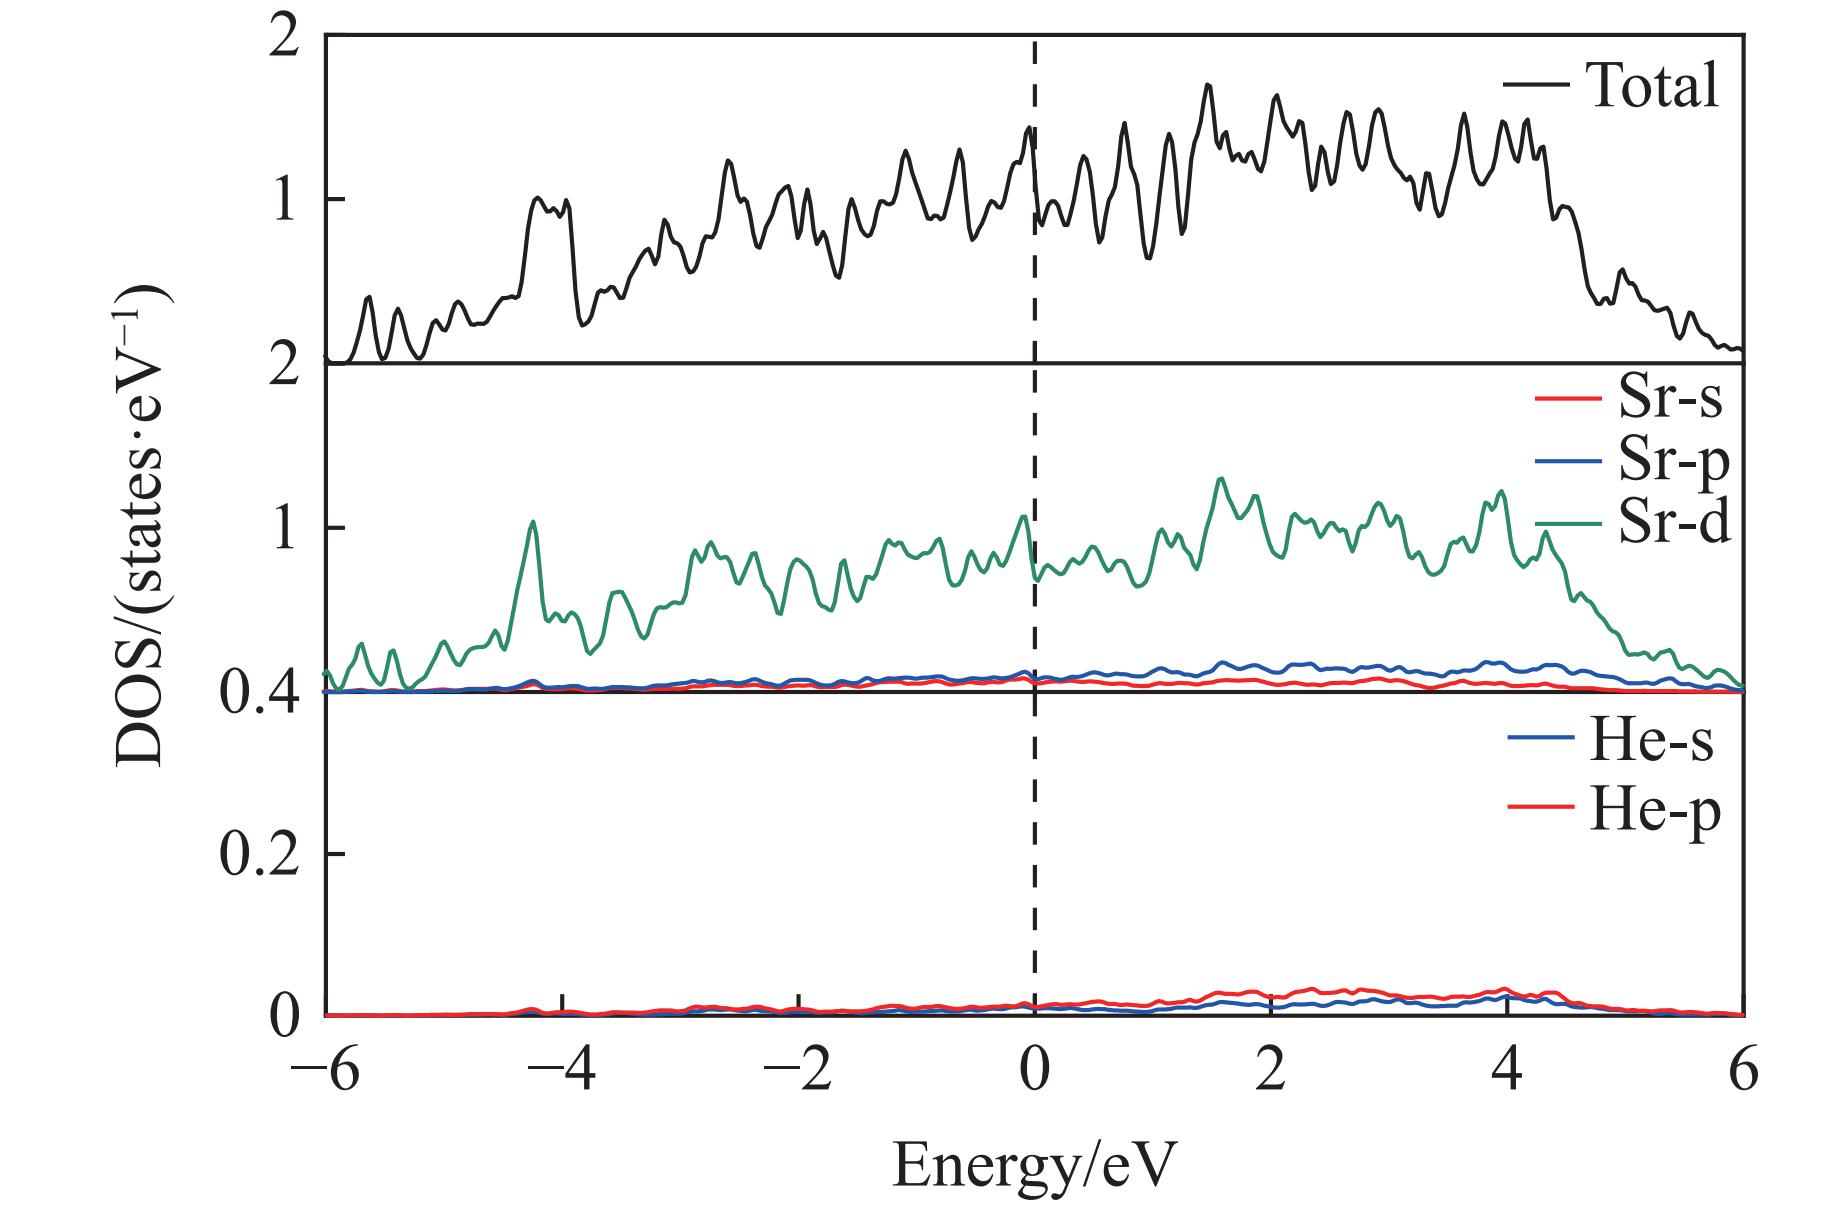
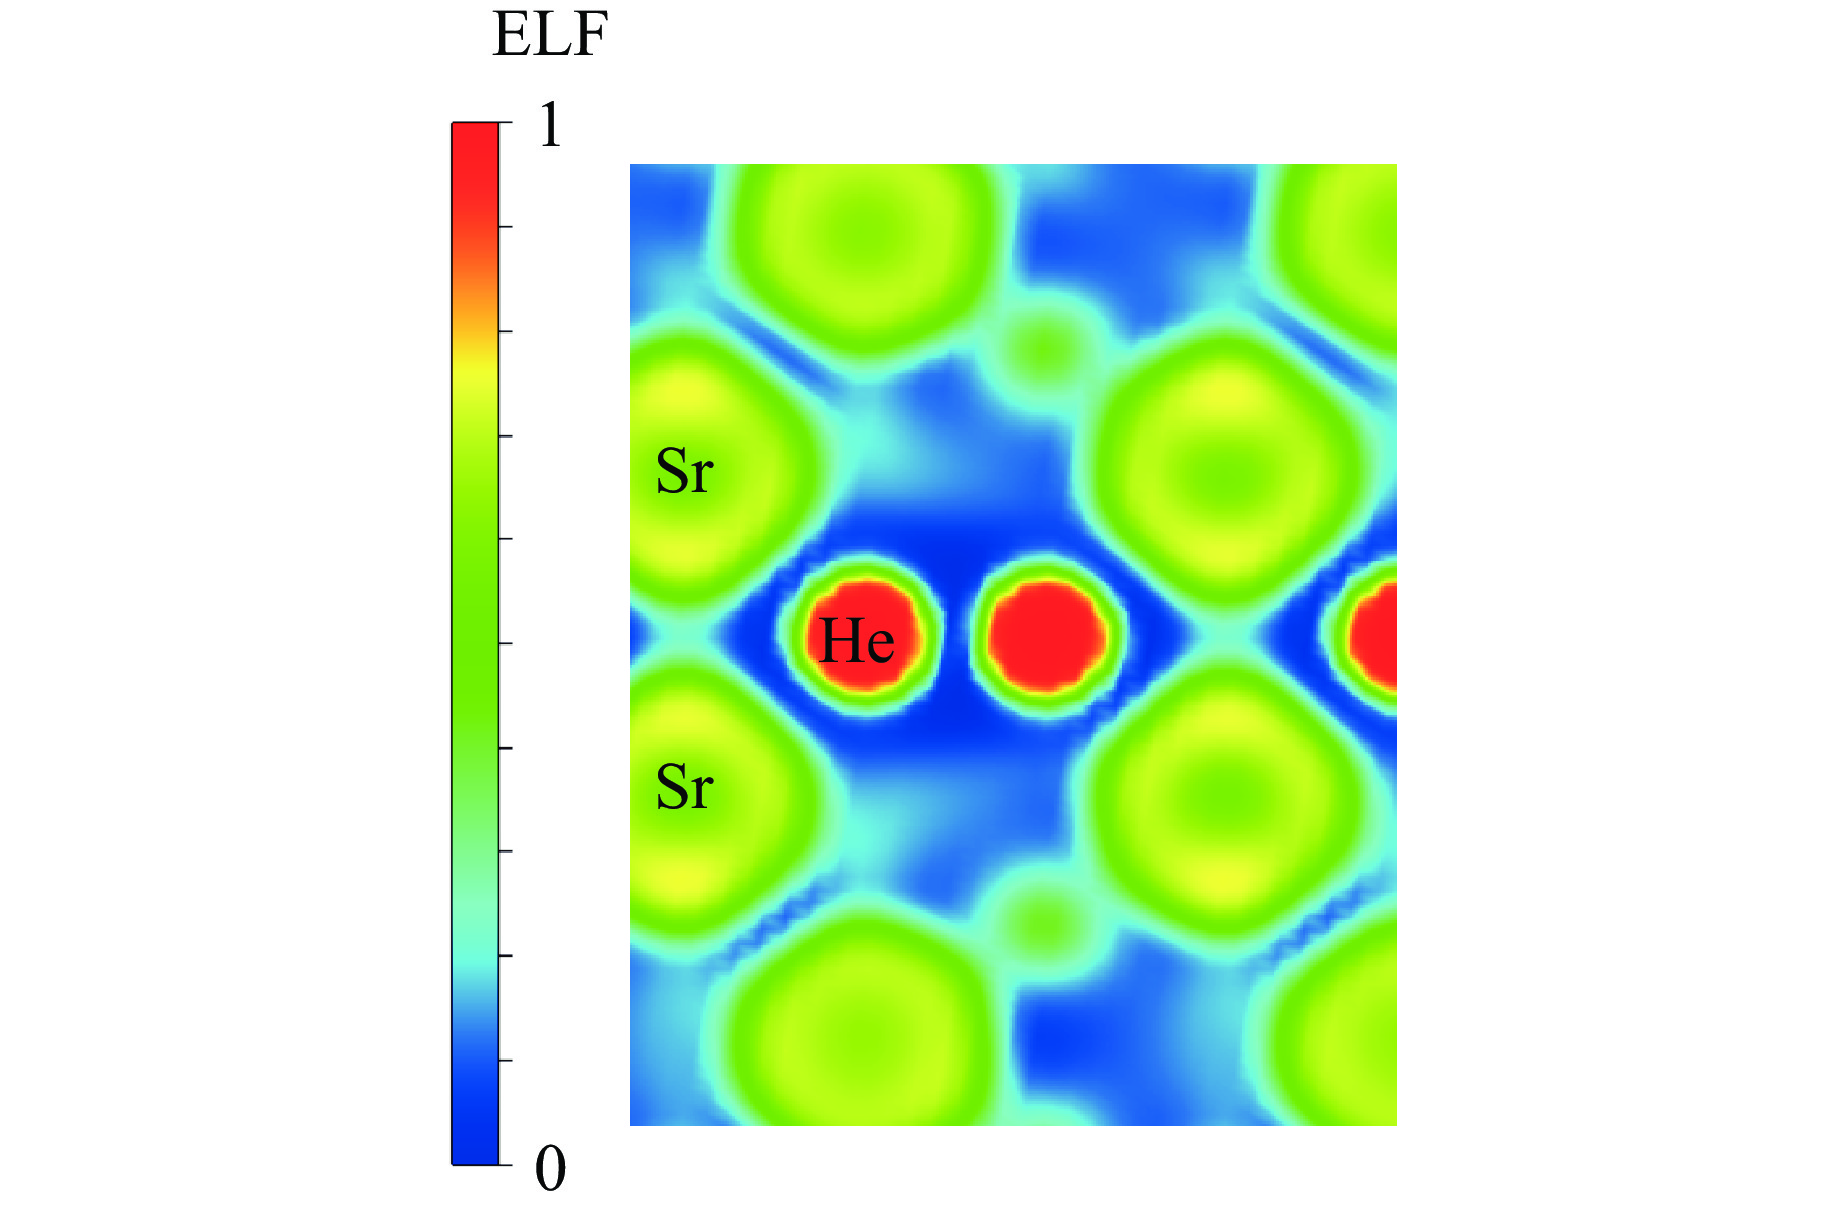
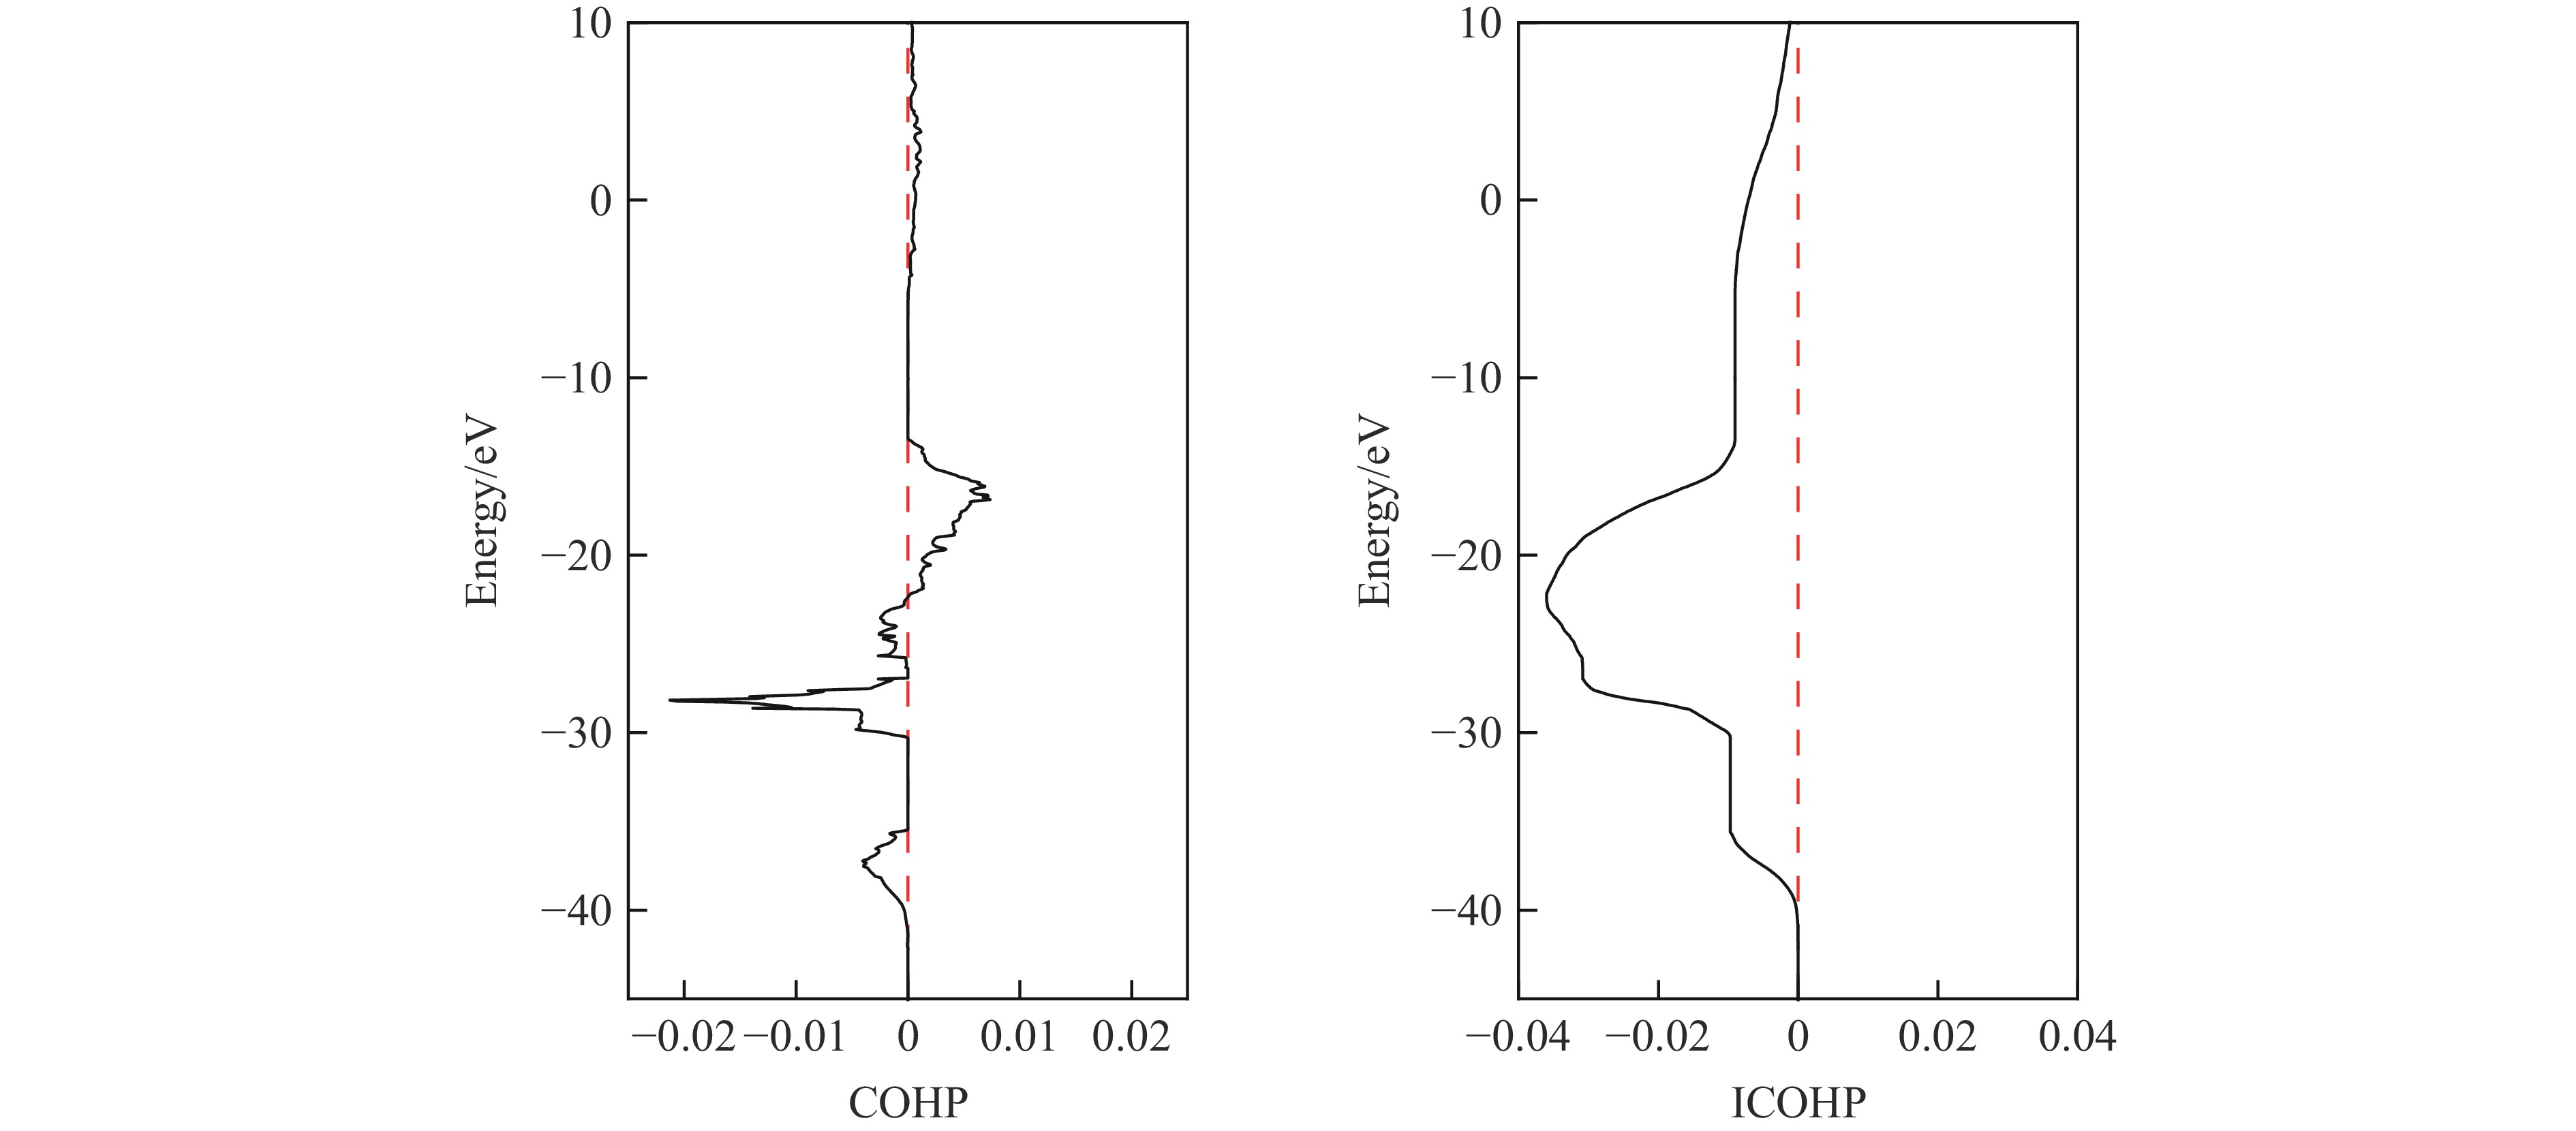
By combining first-principles calculations under the framework of density functional theory (DFT) and the CALYPSO crystal structure prediction method, the structural stability of the inert element helium (He) and alkaline-earth metals under high-pressure conditions has been systematically investigated. The calculations reveal that among the alkaline-earth metals, strontium (Sr) forms compounds with He exhibiting relatively low energy values. Consequently, the crystal structure of Sr2He at 400 GPa was predicted. Electron localization function (ELF) and density of states (DOS) analyses show no tendency for covalent bond formation between Sr and He atoms. Furthermore, Bader charge analysis reveals ionic bonding between Sr and He atoms, with charge transfer occurring from He to Sr. These results provide key insights into the bonding mechanism of Sr2He. This study elucidates the crystal structure, bonding nature, and electronic properties of Sr2He, offering theoretical support for understanding the stability and physical properties of such metastable materials and providing important guidance for their experimental synthesis.

First-Principles Study of the Crystal Structure and Elastic Properties of FeO2H2He under High Pressure
CHEN Haonan, XIONG Yanhao, LIU Haibo, XU Guomiao
2026, 40(11): 110108. doi: 10.11858/gywlxb.20261111
Abstract:
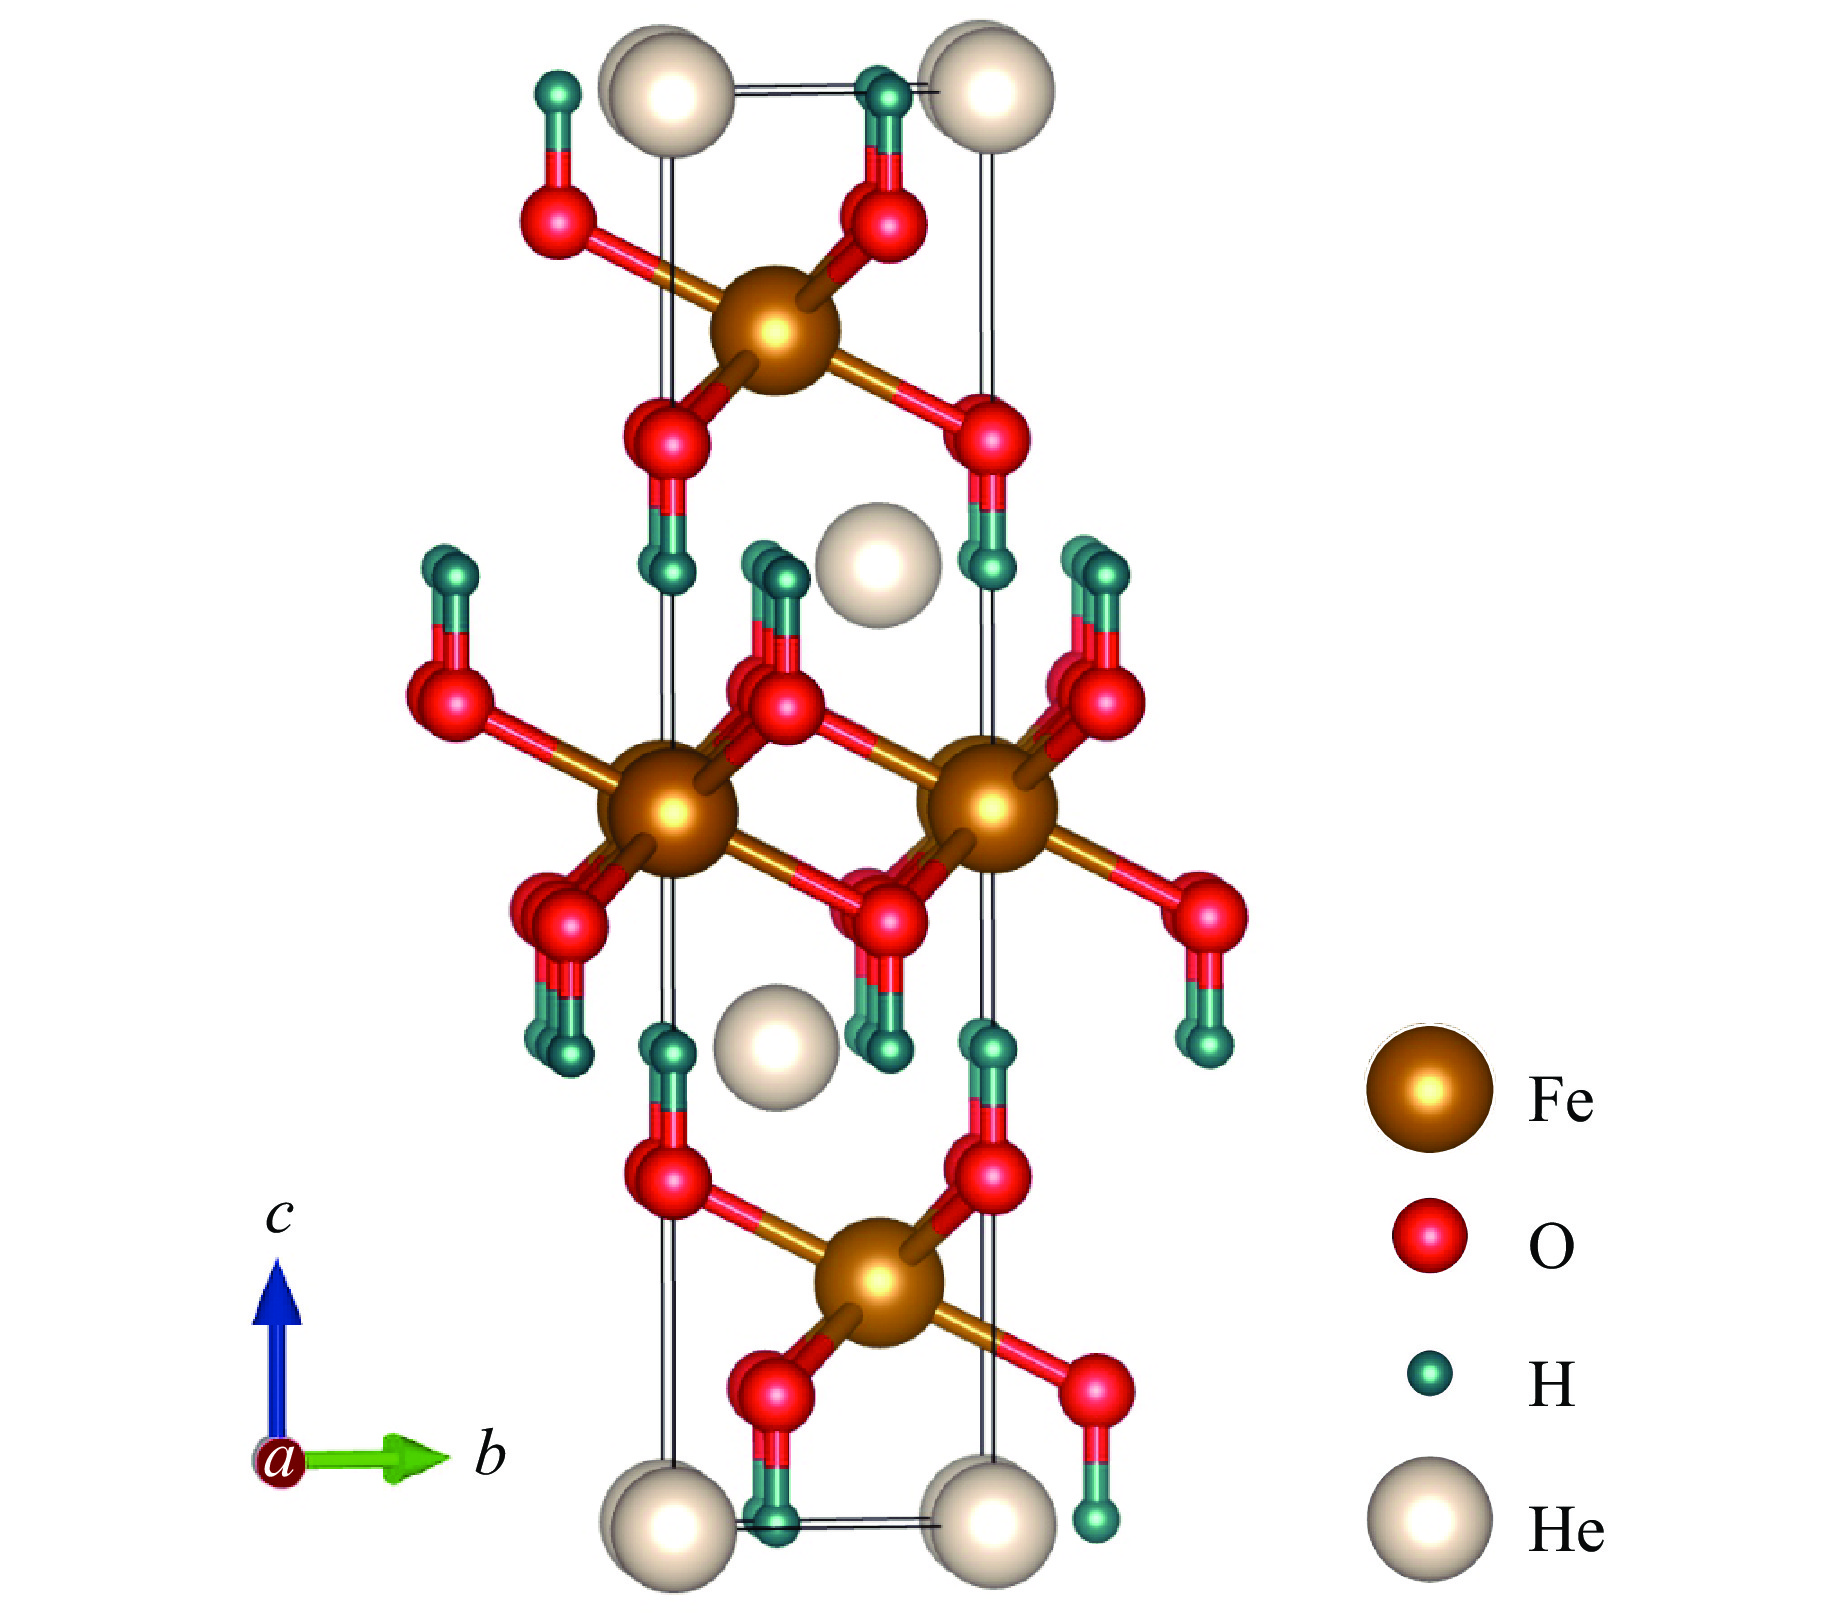
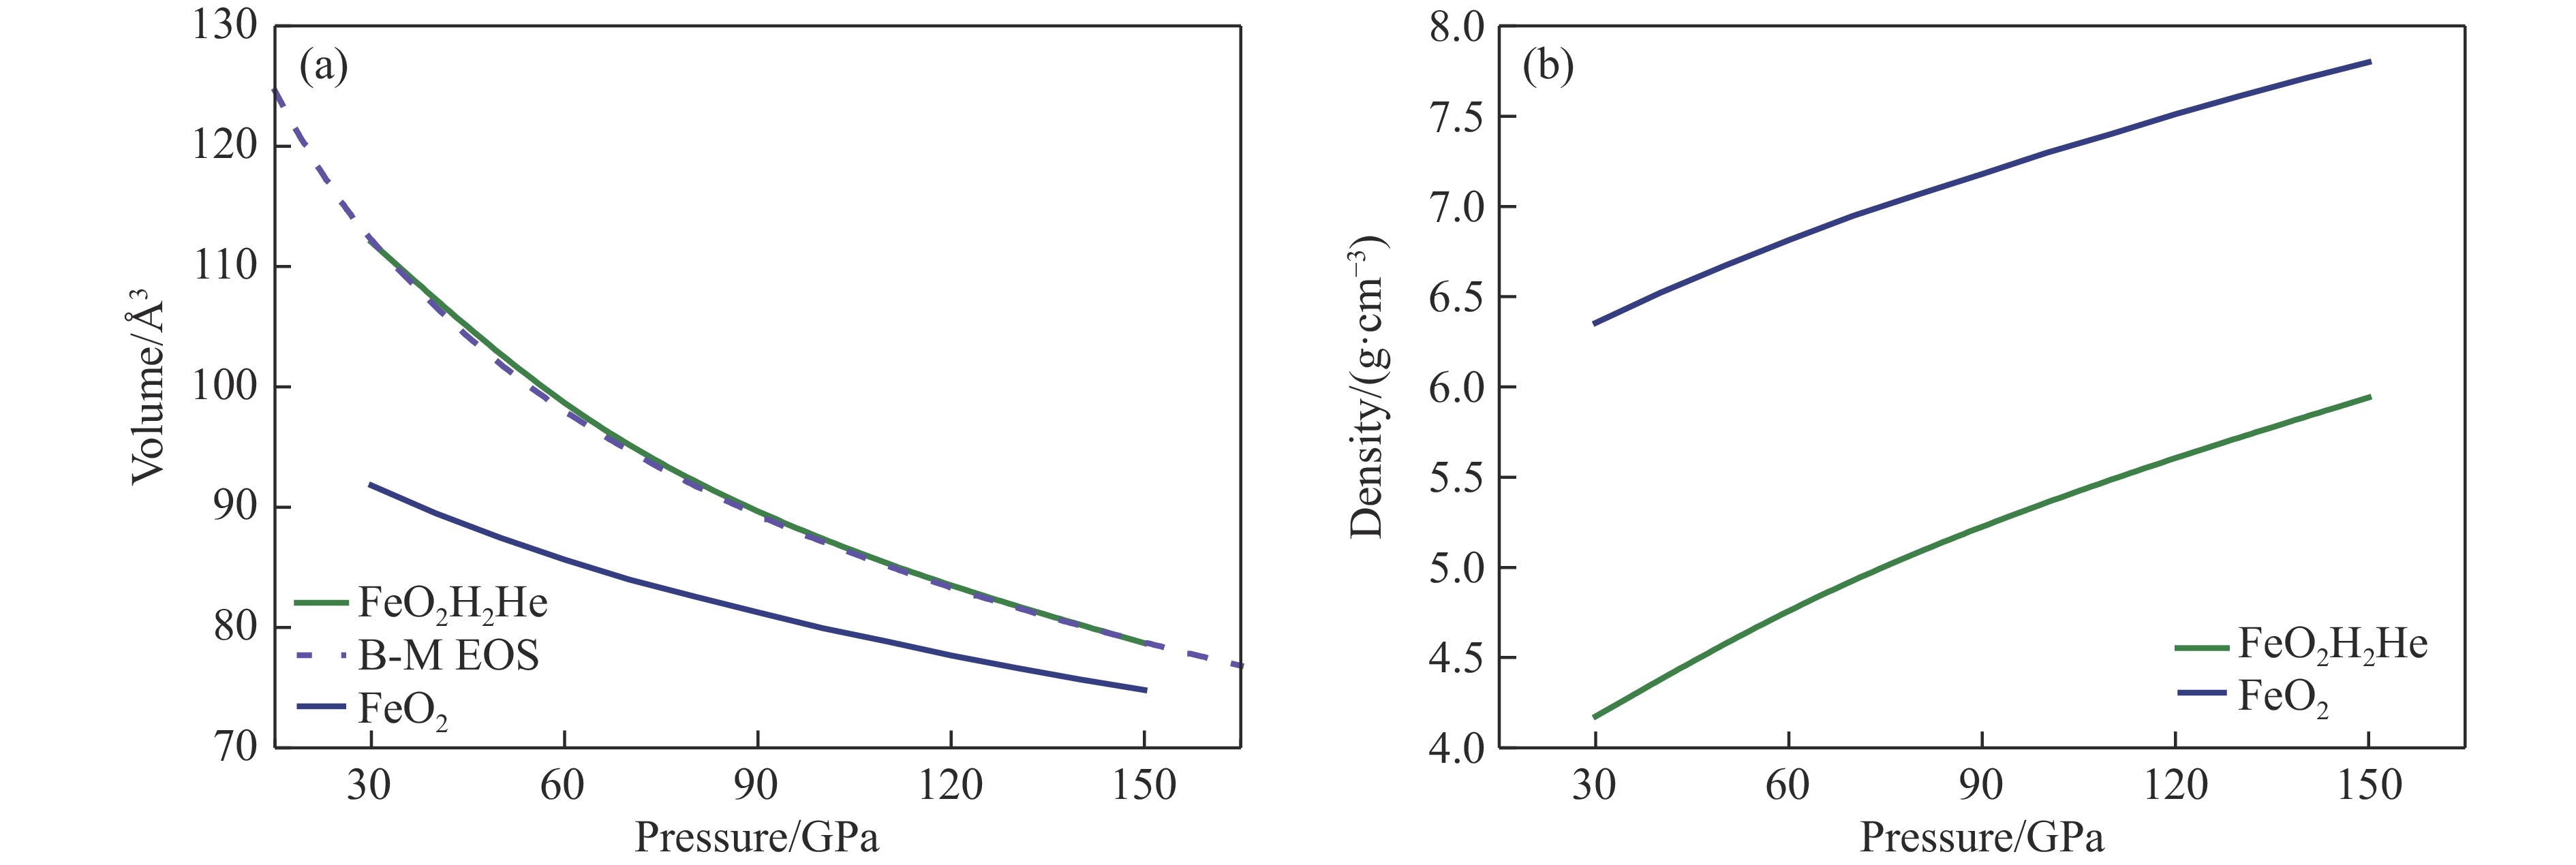
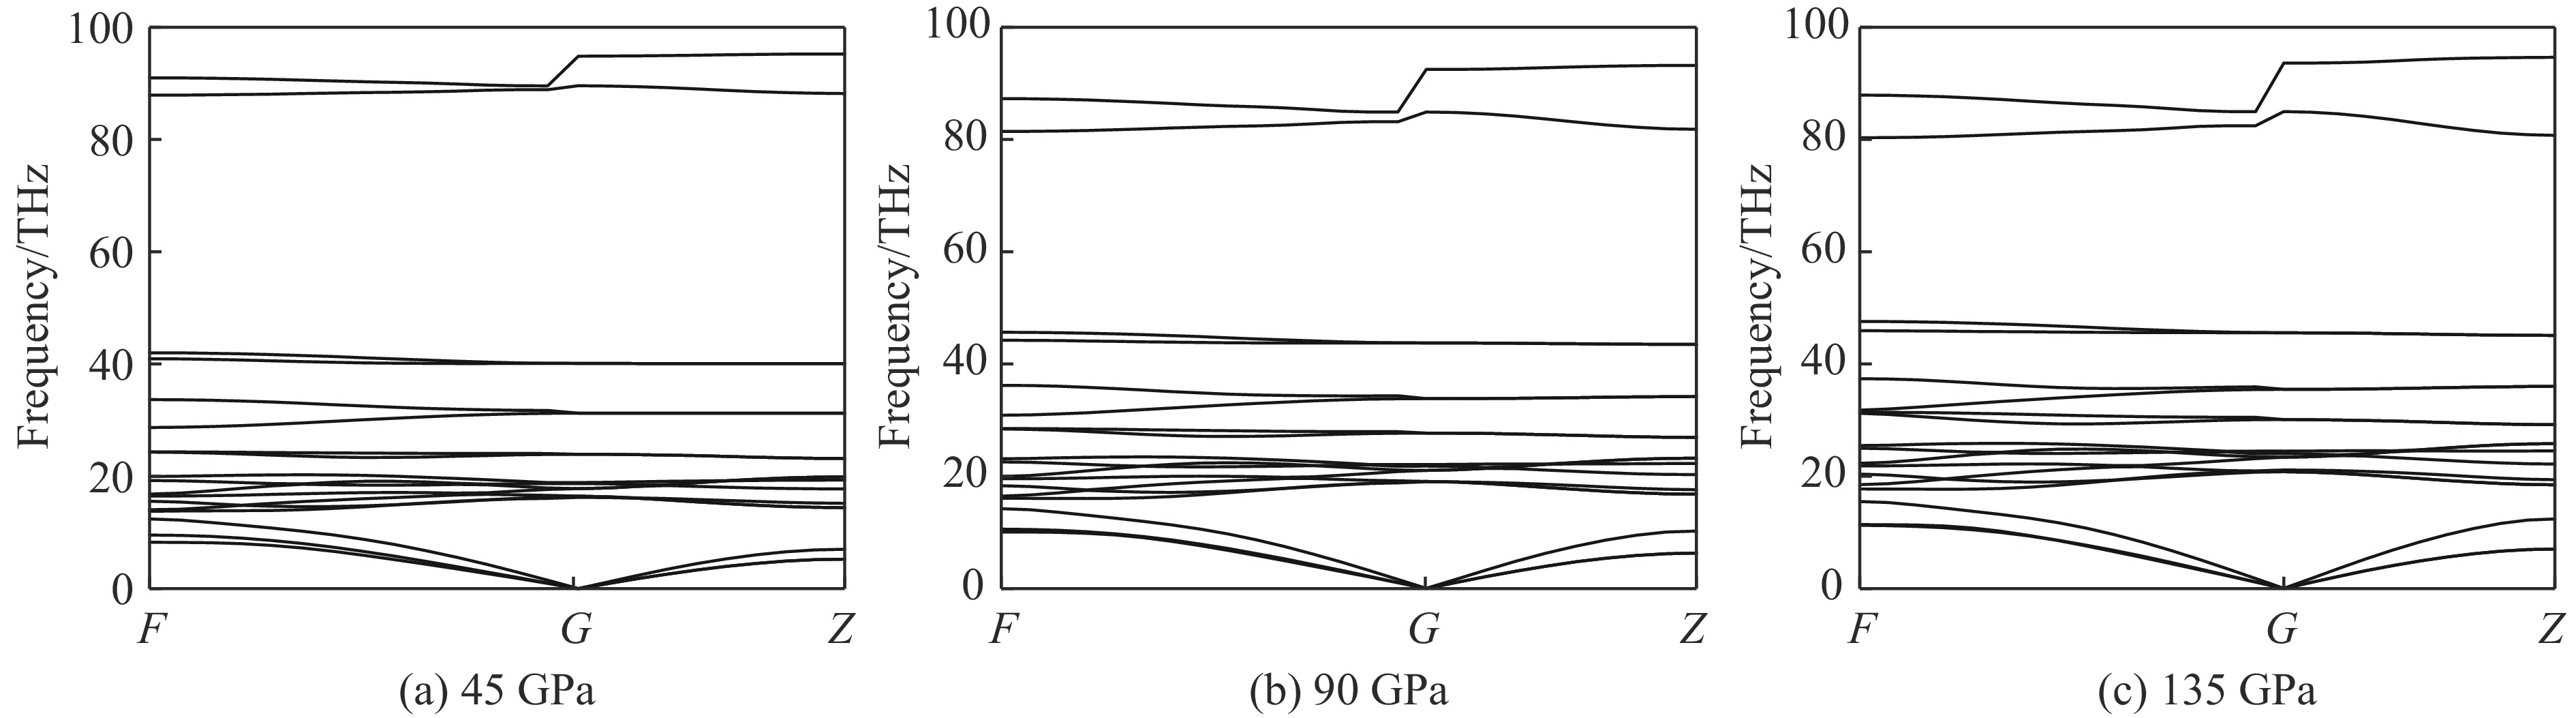
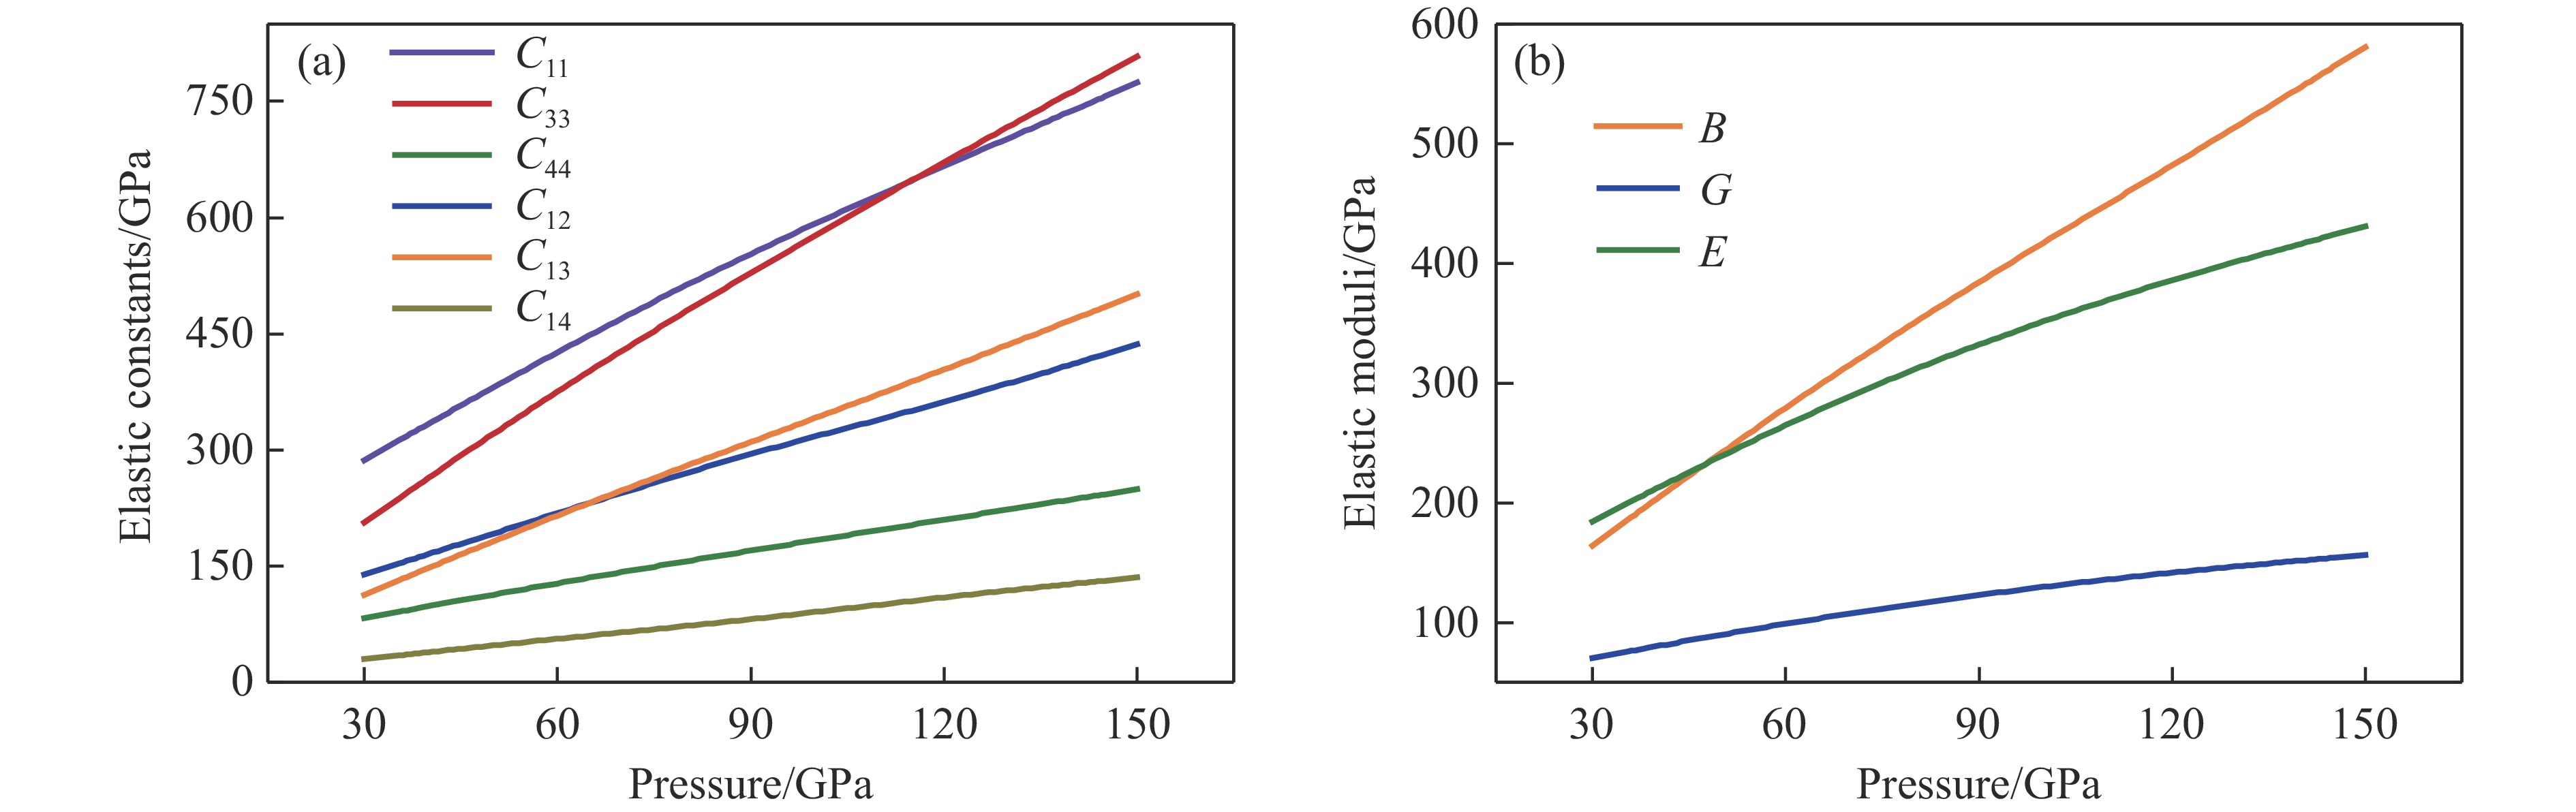
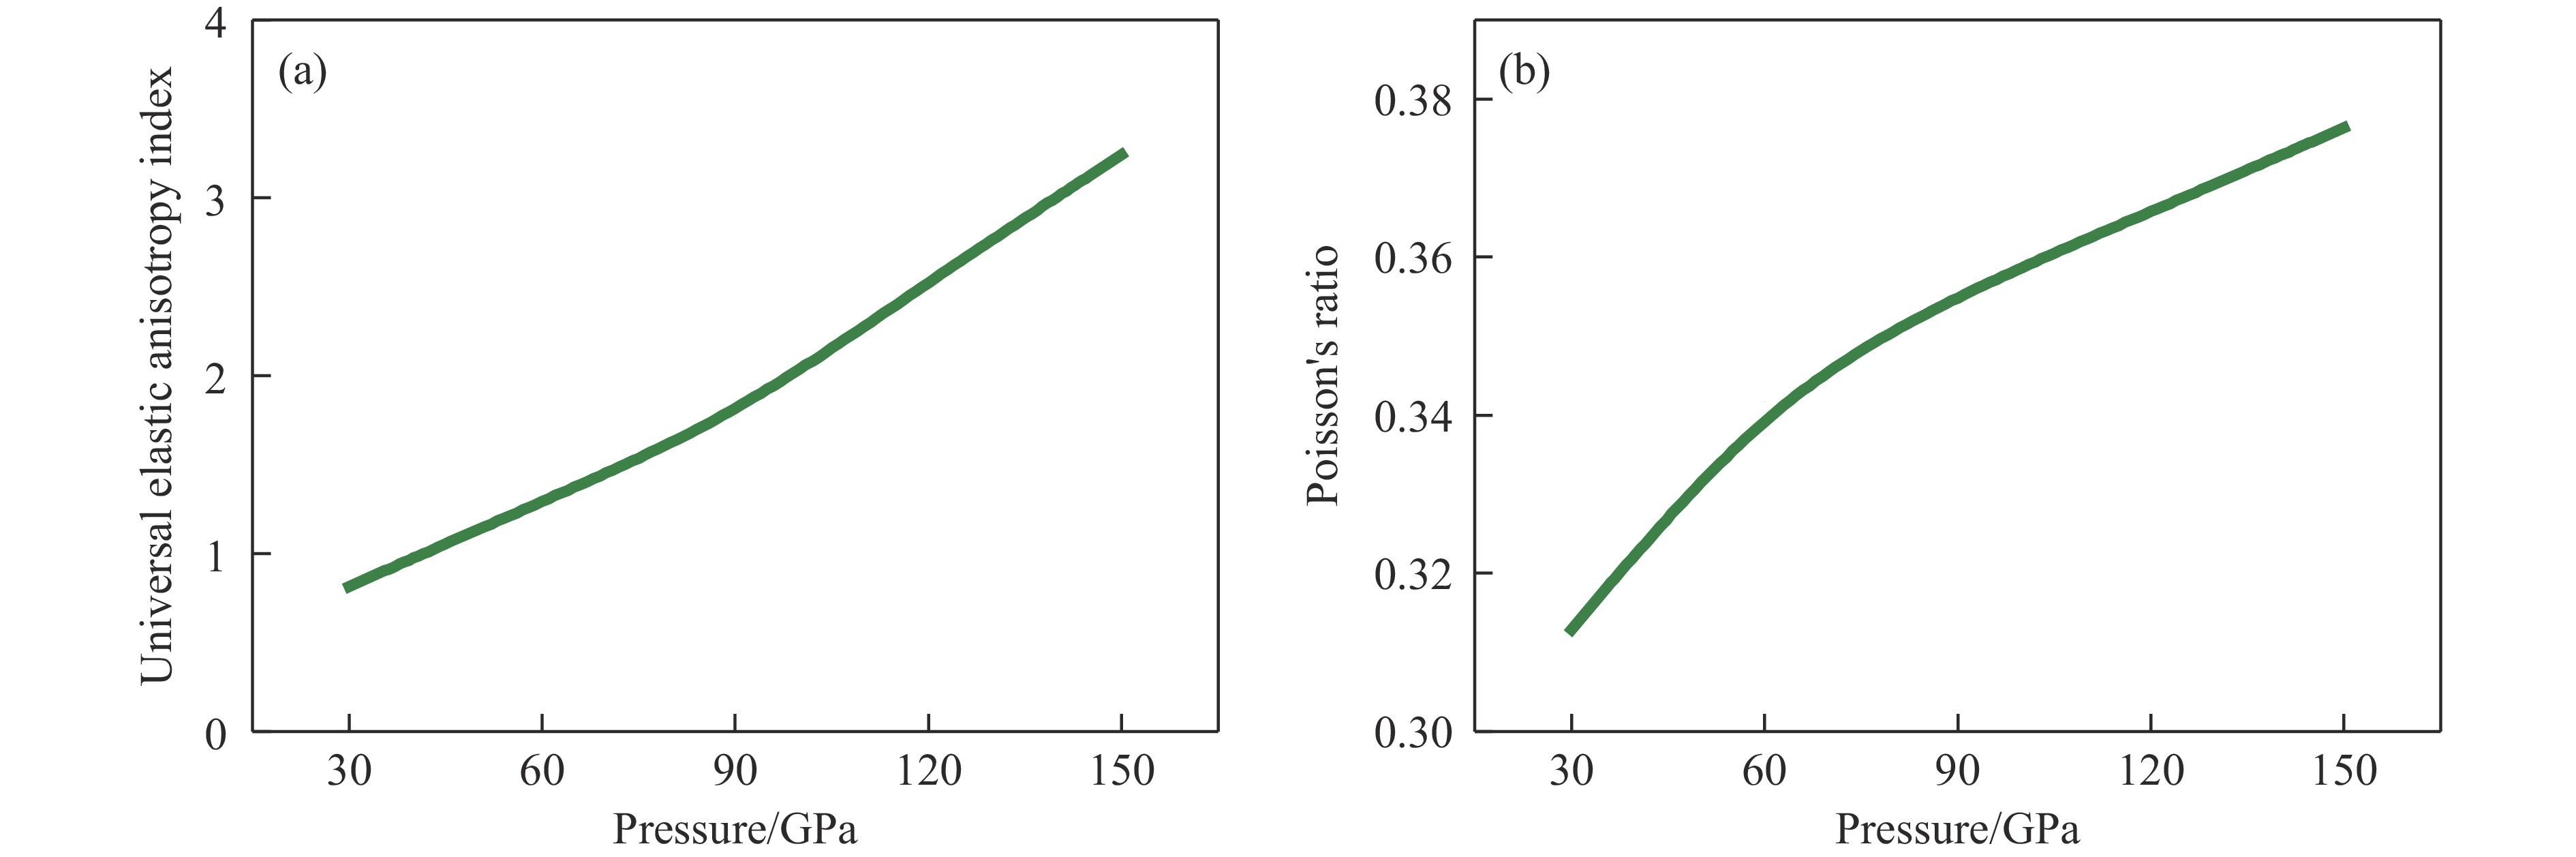
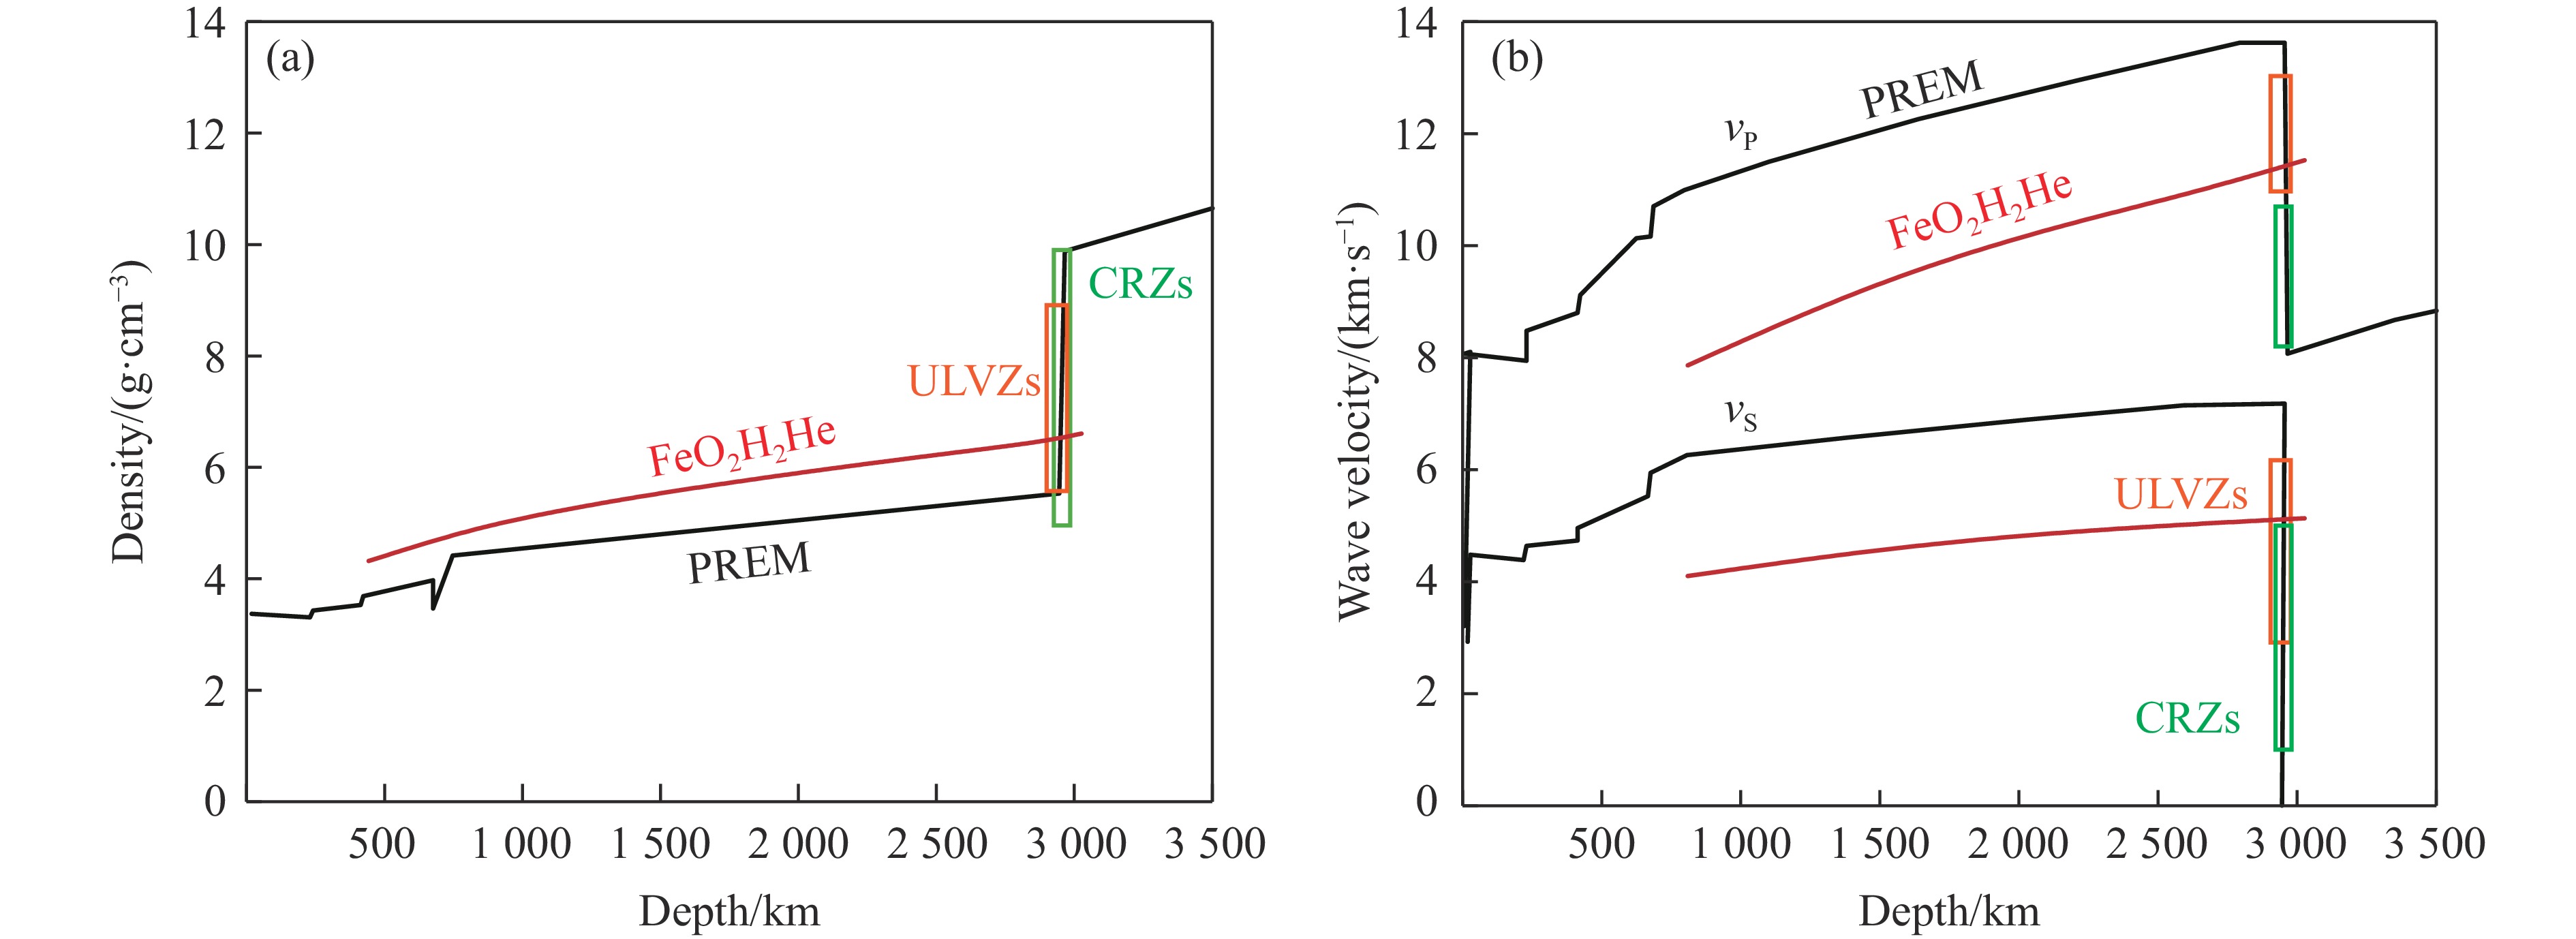
The composition and seismic anomalies of the deep Earth serve as key research topics, and the high 3He/4He ratio in ocean island basalts indicates the presence of the primordial helium reservoir in the Earth’s deep interior. However, our understanding of stable helium-bearing compounds in the deep Earth is still limited. This study systematically calculated the crystal structure and elastic properties of FeO2H2He at 30–150 GPa using first principles methods based on density functional theory. The static calculation results of phonon dispersion and elastic constants indicate that the R3m cubic crystal structure of FeO2H2He exhibits excellent local dynamic stability and mechanical stability within the studied pressure range. Poisson’s ratio and Pugh ratio analyses consistently show that FeO2H2He exhibits a ductile state throughout the investigated pressure range. The comprehensive characteristics of density and wave velocity reveal that FeO2H2He could potentially serve as one of the helium-bearing components near the core-mantle boundary, and may represent a potential mineral component for explaining the wave velocity anomalies observed in ultra-low velocity zones (ULVZs). This research enriches our understanding of the core-mantle boundary composition and provides new mineralogical constraints on the occurrence and distribution of helium in the deep Earth.

Structural Phase Transitions and Physical Properties of TaB4 under High Pressure
SHI Limin, YIN Chaoyi, CHEN Xin
2026, 40(11): 110109. doi: 10.11858/gywlxb.20261181
Abstract:
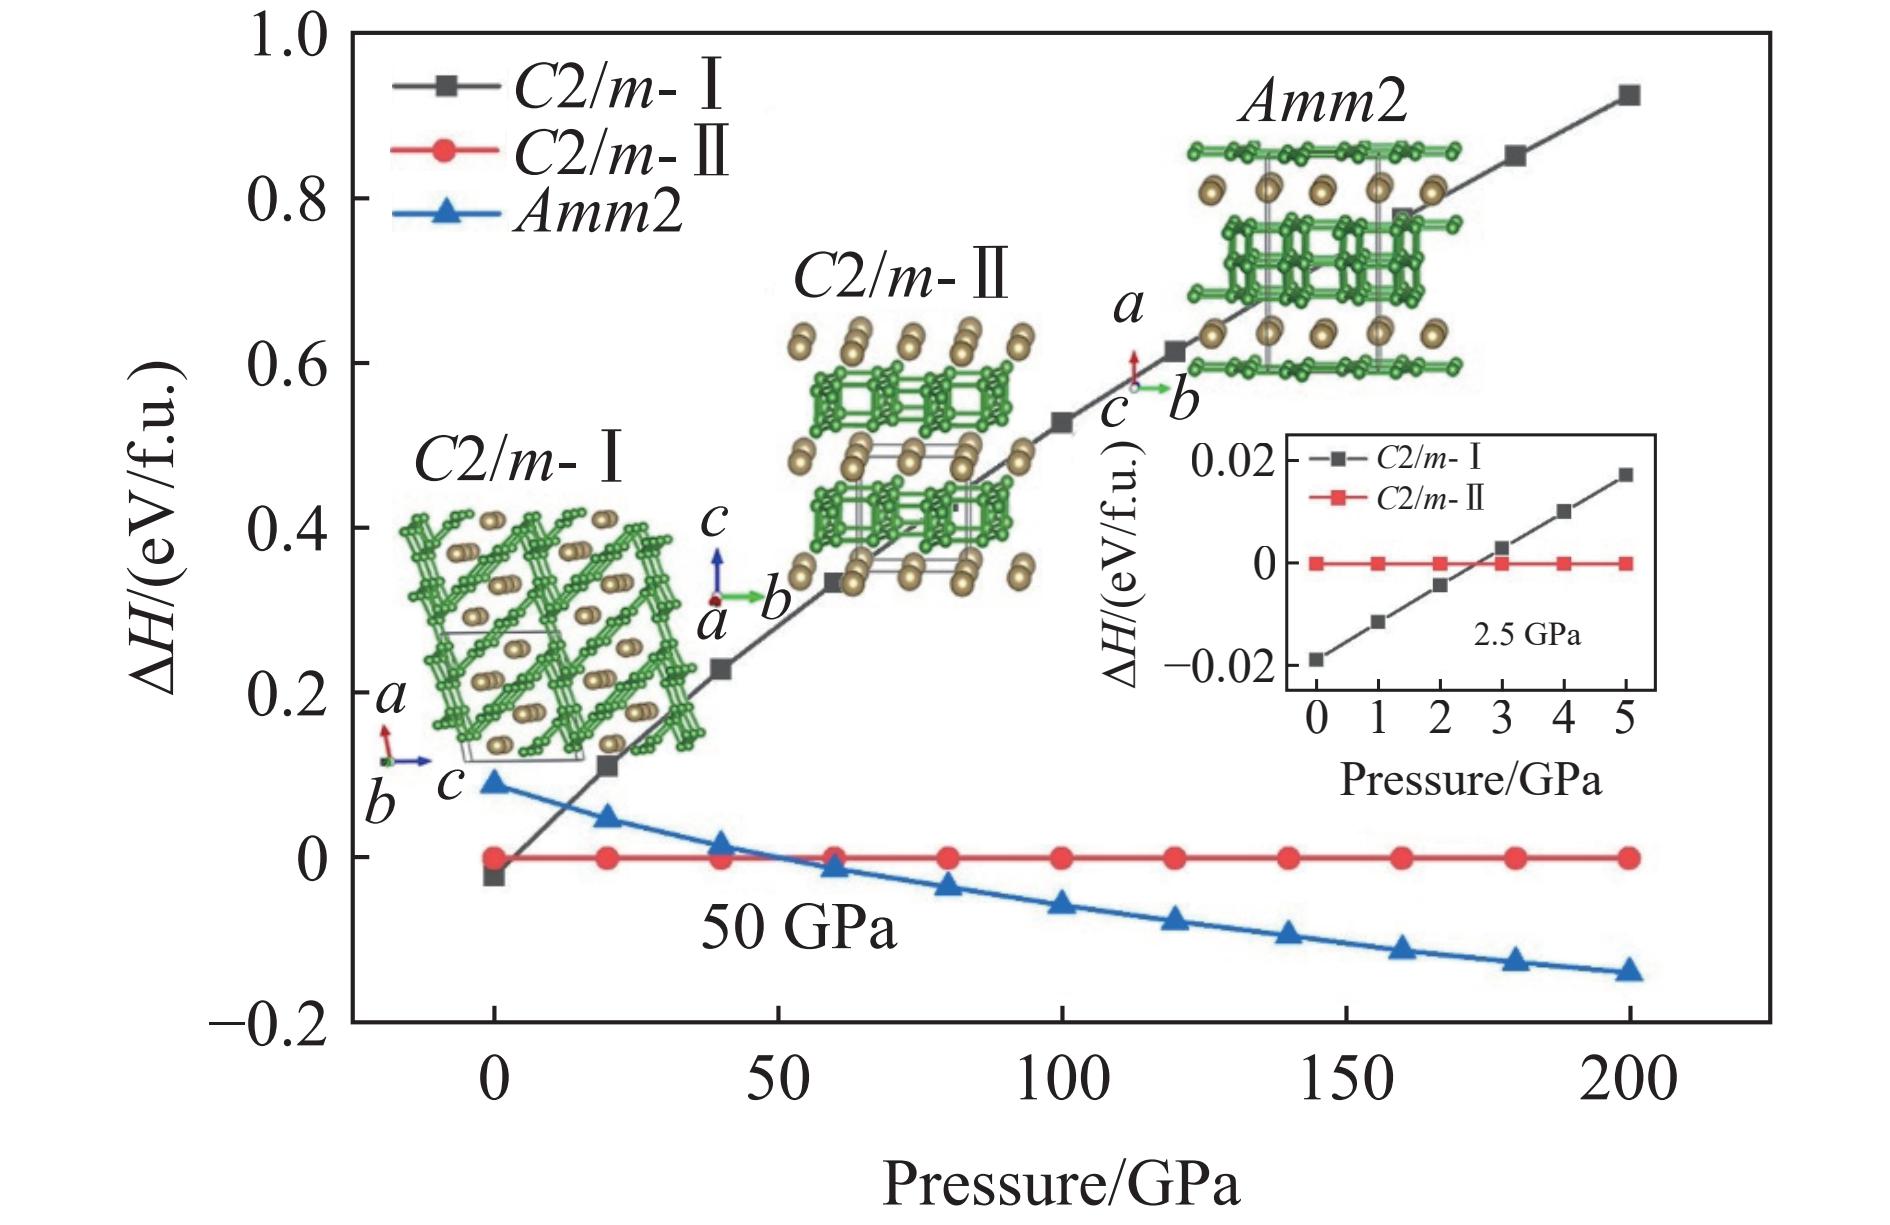
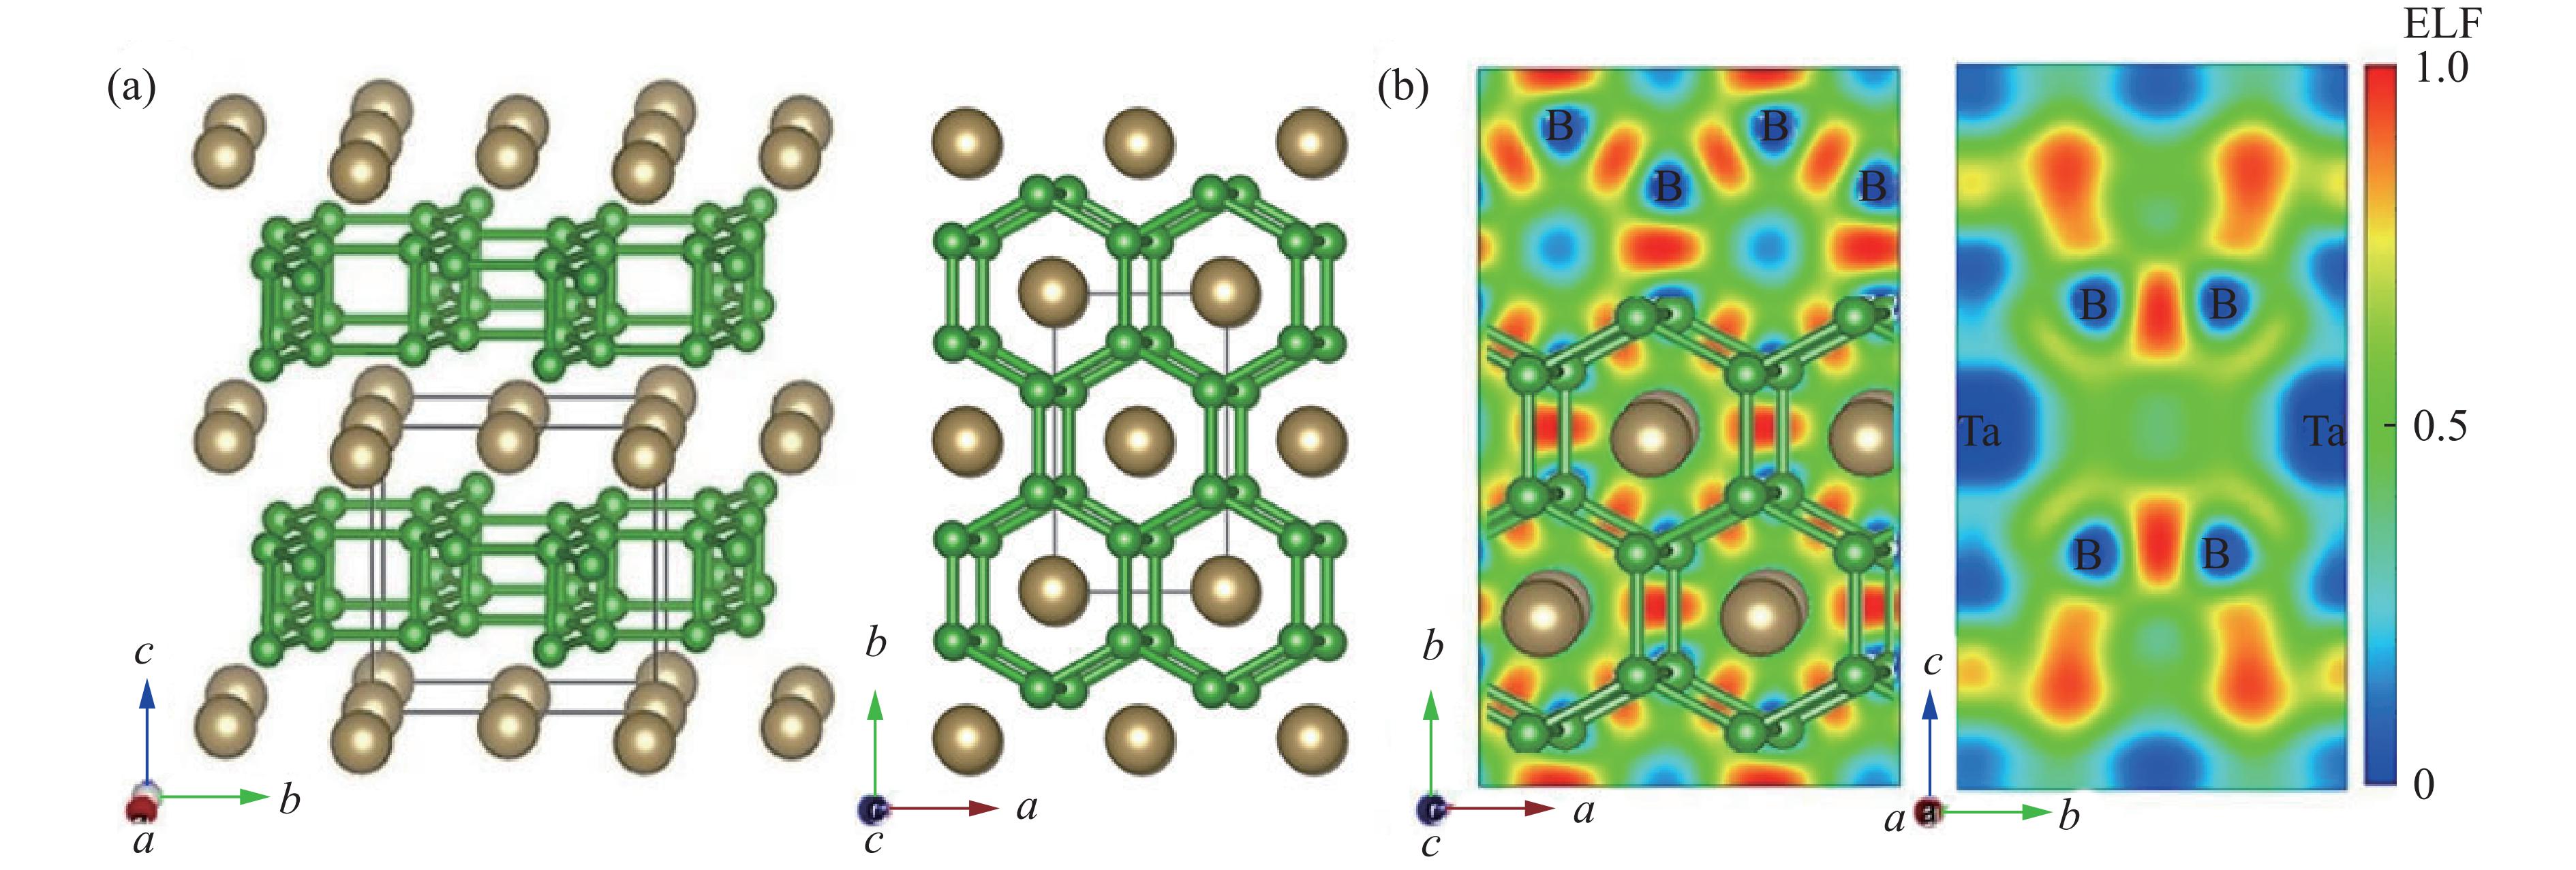
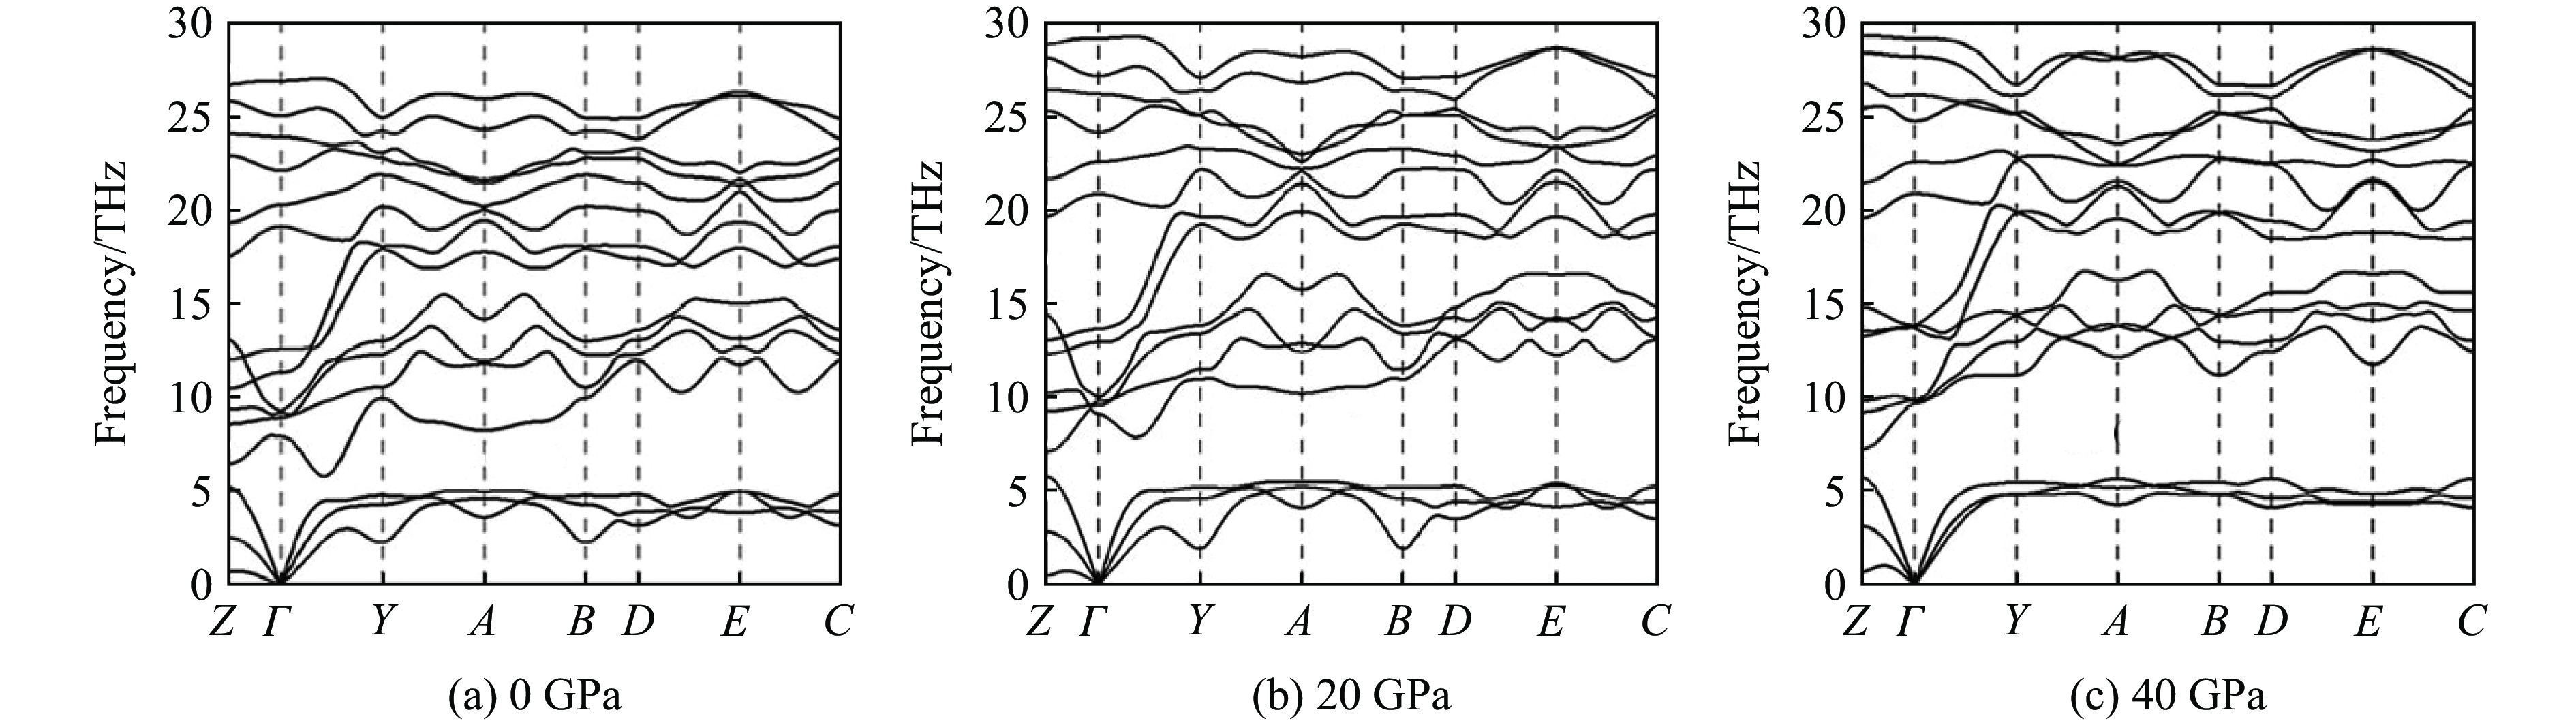
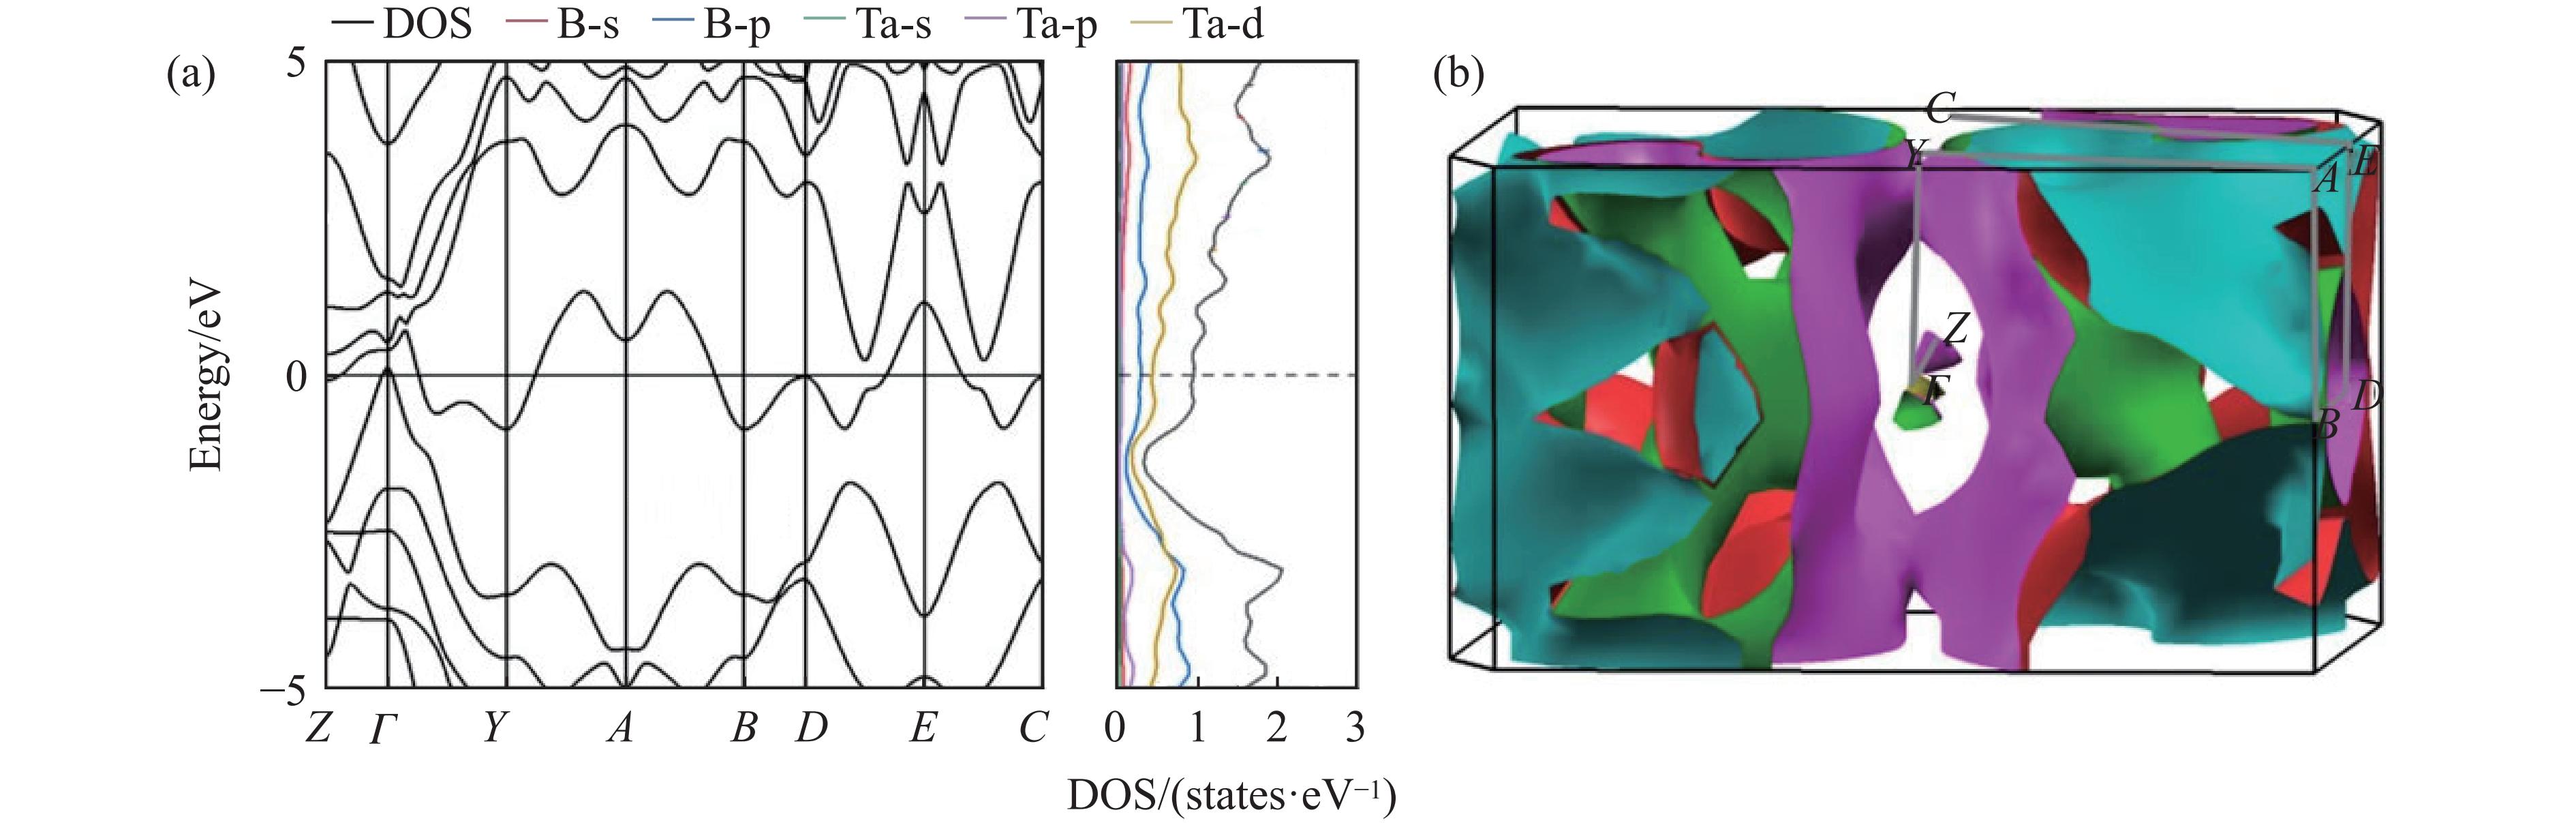
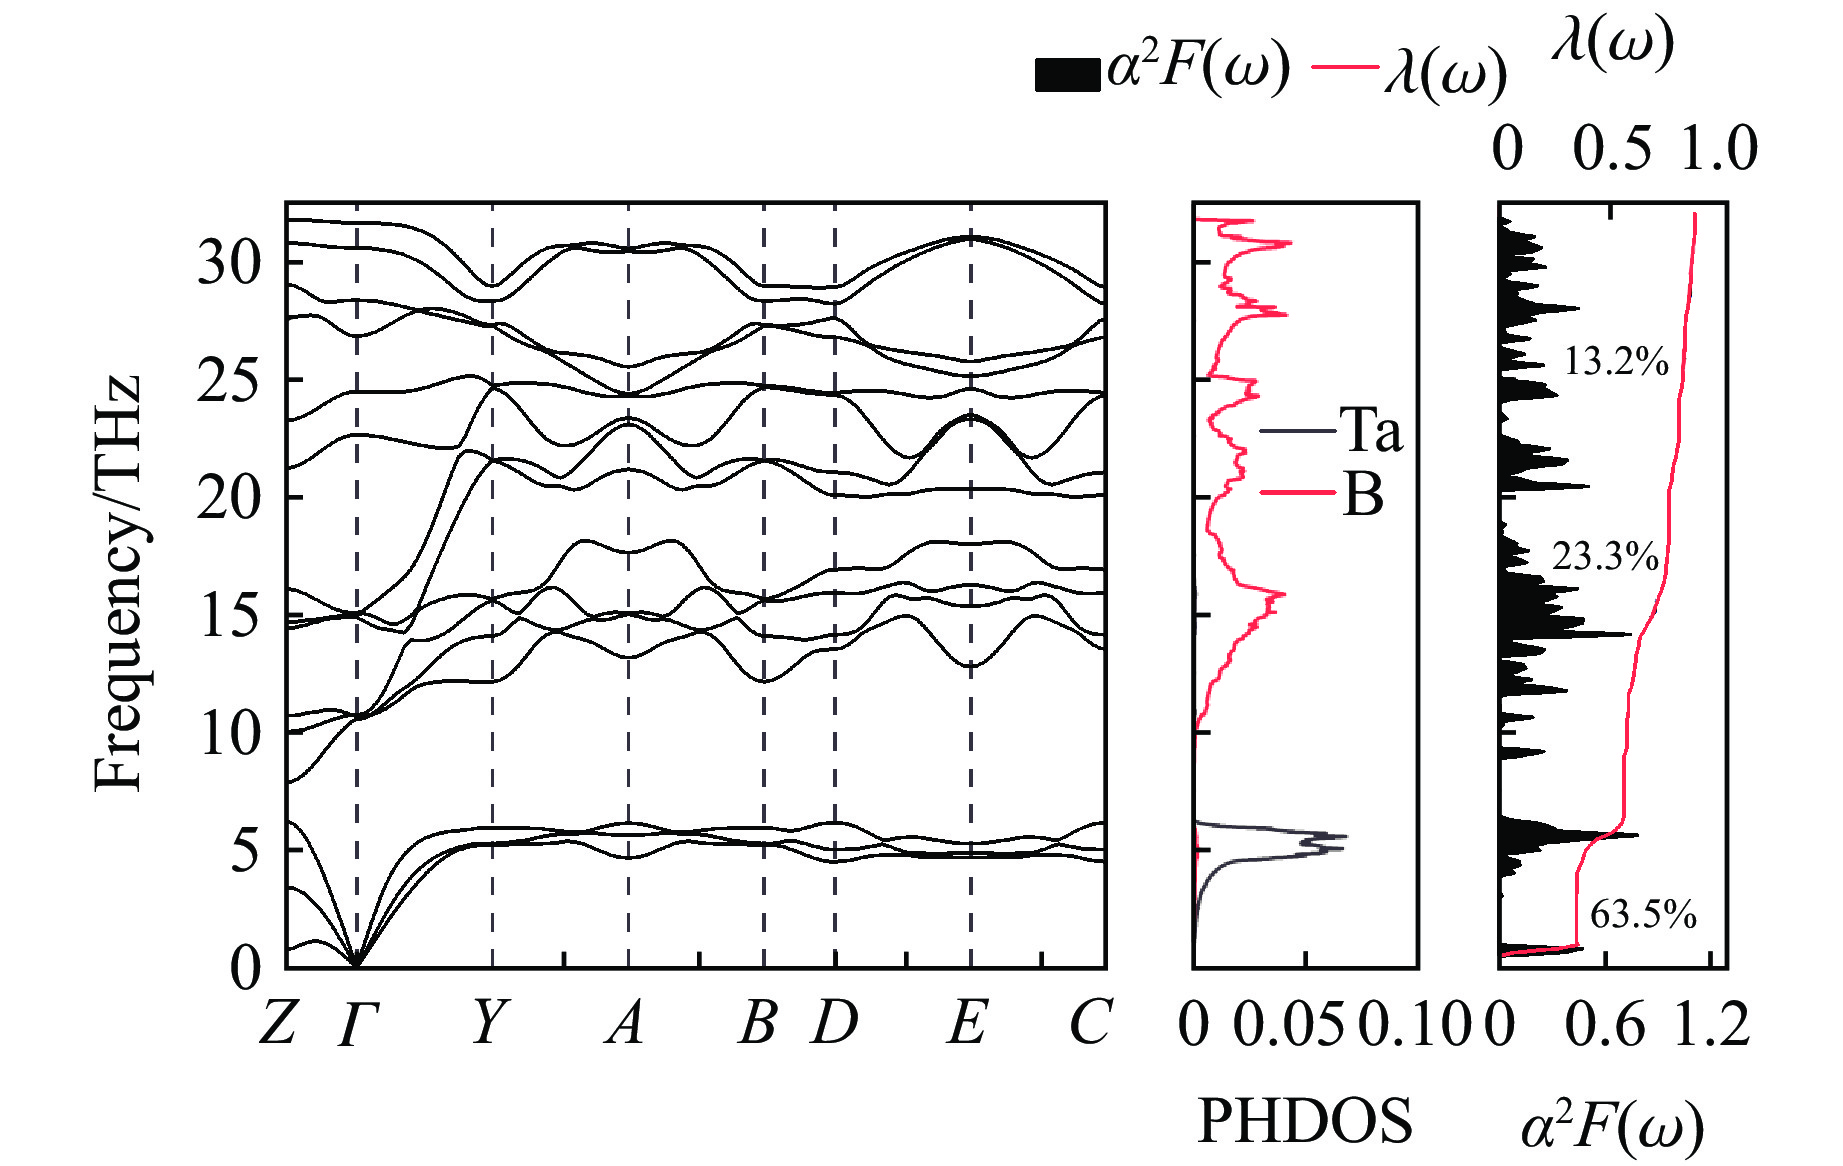
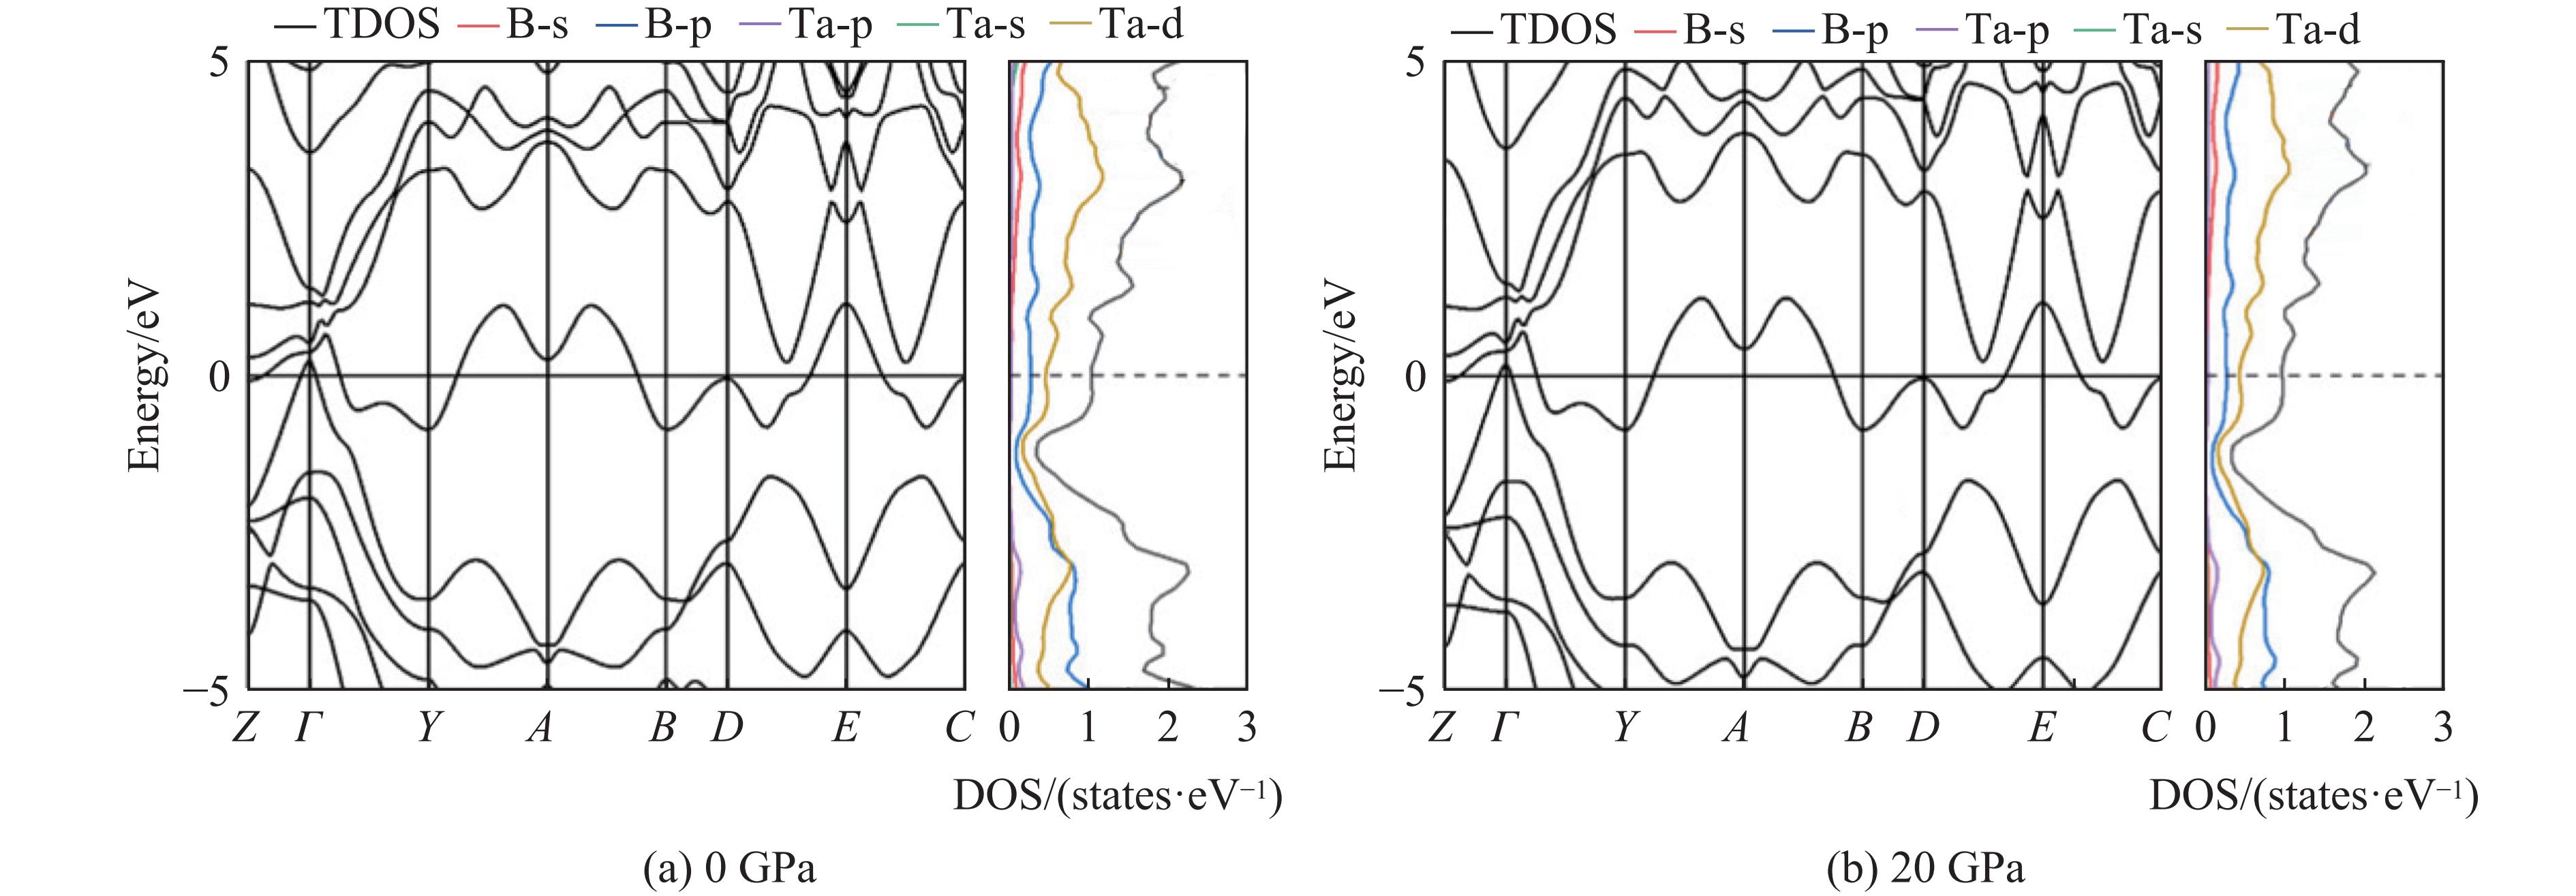
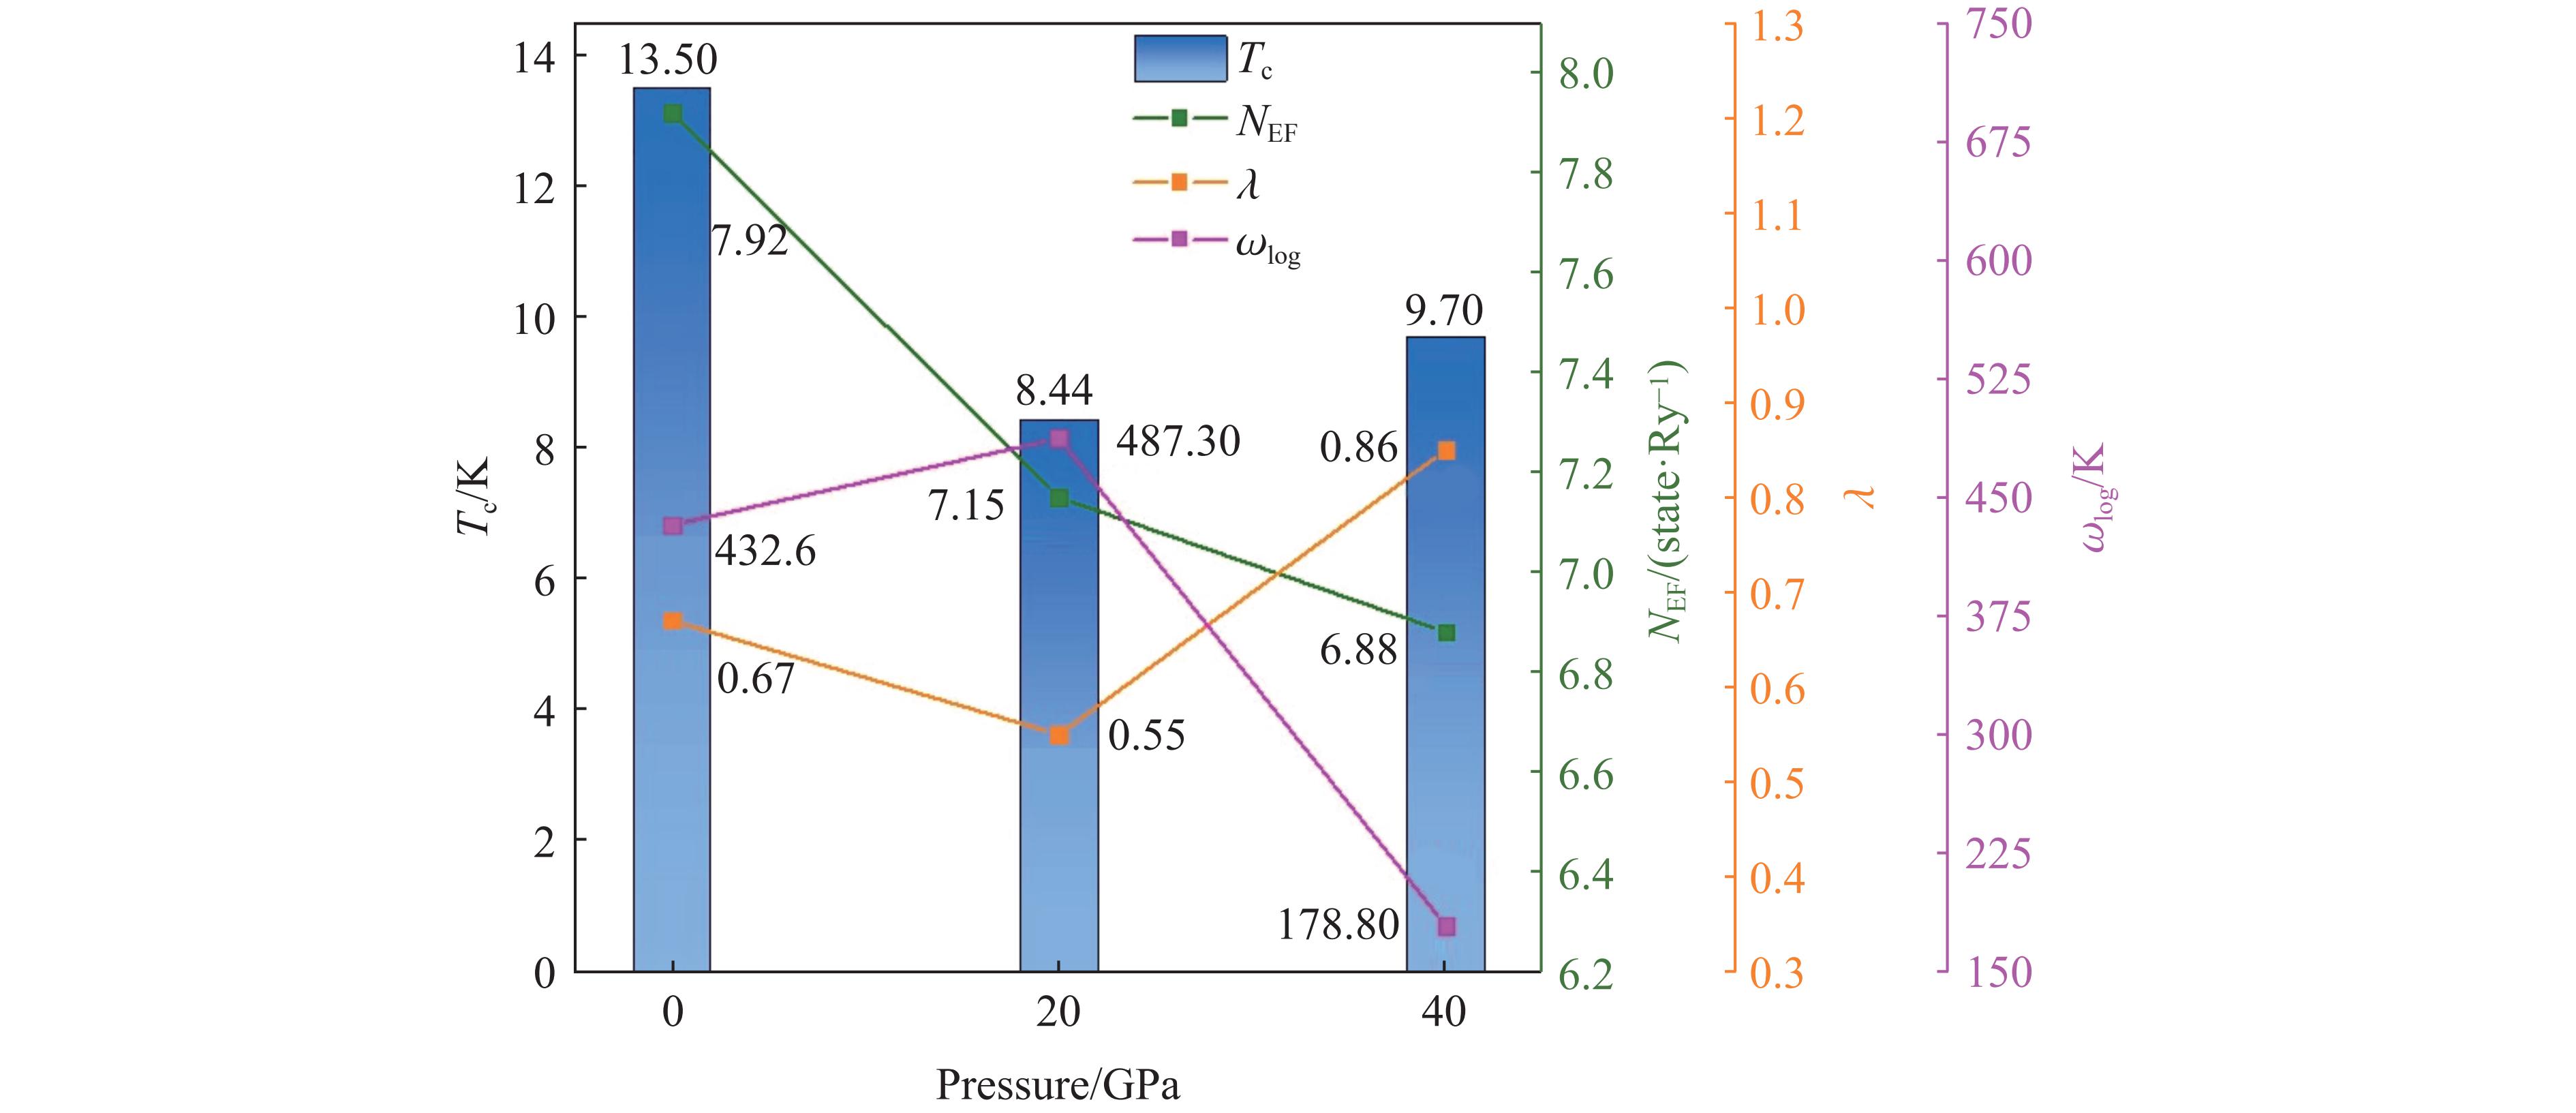
Crystal structure searches based on the particle swarm optimization algorithm combined with first-principles calculations were performed to systematically investigate the crystal structures, stability, electronic properties, mechanical properties, and superconductivity of the boron-rich transition-metal boride TaB4 under high pressure, with particular emphasis on the intrinsic relationship between crystal structure and superconductivity. The results show that the C2/m-Ⅰ phase, which is stable at ambient pressure, undergoes a structural phase transition at approximately 2.5 GPa, giving rise to a new monoclinic C2/m-Ⅱ phase. Phonon calculations reveal that this new phase remains dynamically stable after decompression to ambient pressure, suggesting the possibility of its metastable existence under ambient conditions. Electronic structure calculations indicate that C2/m-Ⅱ-TaB4 exhibits metallic behavior. Electron-phonon coupling calculations further demonstrate that this phase exhibits phonon-mediated superconductivity, with a superconducting transition temperature of up to 13.5 K at ambient pressure. The electron-phonon coupling is mainly associated with low-frequency phonon modes involving Ta atoms. Mechanical property calculations show that C2/m-Ⅱ-TaB4 has a Vickers hardness of approximately 14.5 GPa, indicating a moderate hardness. These results expand our understanding of the structural diversity of boron-rich compounds in the Ta-B system and provide a theoretical basis for elucidating the intrinsic relationships among crystal structure, lattice vibrations, and superconductivity in transition-metal borides.

Novel Non-Diamond Isoelectronic B-C-O Metastable Phases under High Pressure: B2CO5 and B3CO3
LIU Dan, GAO Pengyue
2026, 40(11): 110110. doi: 10.11858/gywlxb.20261148
Abstract:
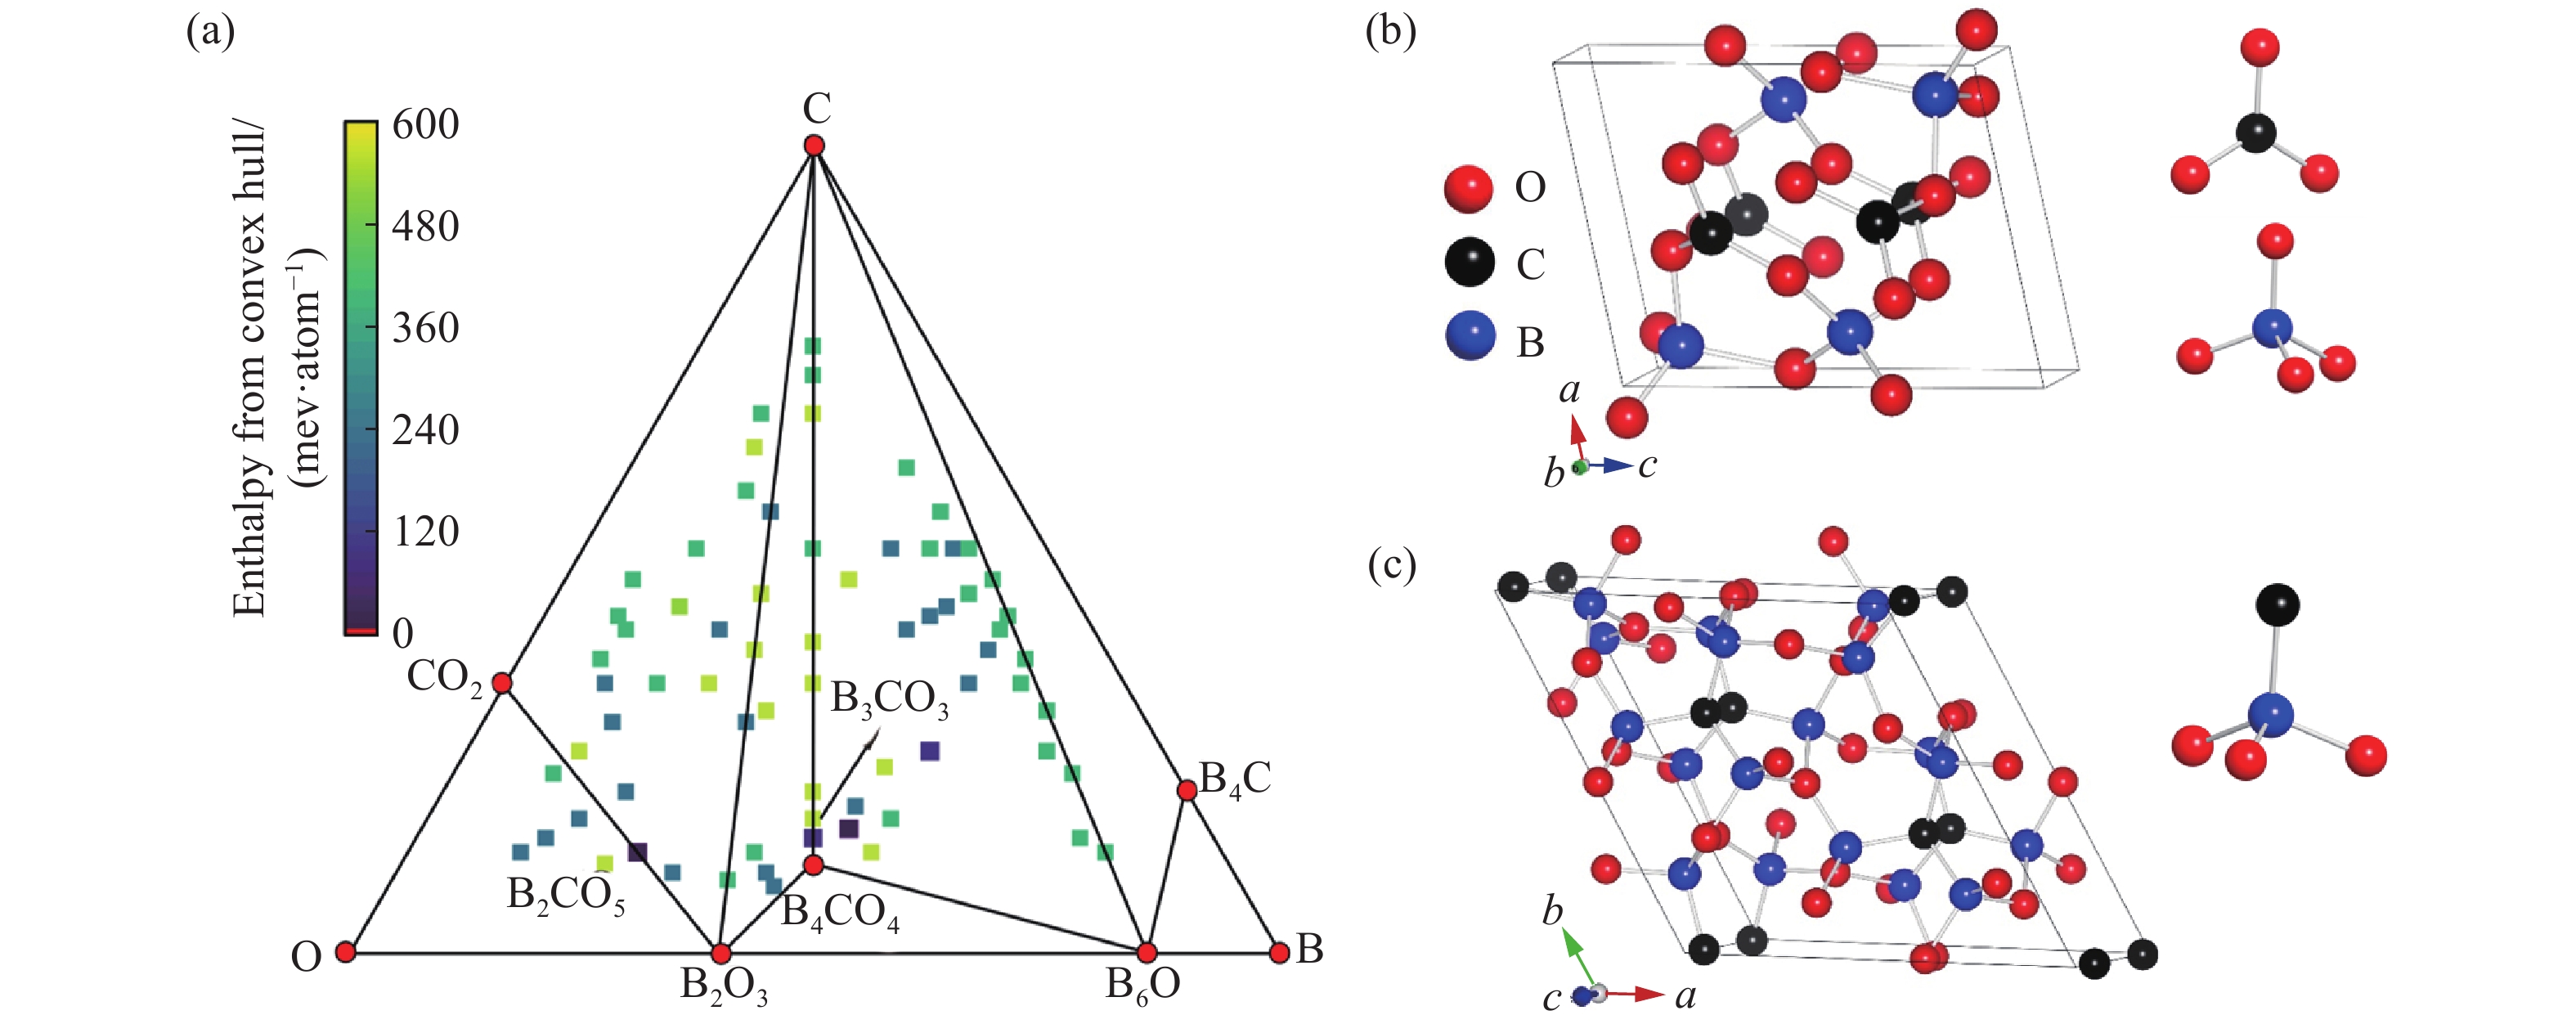
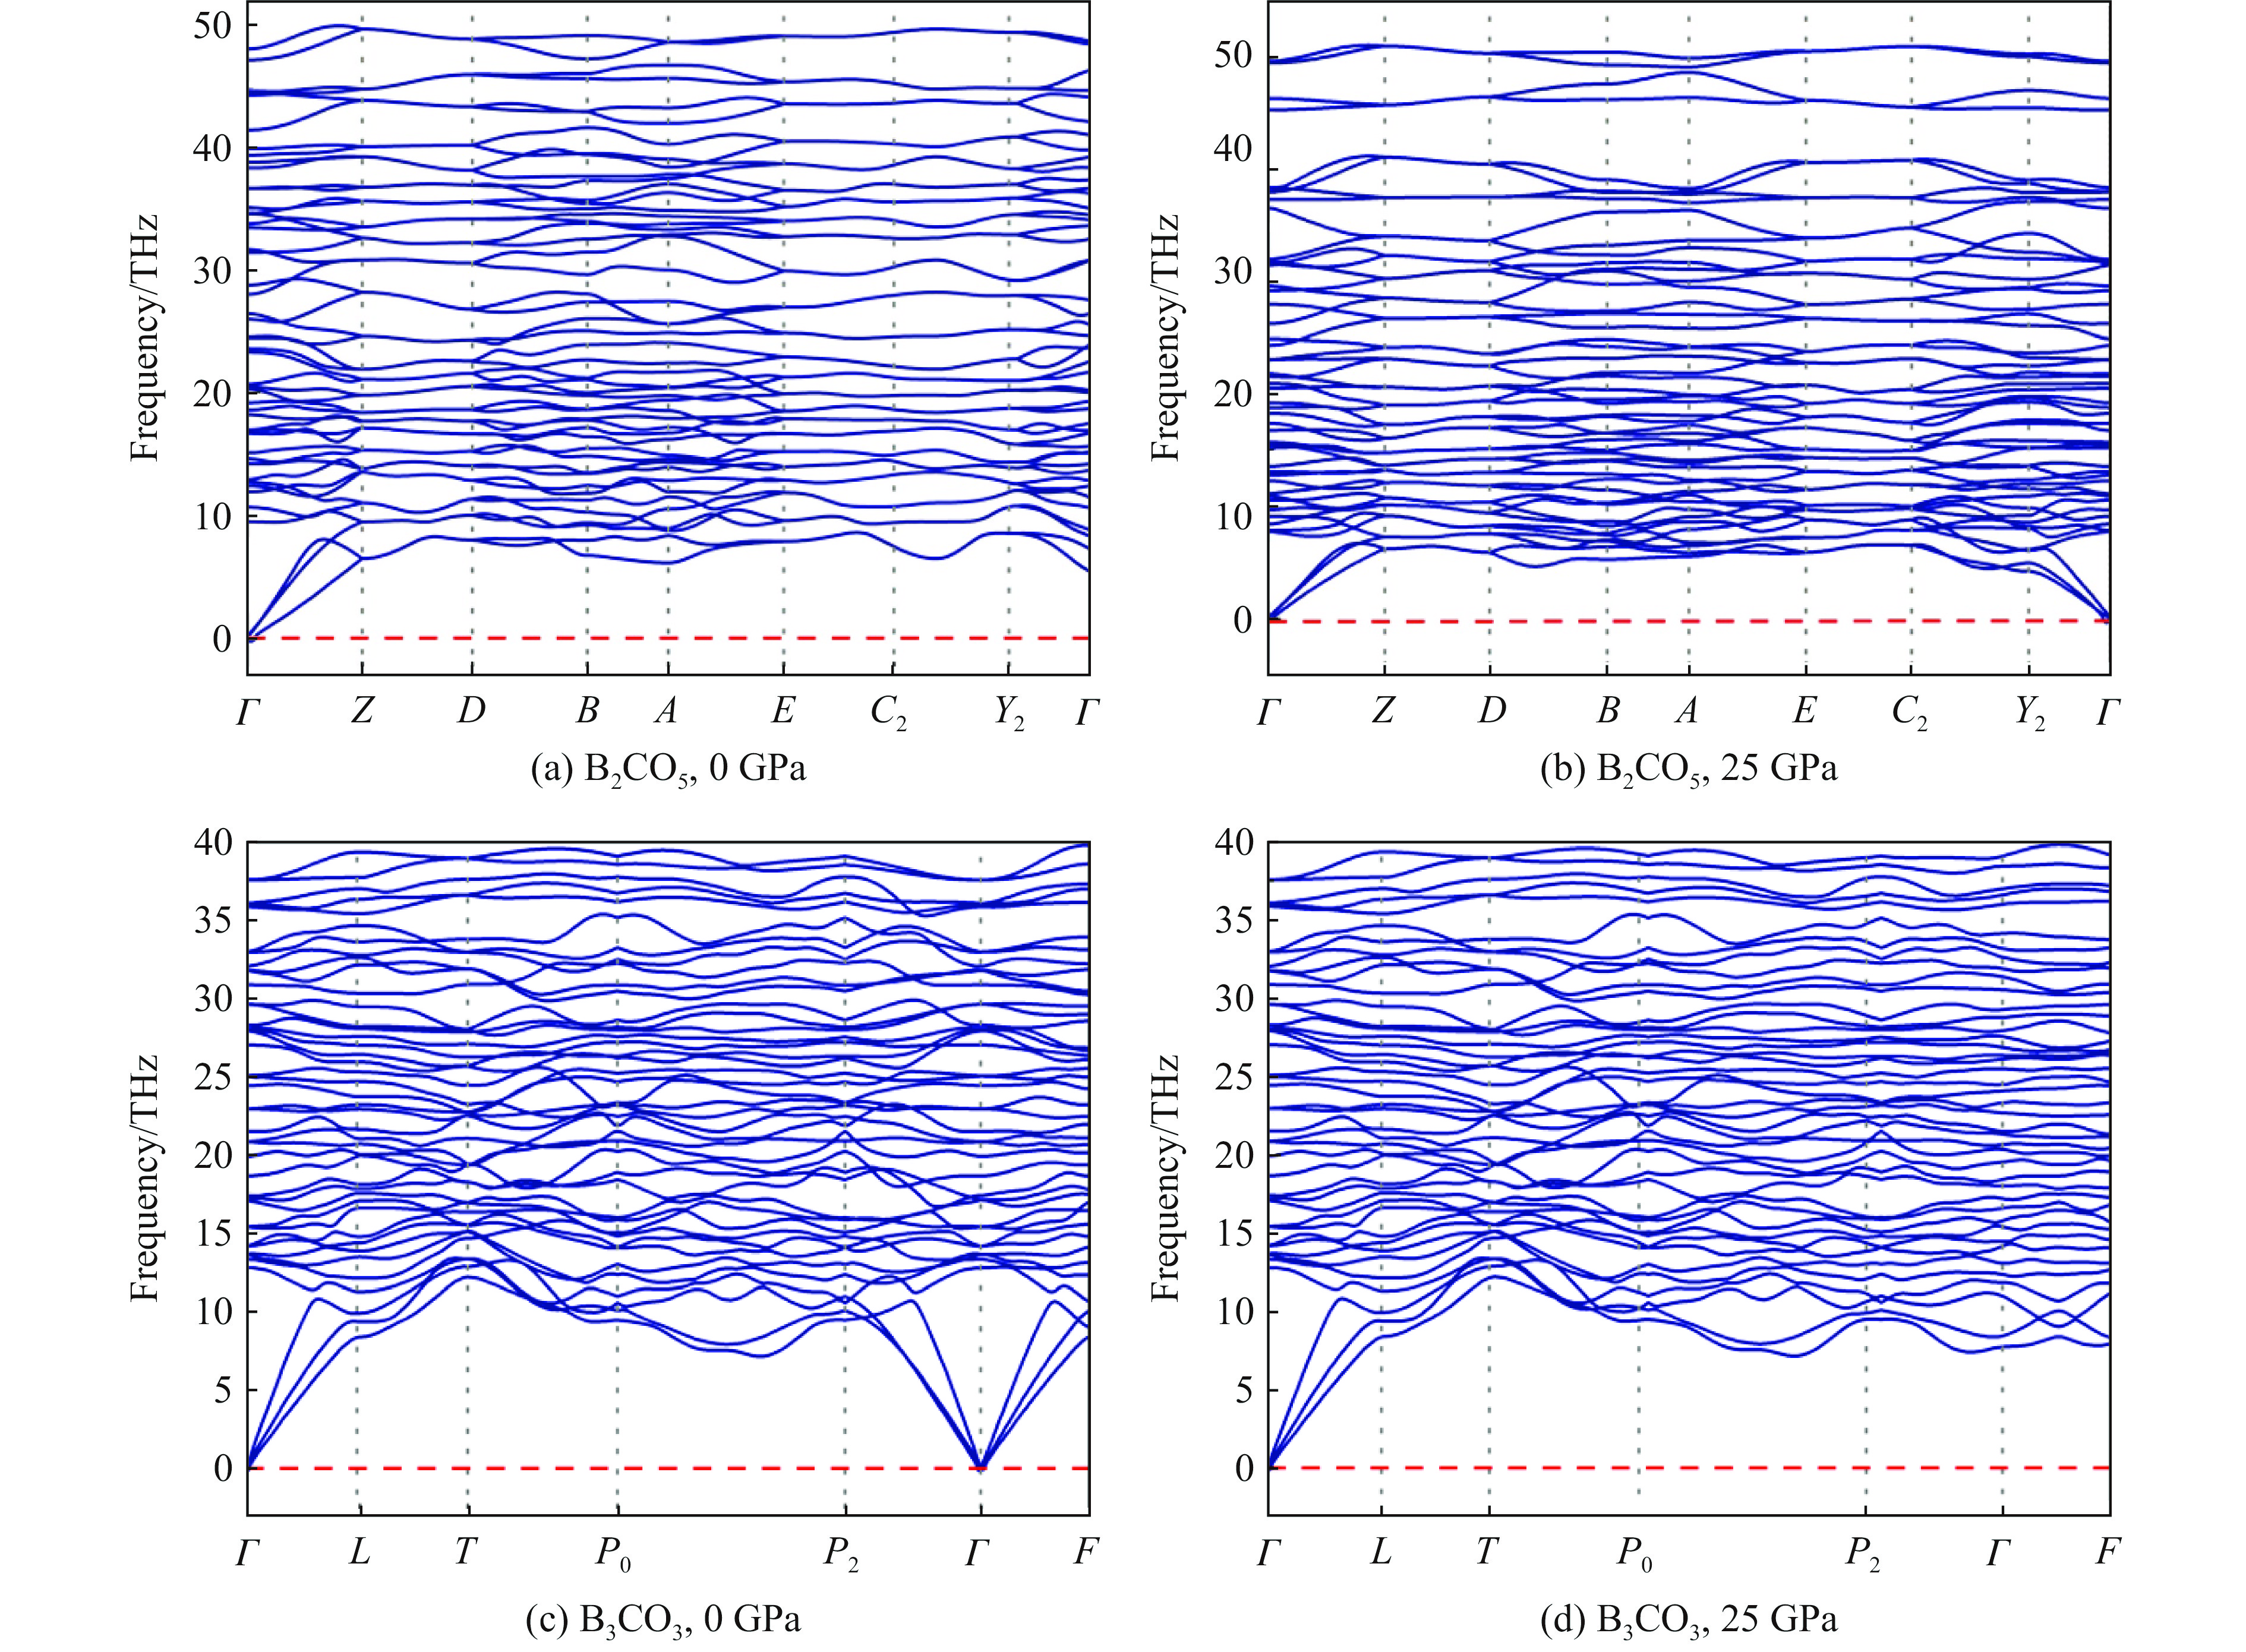
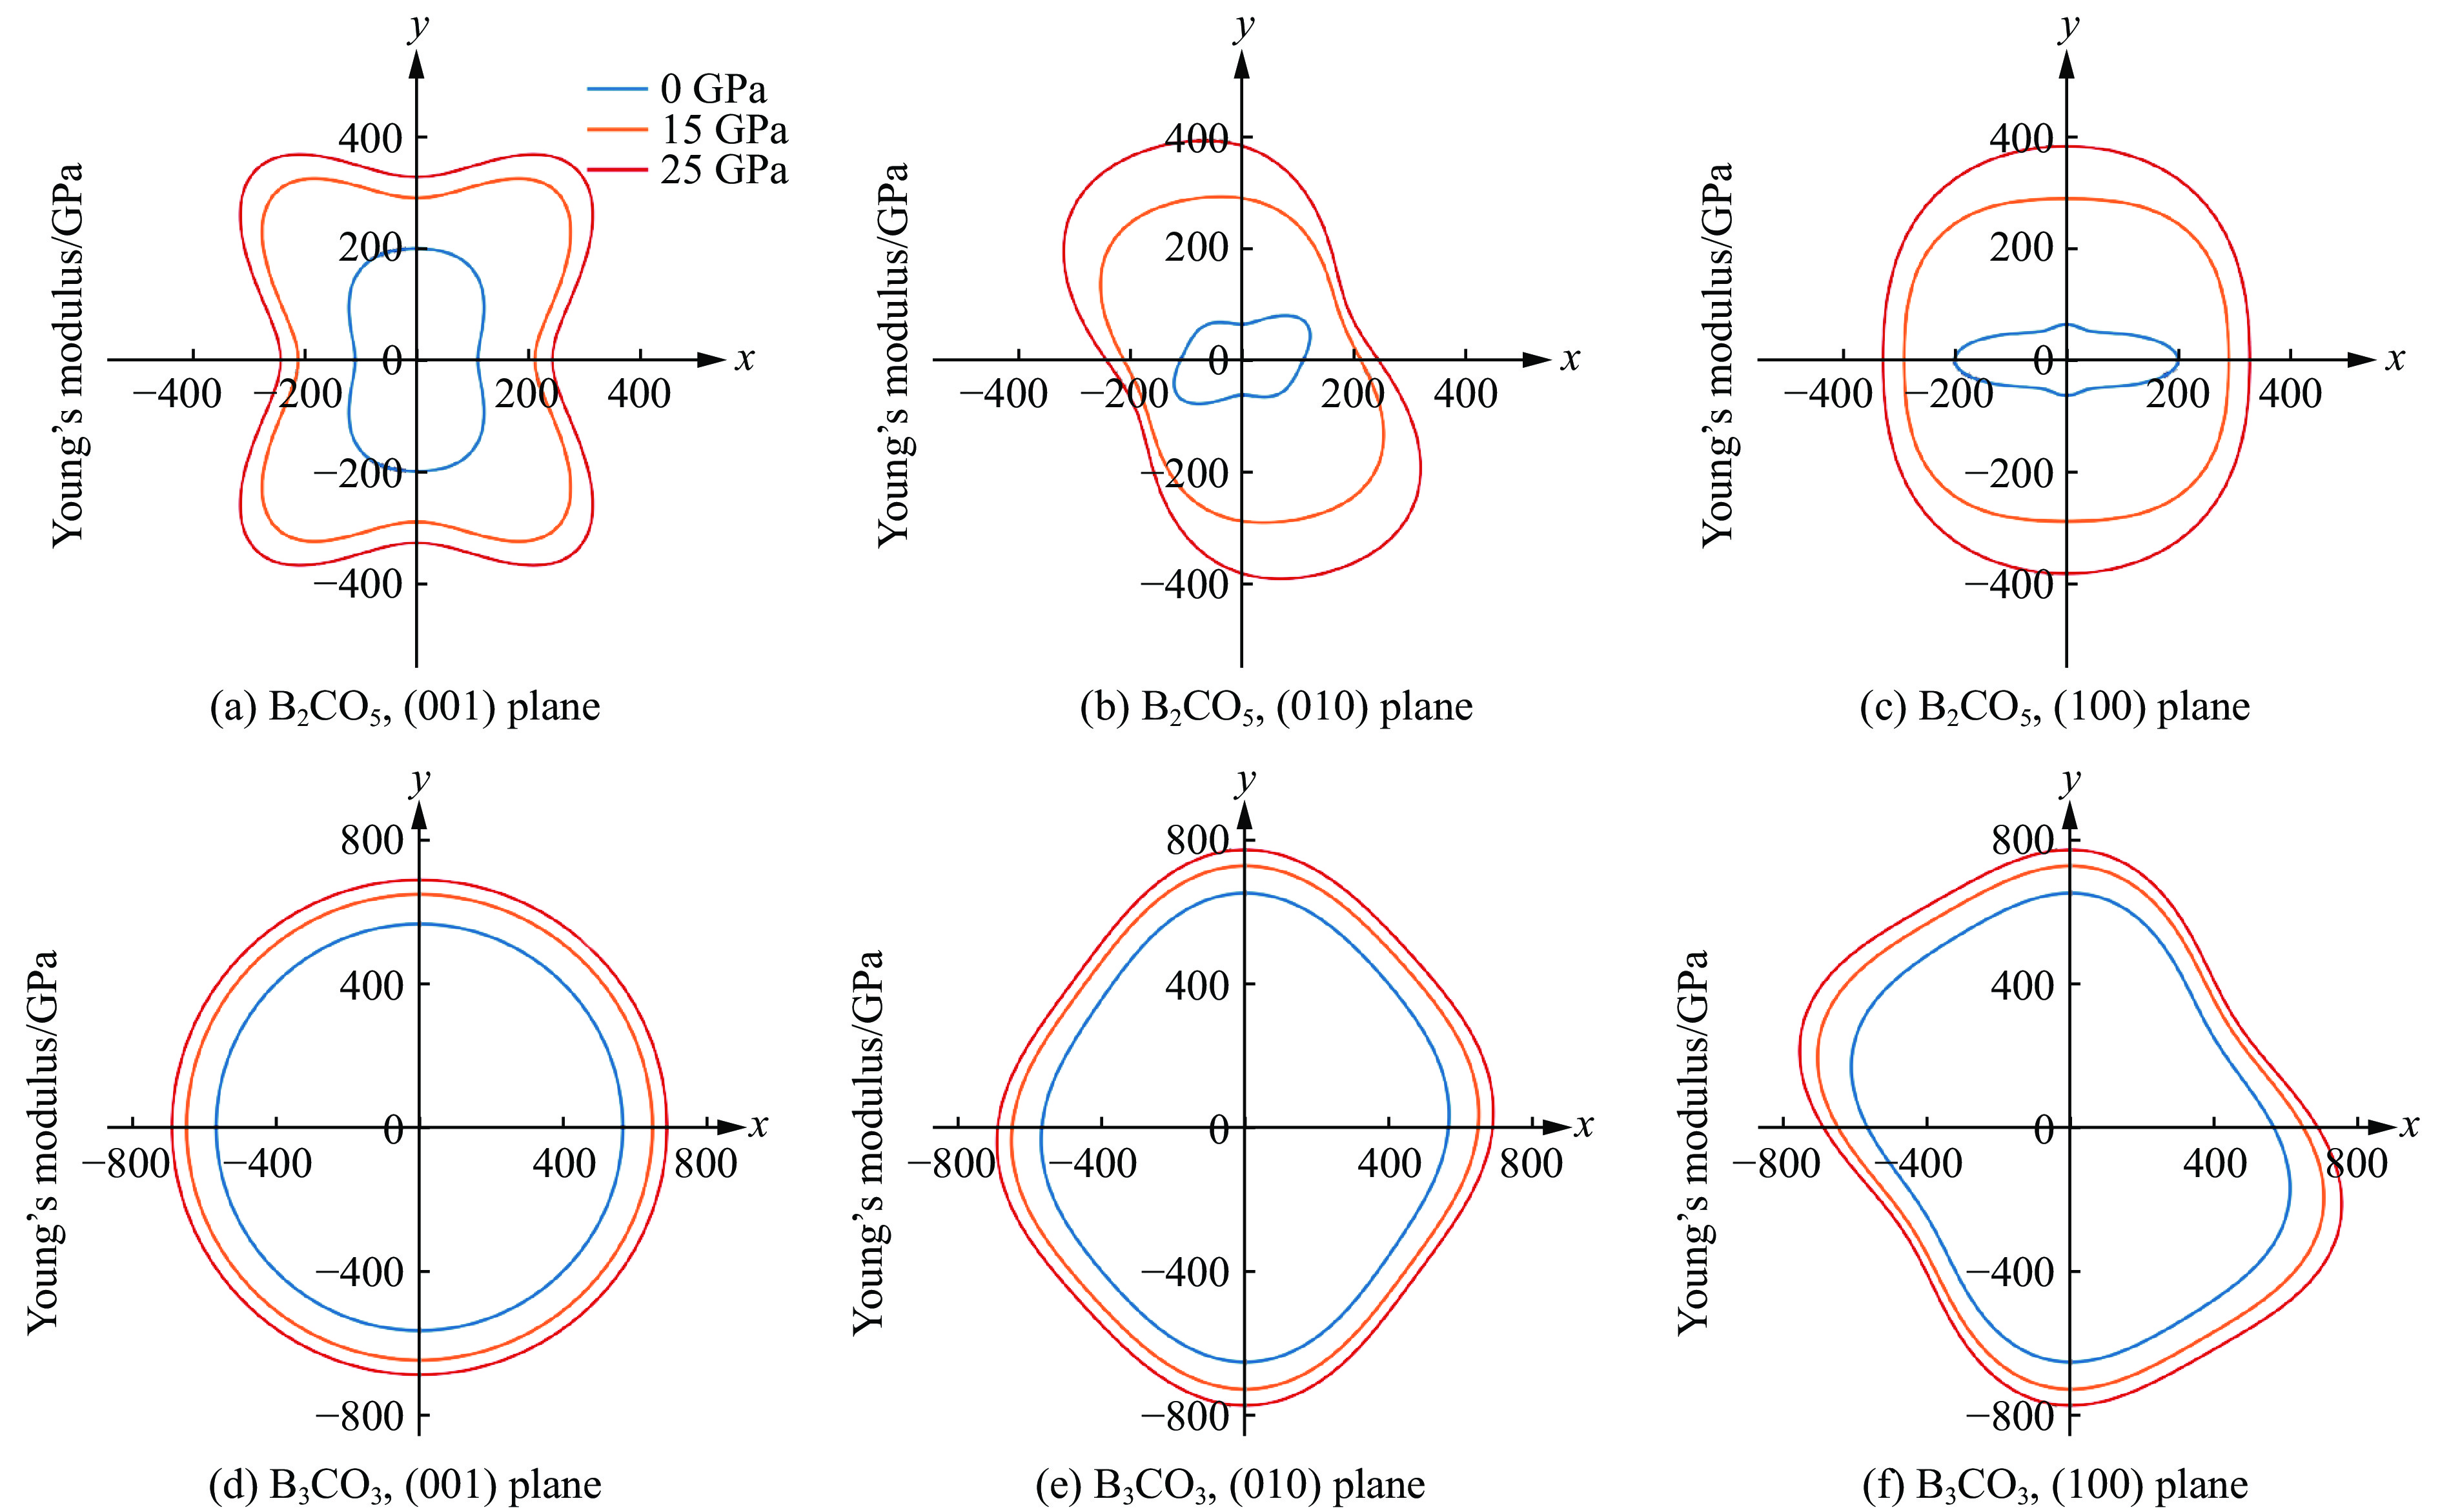
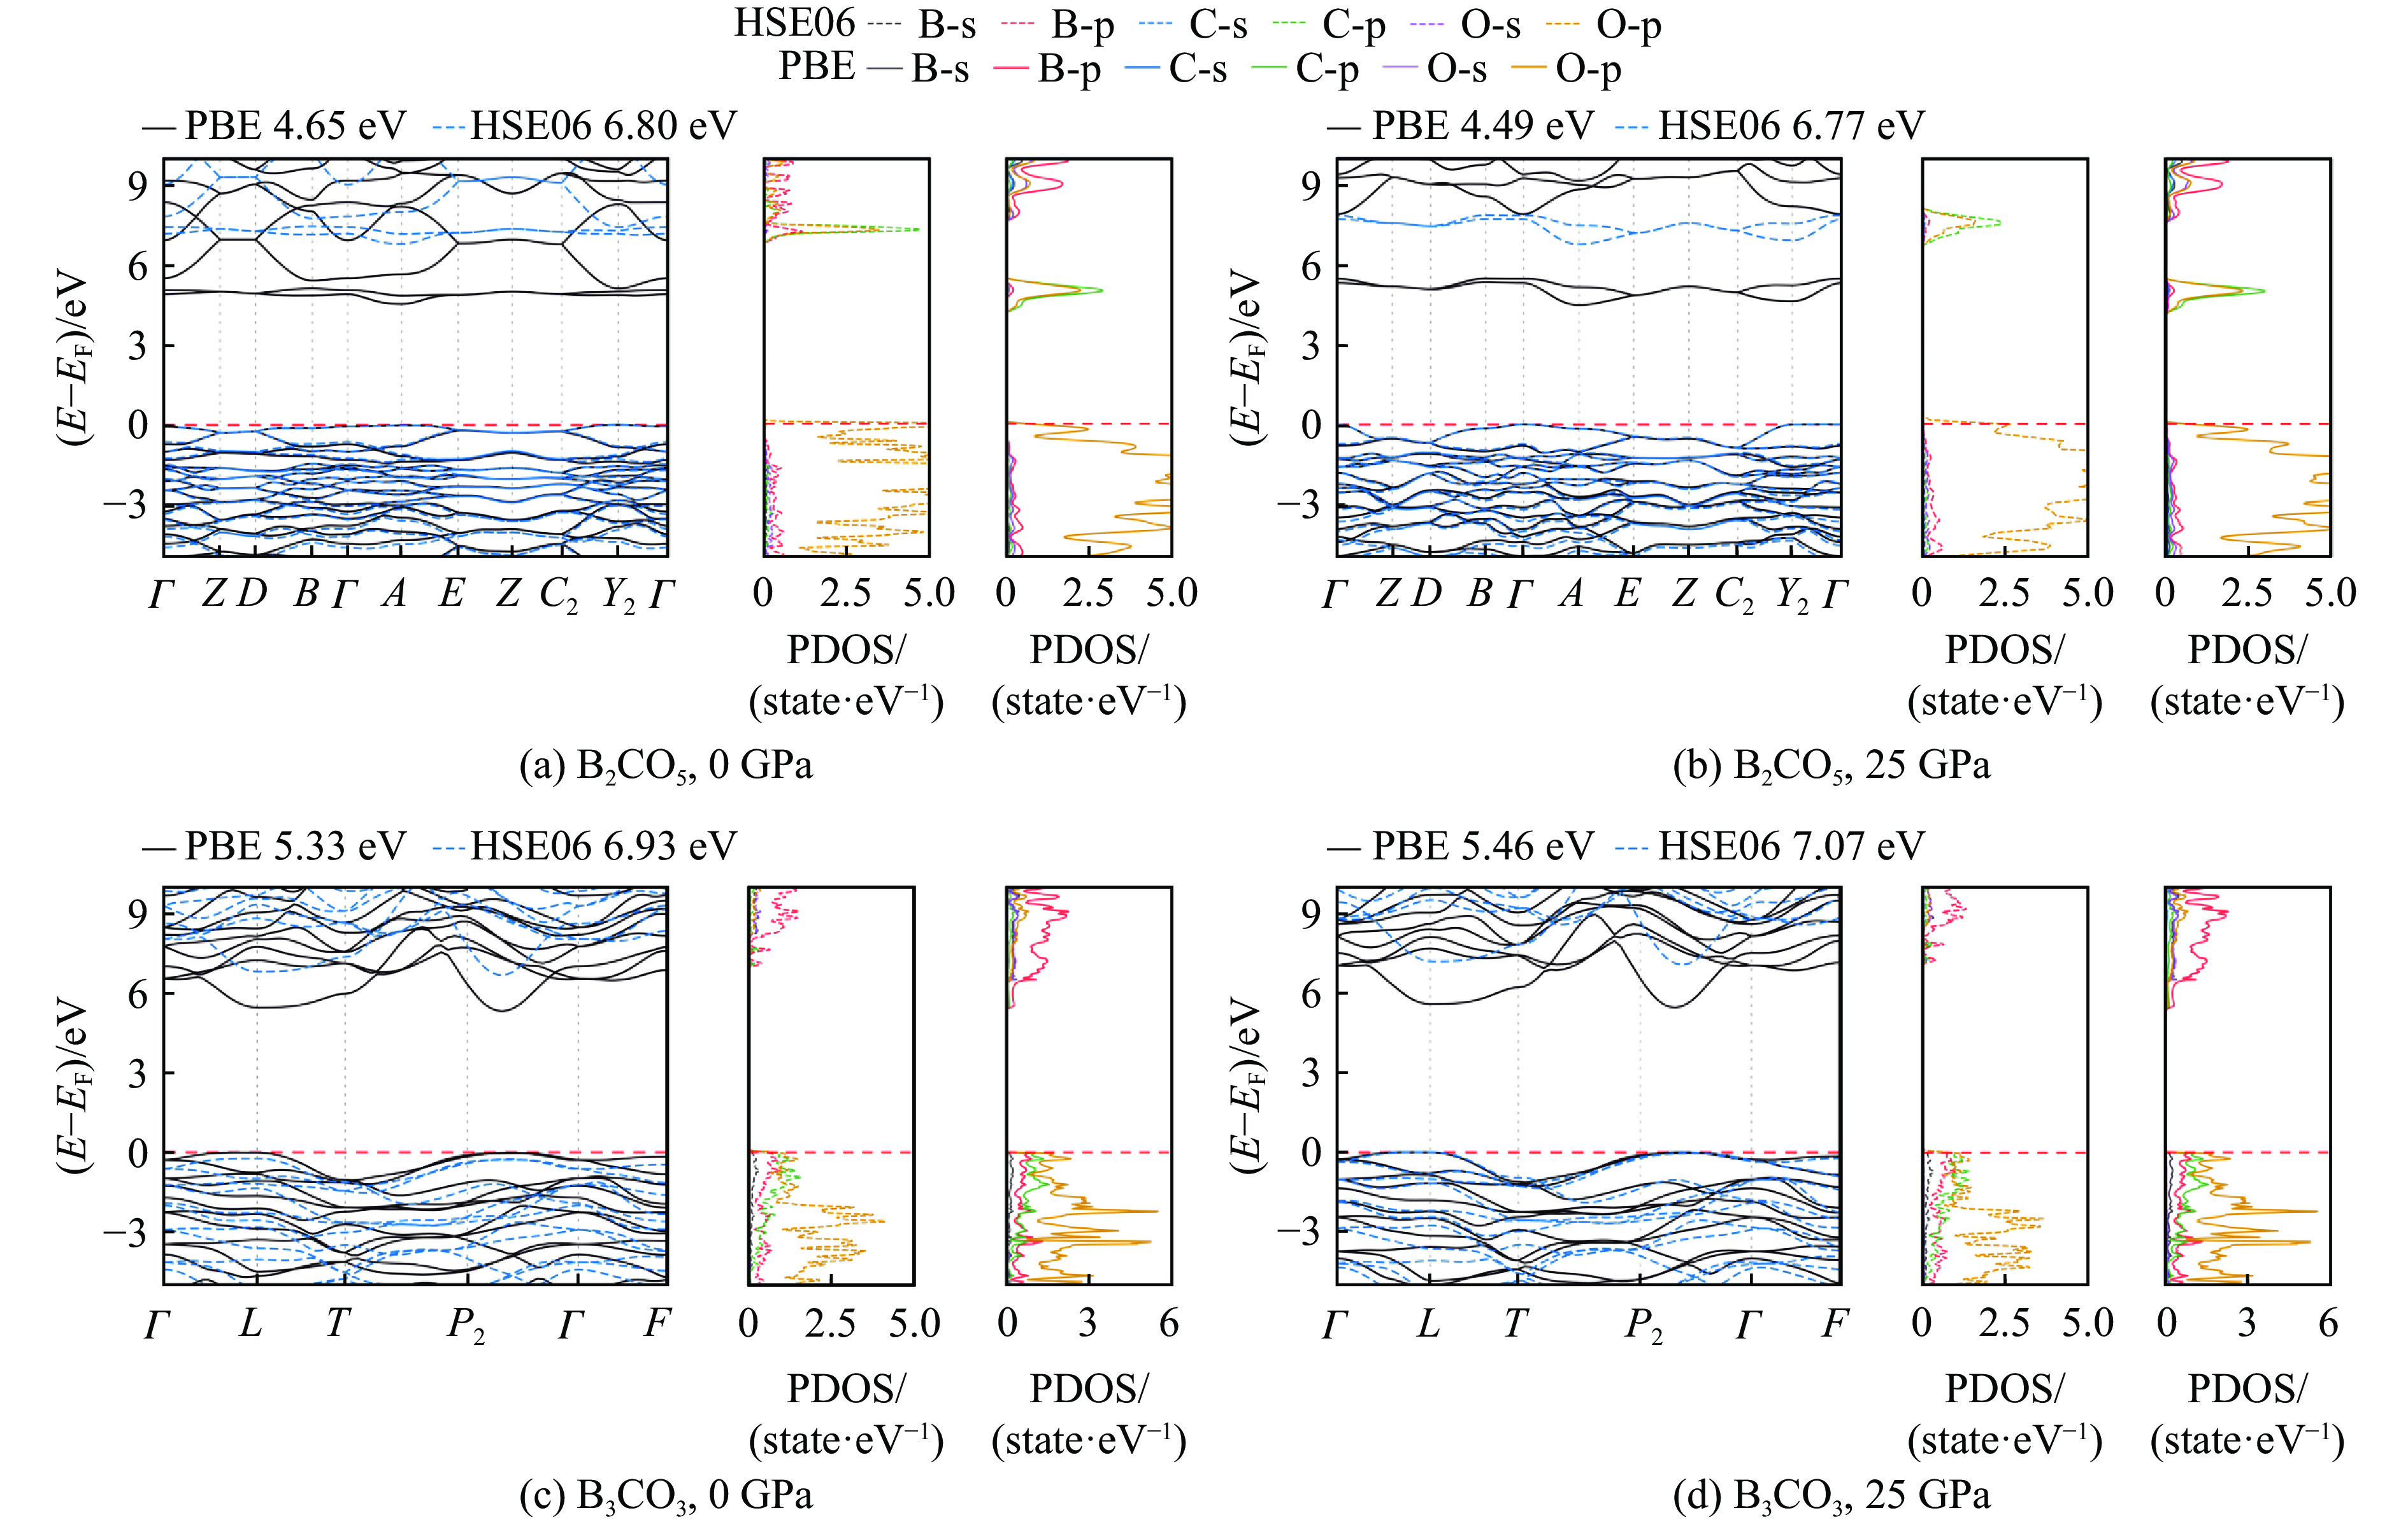
B-C-O compounds are considered one of the ideal systems for exploring and designing novel superhard materials due to the readily formed strong covalent bonds among the light elements B, C, and O. In this work, the CALYPSO structure prediction method, combined with first-principles calculations, was employed to systematically investigate the stability of ternary B-C-O compounds under high pressure. Two novel metastable compounds, B2CO5 and B3CO3, were identified. Among them, the formation energy of B2CO5 is only 6 meV/atom above the thermodynamic convex hull. Both B2CO5 and B3CO3 are non-diamond isoelectronic compounds and exhibit novel bonding configurations. The crystal structure of B2CO5 consists of sp2-hybridized planar triangular CO3 units and sp3-hybridized BO4 tetrahedra, which are interconnected through shared O atoms. In contrast, the crystal structure of B3CO3 contains sp3-hybridized BCO3 tetrahedral units. Mechanical property analysis reveals that B2CO5 exhibits pronounced elastic anisotropy along different crystallographic planes, whereas B3CO3 maintains elastic isotropy on the (001) plane. Hardness calculations indicate that B3CO3 has a Vickers hardness of approximately 36.4 GPa at 0 GPa, suggesting its potential for applications as a superhard material. Electronic structure calculations show that both B2CO5 and B3CO3 exhibit wide-band-gap semiconducting characteristics at 25 GPa, with PBE-calculated (Perdew-Burke-Ernzerhof-calculated) band gaps of 4.49 and 5.46 eV, respectively, while the corresponding values obtained using the HSE06 (Heyd-Scuseria-Ernzerhof 2006) hybrid functional are 6.77 and 7.07 eV. This study enriches the diversity of bonding configurations in B-C-O compounds and provides theoretical insights and data for the design of novel superhard materials under high-pressure conditions.

First-Principles Study of the Effects of Phase Transitions and Decomposition on the Lattice Thermal Conductivity of Calcite
HONG Zheng, ZHU Yongqiang, XIONG Yuanmeng, LU Cheng, HE Kaihua
2026, 40(11): 110111. doi: 10.11858/gywlxb.20251228
HTML PDF (127)
Abstract:

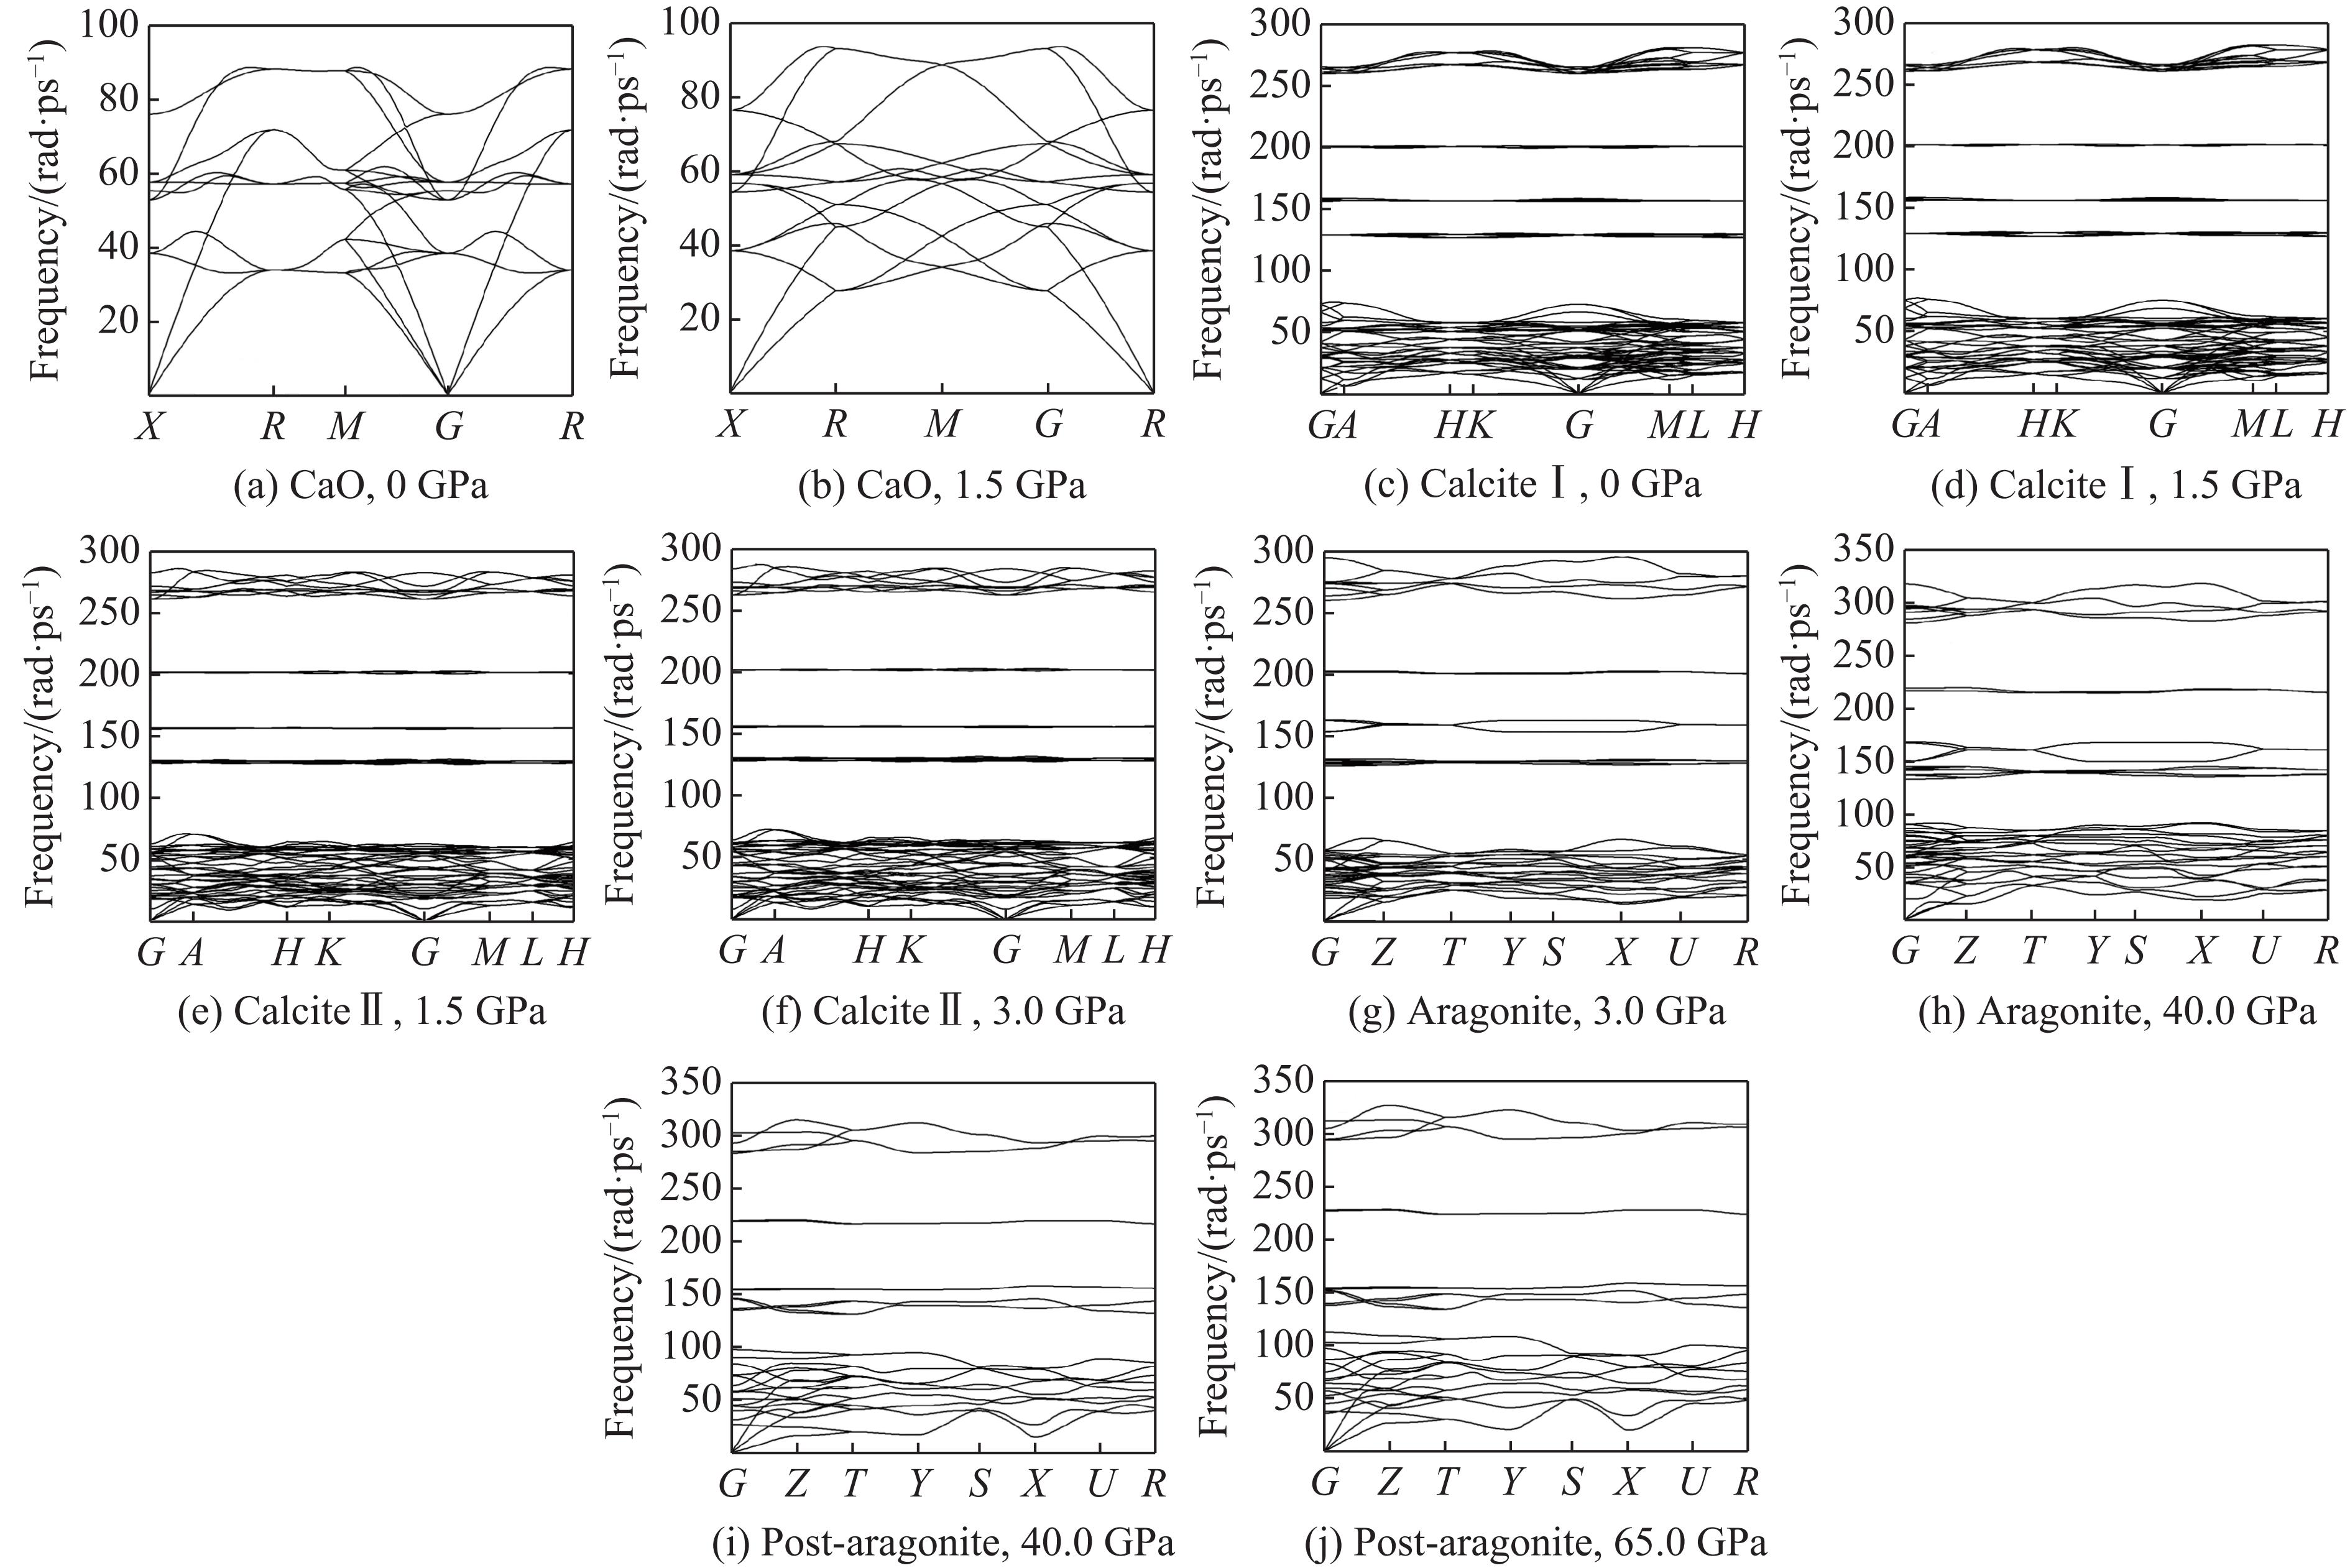
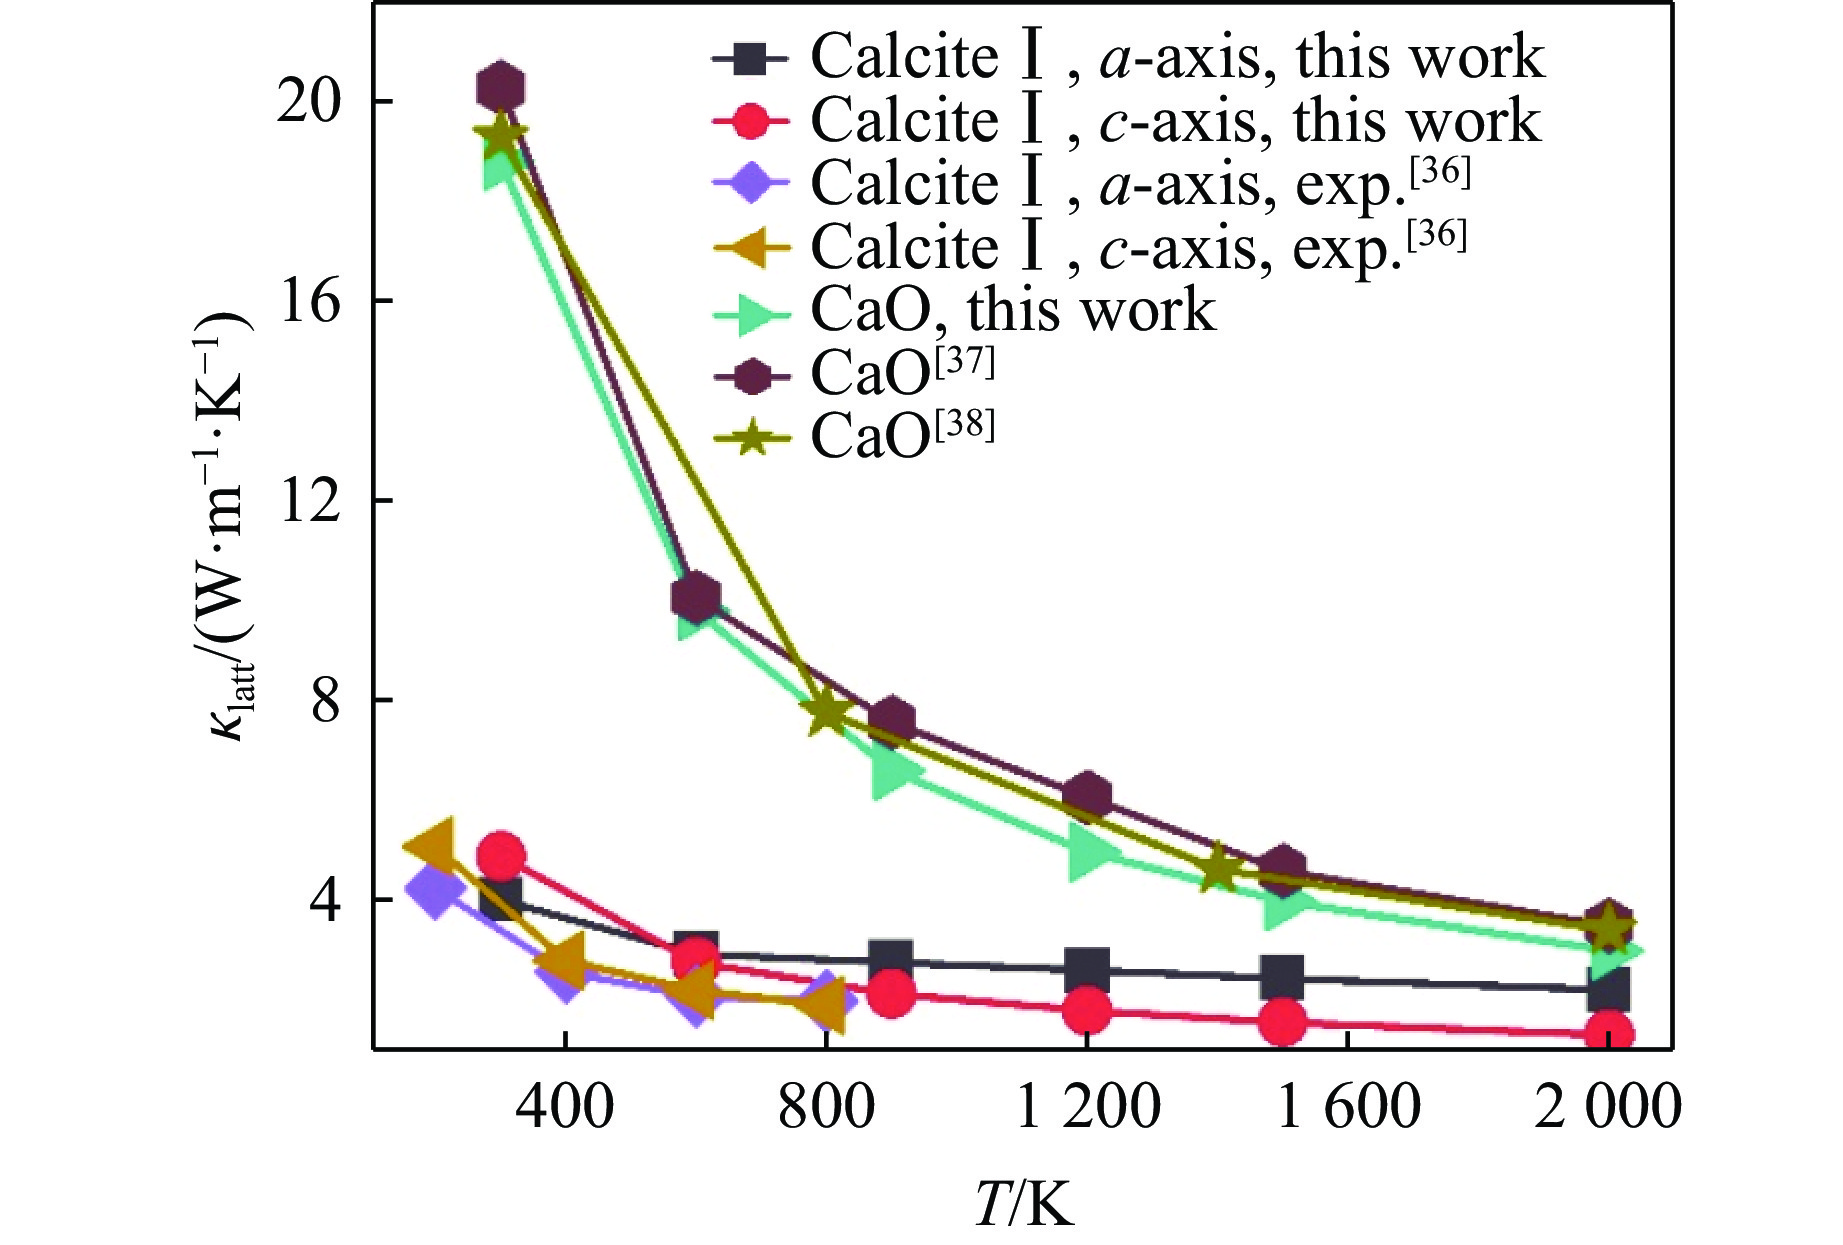
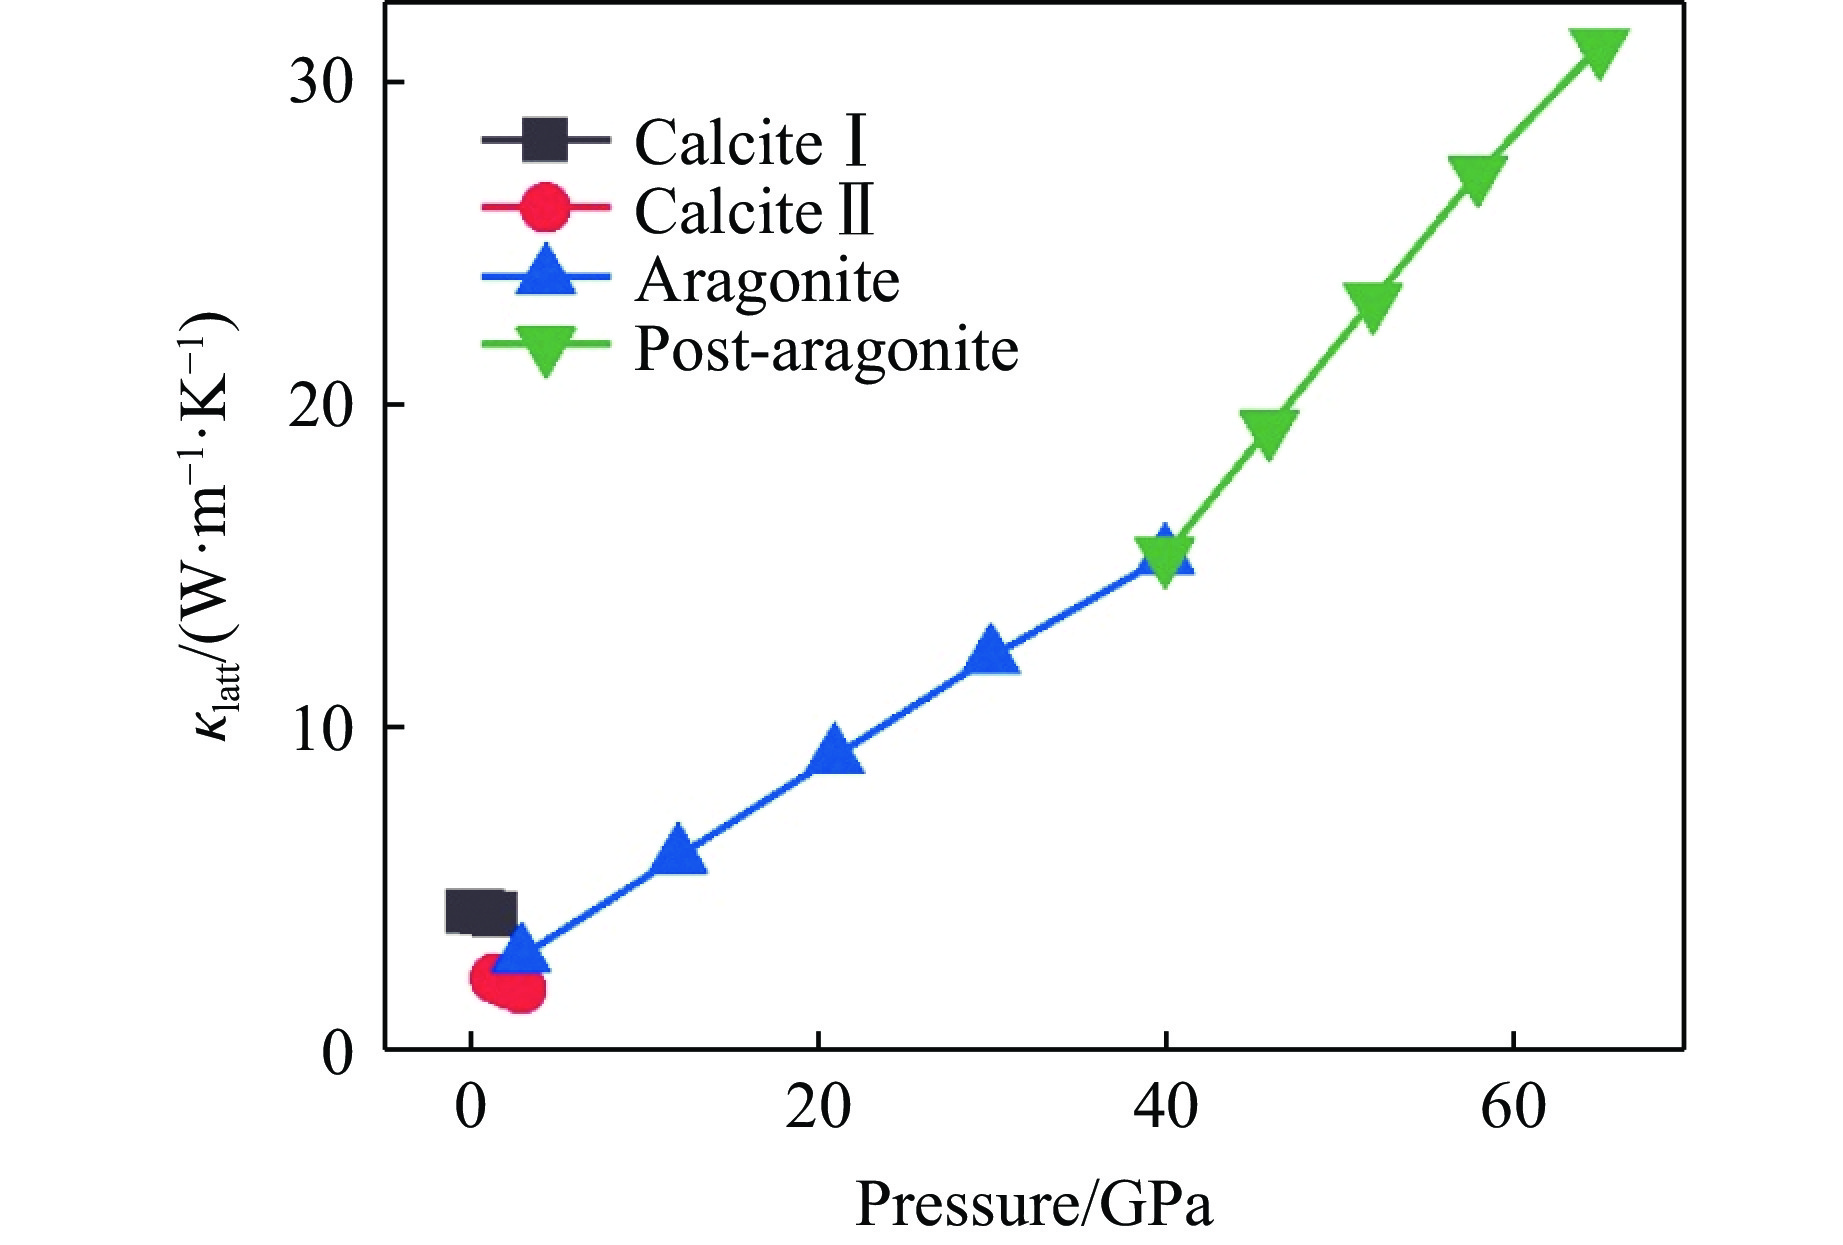
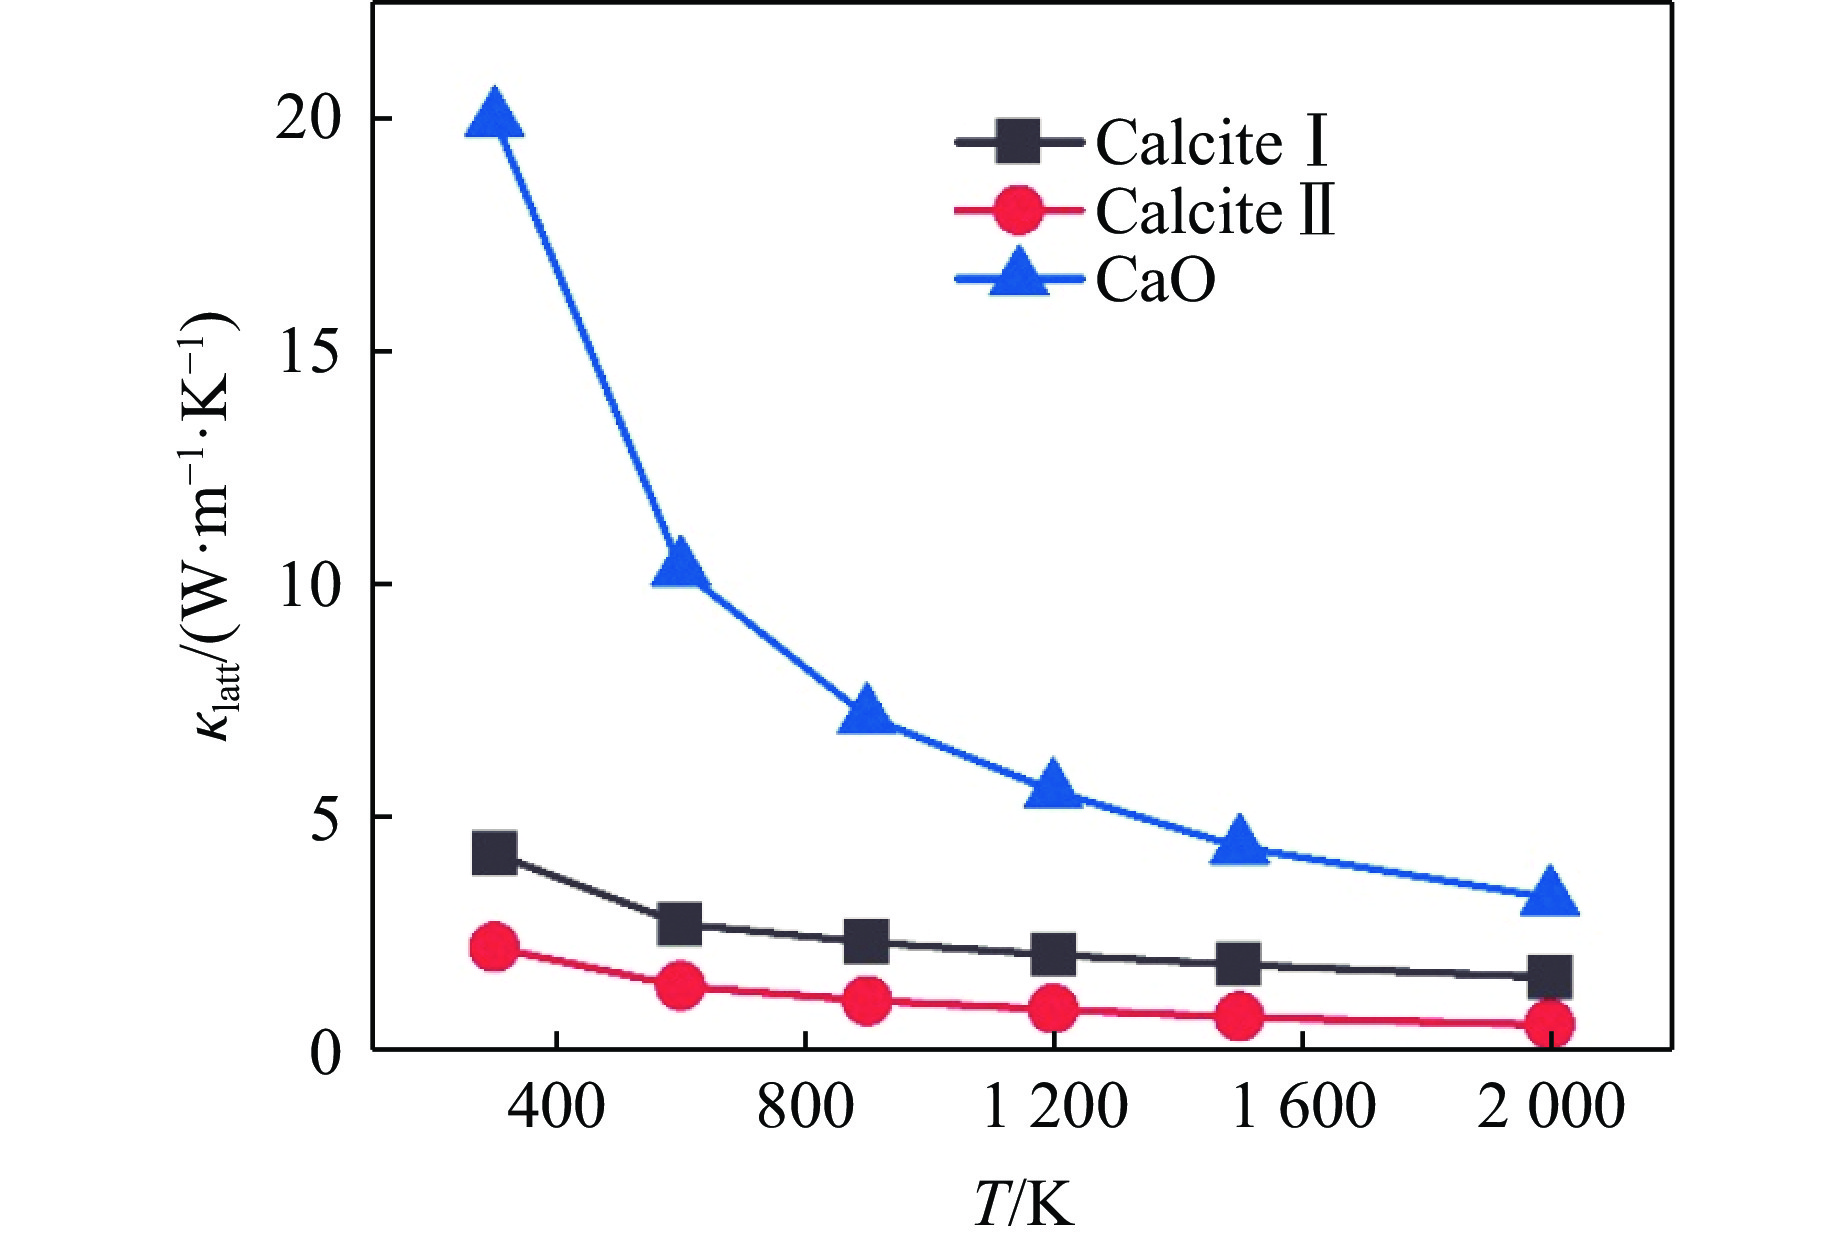
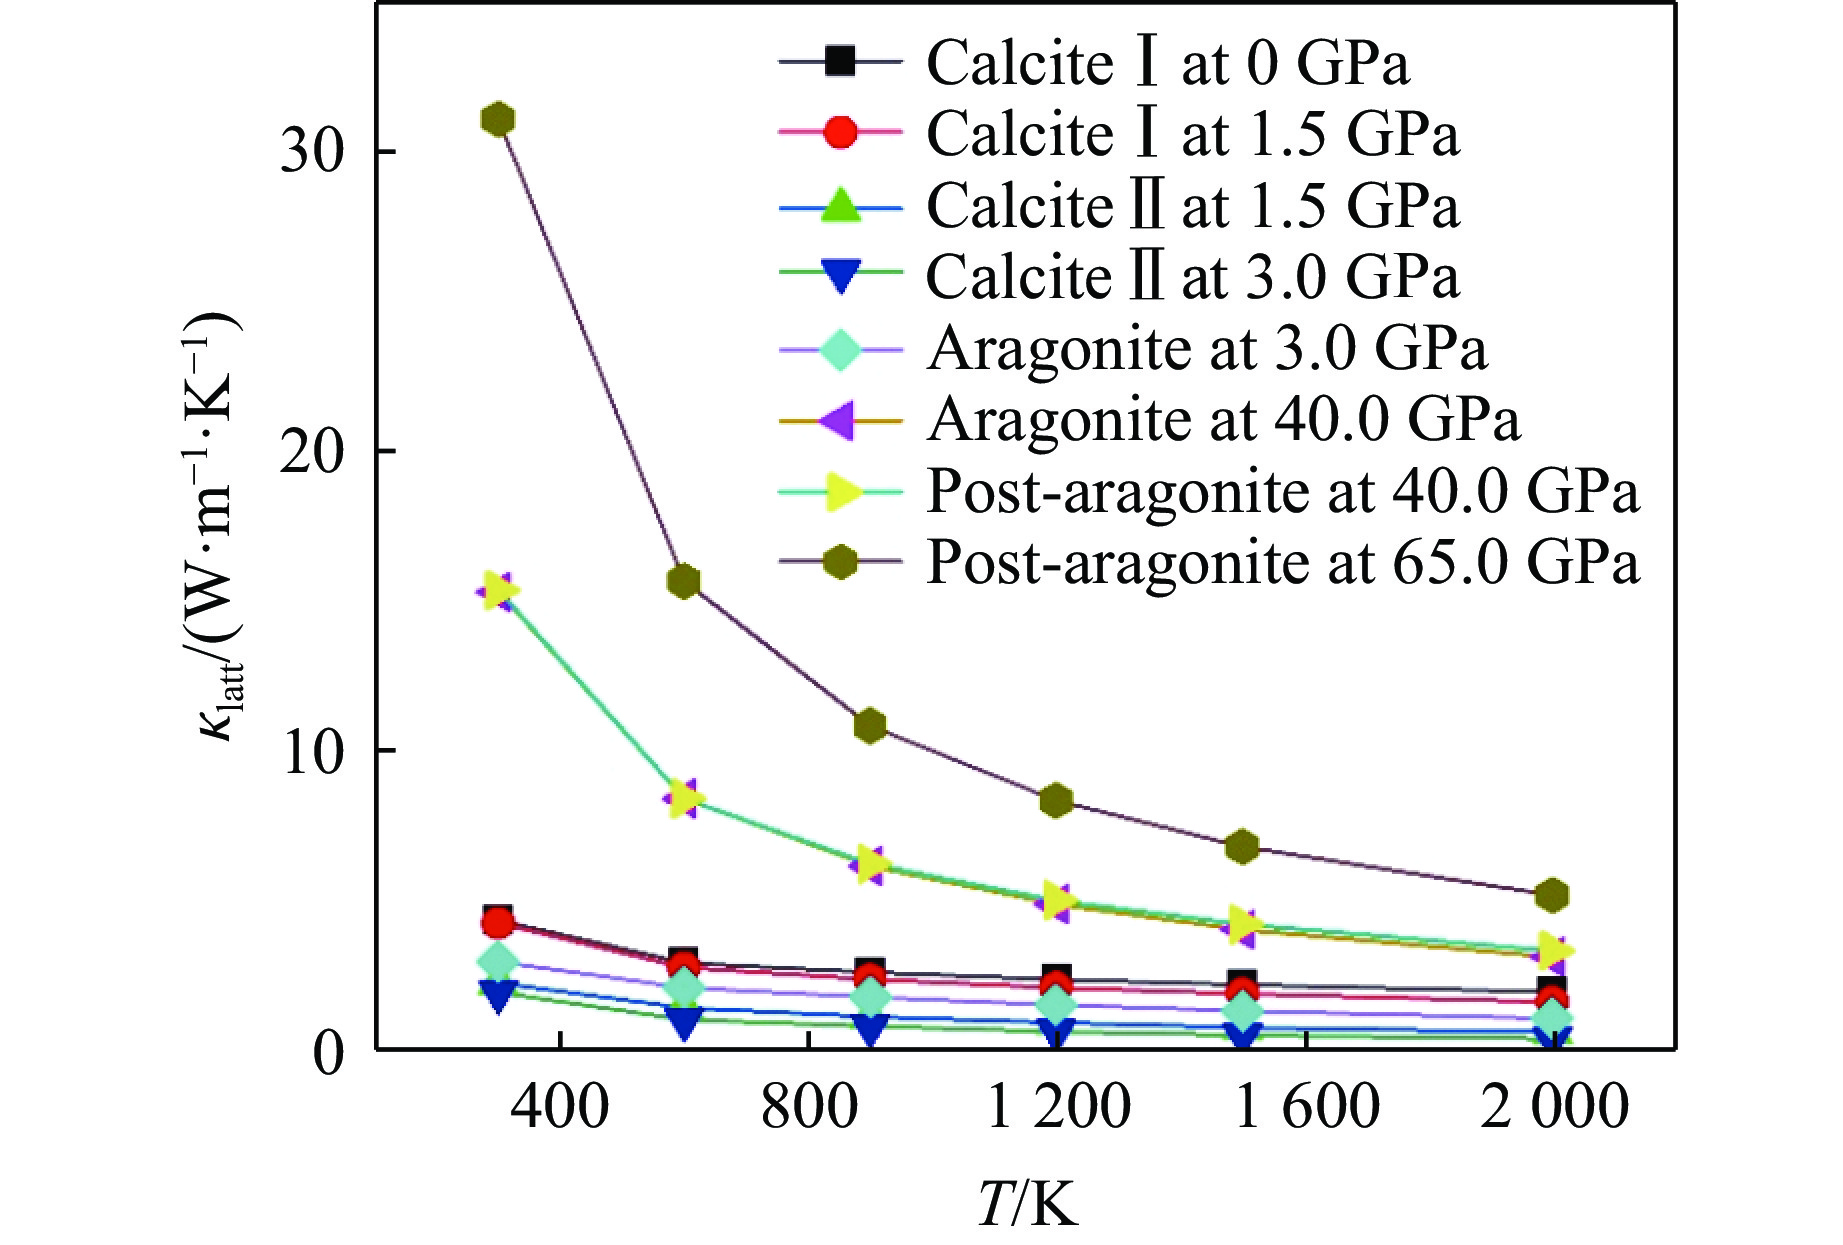
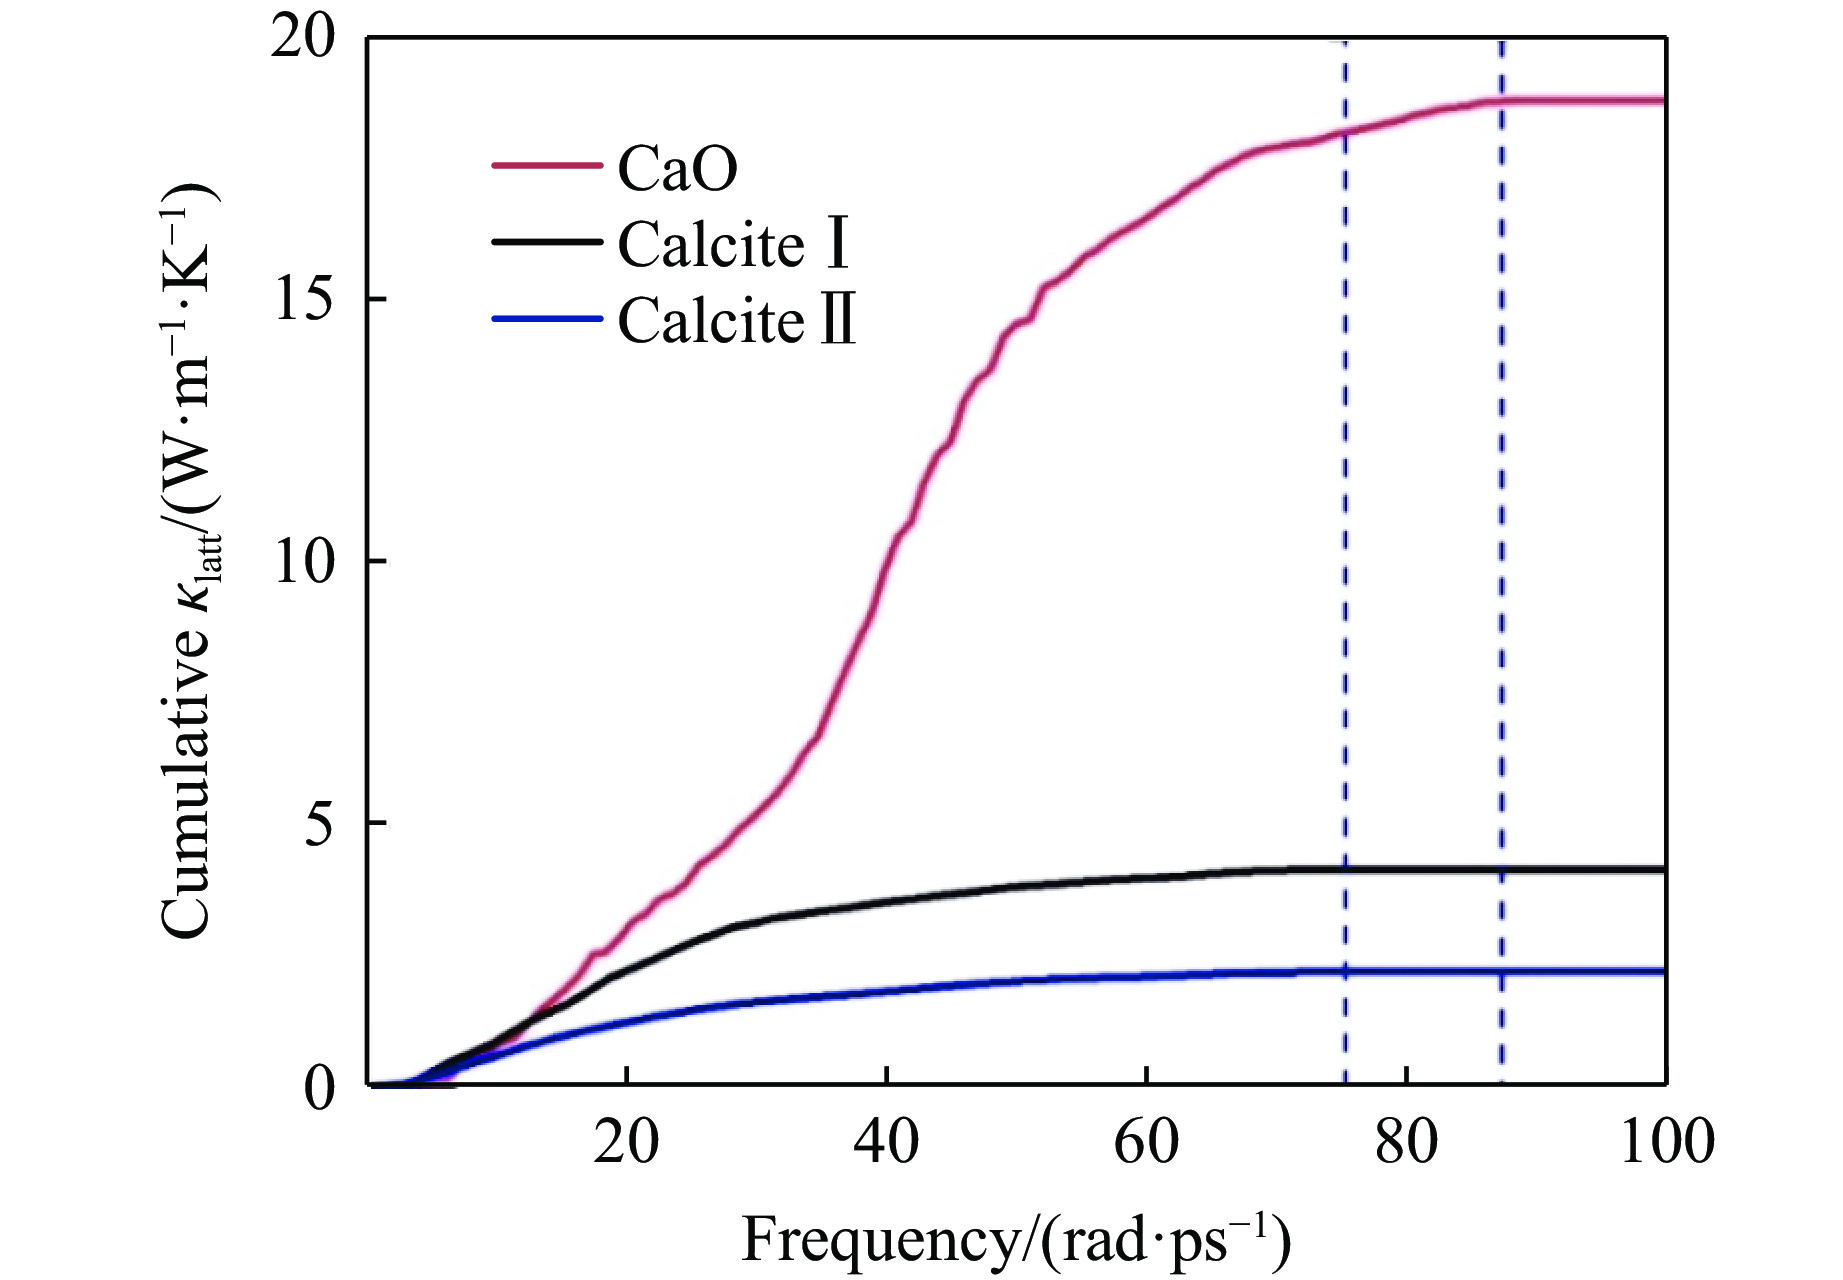
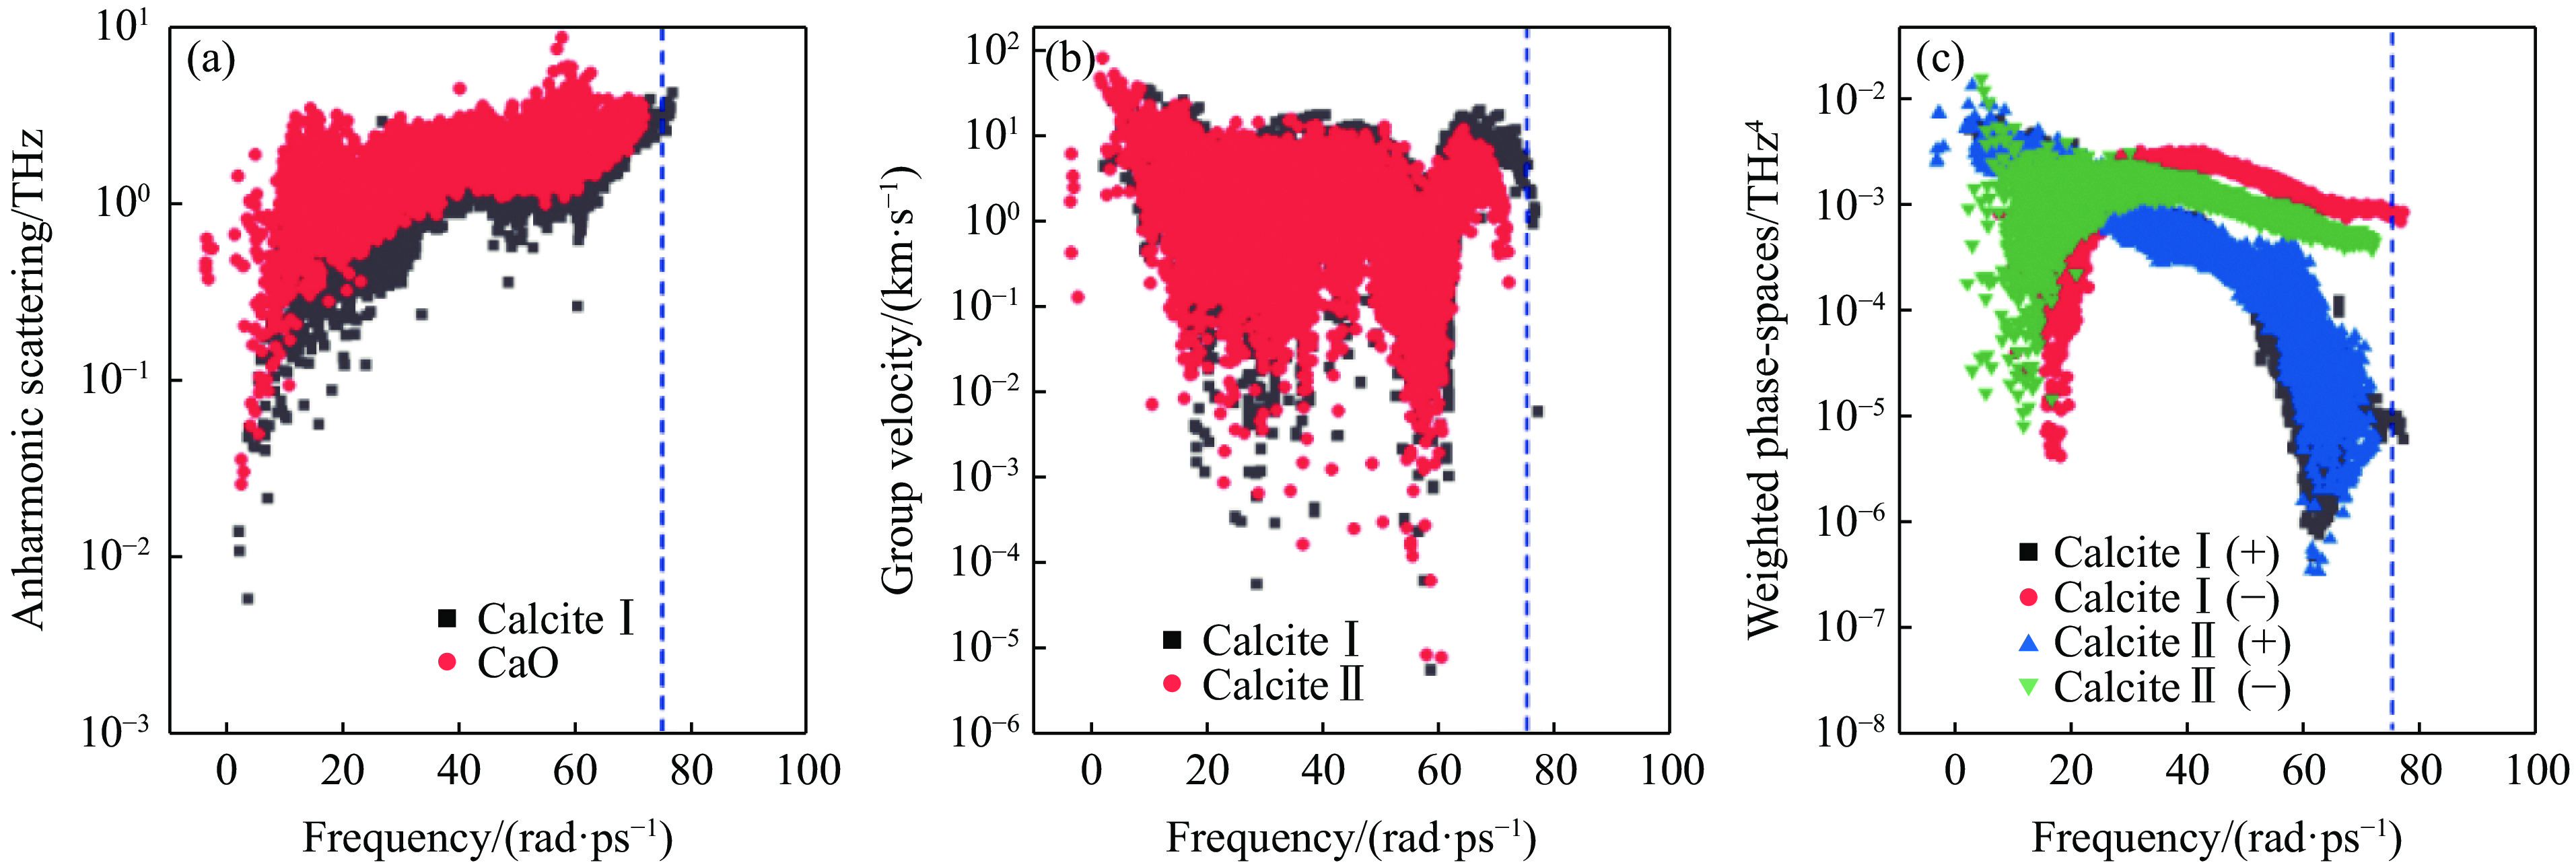
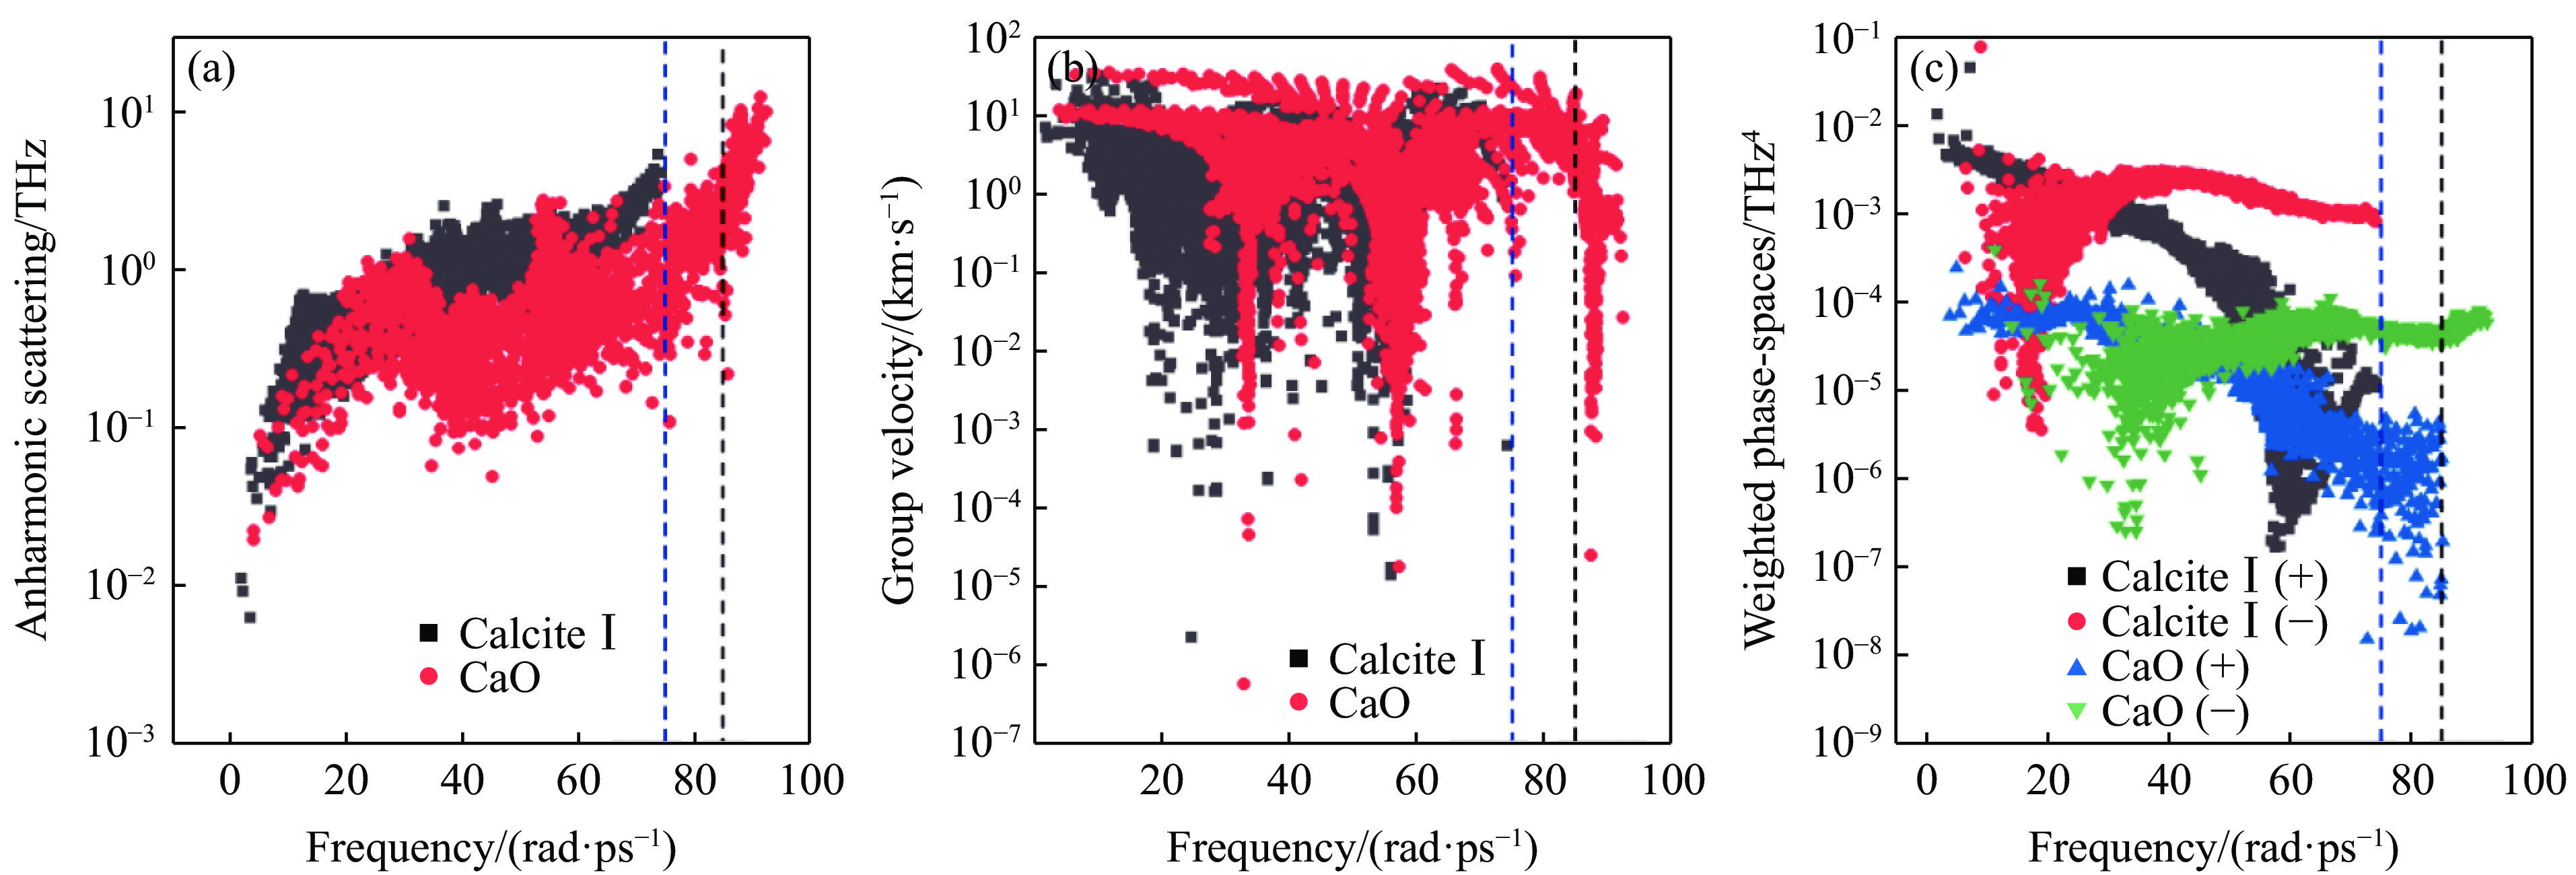
The lattice thermal conductivity of minerals plays a critical role in controlling heat flow and temperature distribution in the Earth’s interior. Calcite, primarily composed of calcium carbonate (CaCO3), can be subducted into the deep Earth and serves as an important carbon source. As pressure and temperature conditions change with depth, CaCO3 undergoes phase transitions and thermal decomposition, which significantly affect its physical properties. In this study, we investigate the effects on thermal conductivity of calcite induced phase transitions and thermal decomposition using first-principles calculations combined with lattice dynamics. Our results show that the calcite Ⅰ → calcite Ⅱ phase transition leads to a reduction in thermal conductivity, whereas subsequent phase transitions at higher pressures result in increase. The thermal conductivity of aragonite and post-aragonite increases nearly linearly with pressure increasing, and the latter exhibiting a stronger pressure dependence. Upon thermal decomposition, the CaO exhibits significantly higher thermal conductivity than that of calcite, which may enhance local heat transfer. Analysis of relevant thermodynamic parameters indicates that the changes in thermal conductivity induced by phase transitions and decomposition are collectively determined by phonon group velocity and anharmonic scattering rates.